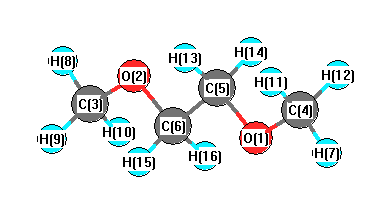
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3070 |
3020 |
0.00 |
|
|
|
| 2 |
Ag |
2914 |
2866 |
0.00 |
|
|
|
| 3 |
Ag |
2899 |
2851 |
0.00 |
|
|
|
| 4 |
Ag |
1510 |
1485 |
0.00 |
|
|
|
| 5 |
Ag |
1484 |
1460 |
0.00 |
|
|
|
| 6 |
Ag |
1456 |
1432 |
0.00 |
|
|
|
| 7 |
Ag |
1412 |
1388 |
0.00 |
|
|
|
| 8 |
Ag |
1210 |
1190 |
0.00 |
|
|
|
| 9 |
Ag |
1147 |
1128 |
0.00 |
|
|
|
| 10 |
Ag |
1064 |
1046 |
0.00 |
|
|
|
| 11 |
Ag |
1003 |
987 |
0.00 |
|
|
|
| 12 |
Ag |
390 |
384 |
0.00 |
|
|
|
| 13 |
Ag |
326 |
321 |
0.00 |
|
|
|
| 14 |
Au |
2955 |
2906 |
11.71 |
|
|
|
| 15 |
Au |
2949 |
2901 |
255.91 |
|
|
|
| 16 |
Au |
1462 |
1438 |
11.64 |
|
|
|
| 17 |
Au |
1206 |
1186 |
10.42 |
|
|
|
| 18 |
Au |
1150 |
1131 |
1.41 |
|
|
|
| 19 |
Au |
821 |
808 |
0.16 |
|
|
|
| 20 |
Au |
248 |
244 |
7.10 |
|
|
|
| 21 |
Au |
98 |
96 |
6.68 |
|
|
|
| 22 |
Au |
71 |
70 |
2.95 |
|
|
|
| 23 |
Bg |
2953 |
2904 |
0.00 |
|
|
|
| 24 |
Bg |
2924 |
2876 |
0.00 |
|
|
|
| 25 |
Bg |
1462 |
1438 |
0.00 |
|
|
|
| 26 |
Bg |
1263 |
1242 |
0.00 |
|
|
|
| 27 |
Bg |
1169 |
1149 |
0.00 |
|
|
|
| 28 |
Bg |
1128 |
1109 |
0.00 |
|
|
|
| 29 |
Bg |
235 |
231 |
0.00 |
|
|
|
| 30 |
Bg |
128 |
126 |
0.00 |
|
|
|
| 31 |
Bu |
3070 |
3020 |
60.89 |
|
|
|
| 32 |
Bu |
2915 |
2867 |
160.44 |
|
|
|
| 33 |
Bu |
2902 |
2854 |
80.52 |
|
|
|
| 34 |
Bu |
1517 |
1492 |
1.35 |
|
|
|
| 35 |
Bu |
1486 |
1462 |
25.83 |
|
|
|
| 36 |
Bu |
1450 |
1426 |
5.57 |
|
|
|
| 37 |
Bu |
1328 |
1306 |
29.19 |
|
|
|
| 38 |
Bu |
1188 |
1168 |
133.03 |
|
|
|
| 39 |
Bu |
1149 |
1130 |
228.95 |
|
|
|
| 40 |
Bu |
957 |
941 |
57.47 |
|
|
|
| 41 |
Bu |
495 |
487 |
3.49 |
|
|
|
| 42 |
Bu |
143 |
141 |
3.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30352.8 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 29851.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.418 |
|
|
|
| 2 |
O |
-0.418 |
|
|
|
| 3 |
C |
-0.247 |
|
|
|
| 4 |
C |
-0.247 |
|
|
|
| 5 |
C |
-0.074 |
|
|
|
| 6 |
C |
-0.074 |
|
|
|
| 7 |
H |
0.170 |
|
|
|
| 8 |
H |
0.170 |
|
|
|
| 9 |
H |
0.142 |
|
|
|
| 10 |
H |
0.142 |
|
|
|
| 11 |
H |
0.142 |
|
|
|
| 12 |
H |
0.142 |
|
|
|
| 13 |
H |
0.142 |
|
|
|
| 14 |
H |
0.142 |
|
|
|
| 15 |
H |
0.142 |
|
|
|
| 16 |
H |
0.142 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.113 |
-3.800 |
0.000 |
| y |
-3.800 |
-31.659 |
0.000 |
| z |
0.000 |
0.000 |
-38.198 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.184 |
-3.800 |
0.000 |
| y |
-3.800 |
6.996 |
0.000 |
| z |
0.000 |
0.000 |
-2.812 |
|
| Polar |
|---|
| 3z2-r2 | -5.624 |
|---|
| x2-y2 | -7.454 |
|---|
| xy | -3.800 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.991 |
-0.194 |
0.000 |
| y |
-0.194 |
10.208 |
0.000 |
| z |
0.000 |
0.000 |
6.898 |
<r2> (average value of r
2) Å
2
| <r2> |
271.216 |
| (<r2>)1/2 |
16.469 |