return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (cyclohexene)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-233.696029
Energy at 298.15K-233.707852
HF Energy-233.019413
Nuclear repulsion energy237.791364
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3297 3046 41.82      
2 A 3204 2960 48.11      
3 A 3184 2941 1.26      
4 A 3157 2916 19.10      
5 A 3141 2901 0.36      
6 A 1843 1703 1.49      
7 A 1607 1484 1.00      
8 A 1581 1460 0.20      
9 A 1481 1368 0.16      
10 A 1475 1363 0.13      
11 A 1347 1244 0.81      
12 A 1325 1224 0.08      
13 A 1234 1140 0.06      
14 A 1172 1083 0.04      
15 A 1142 1055 0.08      
16 A 1061 980 0.00      
17 A 966 892 0.88      
18 A 881 814 0.36      
19 A 871 804 0.61      
20 A 523 483 0.01      
21 A 422 390 0.01      
22 A 294 271 0.05      
23 B 3273 3023 8.89      
24 B 3210 2965 64.40      
25 B 3183 2940 77.17      
26 B 3157 2916 16.93      
27 B 3140 2901 64.54      
28 B 1594 1473 3.79      
29 B 1589 1468 5.14      
30 B 1510 1394 0.09      
31 B 1477 1364 2.21      
32 B 1436 1326 1.24      
33 B 1368 1264 2.44      
34 B 1231 1137 5.51      
35 B 1111 1026 2.68      
36 B 1077 995 1.27      
37 B 996 920 4.99      
38 B 940 868 3.83      
39 B 776 717 12.80      
40 B 693 640 20.12      
41 B 476 440 0.95      
42 B 177 163 0.16      

Unscaled Zero Point Vibrational Energy (zpe) 33811.4 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 31231.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.15950 0.15299 0.08612

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.006 -0.663 1.297
C2 0.006 0.663 1.297
C3 0.000 1.494 0.043
C4 0.000 -1.494 0.043
C5 -0.376 0.663 -1.181
C6 0.376 -0.663 -1.181
H7 -0.022 -1.195 2.240
H8 0.022 1.195 2.240
H9 0.986 1.945 -0.099
H10 -0.986 -1.945 -0.099
H11 -0.694 2.328 0.159
H12 0.694 -2.328 0.159
H13 -1.449 0.466 -1.166
H14 1.449 -0.466 -1.166
H15 -0.170 1.223 -2.093
H16 0.170 -1.223 -2.093

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.32692.49541.50412.83542.50751.08272.08433.12032.13353.27382.13453.07052.86813.88373.4411
C21.32691.50412.49542.50752.83542.08431.08272.13353.12032.13453.27382.86813.07053.44113.8837
C32.49541.50412.98791.52642.50893.47252.21731.09373.58031.09133.88642.14962.72112.16043.4610
C41.50412.49542.98792.50891.52642.21733.47253.58031.09373.88641.09132.72112.14963.46102.1604
C52.83542.50751.52642.50891.52473.90963.48512.16062.88922.16093.44831.09182.14591.09002.1656
C62.50752.83542.50891.52641.52473.48513.90962.88922.16063.44832.16092.14591.09182.16561.0900
H71.08272.08433.47252.21733.90963.48512.39084.04312.63874.14682.47514.04993.78174.96504.3379
H82.08431.08272.21733.47253.48513.90962.39082.63874.04312.47514.14683.78174.04994.33794.9650
H93.12032.13351.09373.58032.16062.88924.04312.63874.36131.74264.29083.04292.67692.41573.8319
H102.13353.12033.58031.09372.88922.16062.63874.04314.36134.29081.74262.67693.04293.83192.4157
H113.27382.13451.09133.88642.16093.44834.14682.47511.74264.29084.85882.40763.76262.56324.2936
H122.13453.27383.88641.09133.44832.16092.47514.14684.29081.74264.85883.76262.40764.29362.5632
H133.07052.86812.14962.72111.09182.14594.04993.78173.04292.67692.40763.76263.04441.75212.5167
H142.86813.07052.72112.14962.14591.09183.78174.04992.67693.04293.76262.40763.04442.51671.7521
H153.88373.44112.16043.46101.09002.16564.96504.33792.41573.83192.56324.29361.75212.51672.4703
H163.44113.88373.46102.16042.16561.09004.33794.96503.83192.41574.29362.56322.51671.75212.4703

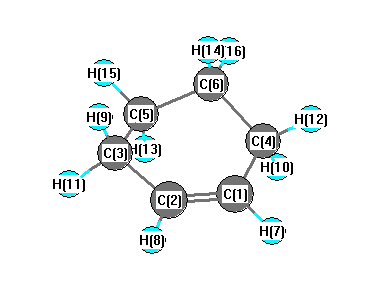
picture of cyclohexene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.513 C1 C2 H8 119.423
C1 C4 C6 111.671 C1 C4 H10 109.414
C1 C4 H12 109.628 C2 C1 C4 123.513
C2 C1 H7 119.423 C2 C3 C5 111.671
C2 C3 H9 109.414 C2 C3 H11 109.628
C3 C2 H8 117.064 C3 C5 C6 110.629
C3 C5 H13 109.247 C3 C5 H15 110.207
C4 C1 H7 117.064 C4 C6 C5 110.629
C4 C6 H14 109.247 C4 C6 H16 110.207
C5 C3 H9 110.001 C5 C3 H11 110.170
C5 C6 H14 109.080 C5 C6 H16 110.739
C6 C4 H10 110.001 C6 C4 H12 110.170
C6 C5 H13 109.080 C6 C5 H15 110.739
H9 C3 H11 105.786 H10 C4 H12 105.786
H13 C5 H15 106.847 H14 C6 H16 106.847
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability