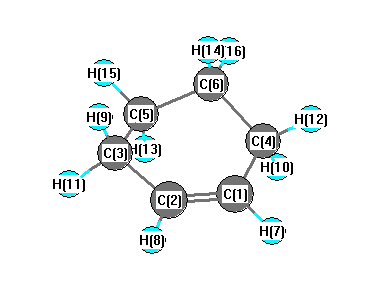
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3182 |
3044 |
47.40 |
|
|
|
| 2 |
A |
3094 |
2960 |
49.95 |
|
|
|
| 3 |
A |
3066 |
2933 |
1.33 |
|
|
|
| 4 |
A |
3043 |
2911 |
19.46 |
|
|
|
| 5 |
A |
3018 |
2888 |
0.01 |
|
|
|
| 6 |
A |
1747 |
1671 |
2.67 |
|
|
|
| 7 |
A |
1526 |
1460 |
1.49 |
|
|
|
| 8 |
A |
1497 |
1432 |
0.03 |
|
|
|
| 9 |
A |
1401 |
1341 |
0.28 |
|
|
|
| 10 |
A |
1392 |
1332 |
0.32 |
|
|
|
| 11 |
A |
1283 |
1227 |
0.73 |
|
|
|
| 12 |
A |
1261 |
1206 |
0.18 |
|
|
|
| 13 |
A |
1172 |
1121 |
0.01 |
|
|
|
| 14 |
A |
1104 |
1056 |
0.01 |
|
|
|
| 15 |
A |
1090 |
1043 |
0.10 |
|
|
|
| 16 |
A |
1000 |
957 |
0.01 |
|
|
|
| 17 |
A |
927 |
887 |
0.74 |
|
|
|
| 18 |
A |
843 |
806 |
0.03 |
|
|
|
| 19 |
A |
831 |
795 |
1.33 |
|
|
|
| 20 |
A |
501 |
479 |
0.04 |
|
|
|
| 21 |
A |
401 |
384 |
0.00 |
|
|
|
| 22 |
A |
279 |
267 |
0.09 |
|
|
|
| 23 |
B |
3158 |
3022 |
10.27 |
|
|
|
| 24 |
B |
3099 |
2965 |
58.01 |
|
|
|
| 25 |
B |
3066 |
2933 |
89.40 |
|
|
|
| 26 |
B |
3046 |
2914 |
12.61 |
|
|
|
| 27 |
B |
3019 |
2888 |
67.66 |
|
|
|
| 28 |
B |
1514 |
1449 |
4.52 |
|
|
|
| 29 |
B |
1505 |
1440 |
5.88 |
|
|
|
| 30 |
B |
1436 |
1373 |
0.10 |
|
|
|
| 31 |
B |
1392 |
1331 |
2.23 |
|
|
|
| 32 |
B |
1361 |
1302 |
1.28 |
|
|
|
| 33 |
B |
1306 |
1250 |
1.92 |
|
|
|
| 34 |
B |
1171 |
1120 |
4.15 |
|
|
|
| 35 |
B |
1064 |
1018 |
4.06 |
|
|
|
| 36 |
B |
1021 |
977 |
1.11 |
|
|
|
| 37 |
B |
940 |
899 |
3.87 |
|
|
|
| 38 |
B |
899 |
861 |
5.08 |
|
|
|
| 39 |
B |
733 |
701 |
8.49 |
|
|
|
| 40 |
B |
656 |
627 |
23.83 |
|
|
|
| 41 |
B |
456 |
437 |
1.40 |
|
|
|
| 42 |
B |
169 |
162 |
0.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32334.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30934.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.134 |
|
|
|
| 2 |
C |
-0.134 |
|
|
|
| 3 |
C |
-0.350 |
|
|
|
| 4 |
C |
-0.350 |
|
|
|
| 5 |
C |
-0.320 |
|
|
|
| 6 |
C |
-0.320 |
|
|
|
| 7 |
H |
0.148 |
|
|
|
| 8 |
H |
0.148 |
|
|
|
| 9 |
H |
0.169 |
|
|
|
| 10 |
H |
0.169 |
|
|
|
| 11 |
H |
0.167 |
|
|
|
| 12 |
H |
0.167 |
|
|
|
| 13 |
H |
0.163 |
|
|
|
| 14 |
H |
0.163 |
|
|
|
| 15 |
H |
0.158 |
|
|
|
| 16 |
H |
0.158 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.244 |
0.244 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.682 |
0.091 |
0.000 |
| y |
0.091 |
-36.210 |
0.000 |
| z |
0.000 |
0.000 |
-37.286 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.934 |
0.091 |
0.000 |
| y |
0.091 |
1.774 |
0.000 |
| z |
0.000 |
0.000 |
0.160 |
|
| Polar |
|---|
| 3z2-r2 | 0.320 |
|---|
| x2-y2 | -2.472 |
|---|
| xy | 0.091 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
154.012 |
| (<r2>)1/2 |
12.410 |