return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (cyclohexene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-234.346067
Energy at 298.15K-234.357671
HF Energy-234.090478
Nuclear repulsion energy236.664378
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3202 3039 46.10      
2 A 3113 2955 48.16      
3 A 3088 2931 1.40      
4 A 3065 2910 19.31      
5 A 3043 2889 0.01      
6 A 1739 1651 2.05      
7 A 1551 1472 0.87      
8 A 1525 1448 0.03      
9 A 1419 1347 0.87      
10 A 1410 1338 0.11      
11 A 1297 1231 0.90      
12 A 1274 1209 0.10      
13 A 1183 1123 0.01      
14 A 1122 1065 0.01      
15 A 1098 1042 0.12      
16 A 1001 950 0.01      
17 A 929 881 0.58      
18 A 847 804 0.47      
19 A 840 797 0.54      
20 A 504 479 0.02      
21 A 401 381 0.00      
22 A 282 268 0.07      
23 B 3179 3017 9.73      
24 B 3119 2961 61.24      
25 B 3087 2931 86.81      
26 B 3067 2912 12.43      
27 B 3043 2889 65.22      
28 B 1540 1462 3.53      
29 B 1532 1455 3.96      
30 B 1451 1378 0.24      
31 B 1408 1337 3.01      
32 B 1381 1311 1.11      
33 B 1320 1253 1.79      
34 B 1185 1125 4.17      
35 B 1071 1016 3.00      
36 B 1036 983 1.03      
37 B 956 907 3.51      
38 B 904 858 3.78      
39 B 742 704 8.78      
40 B 662 628 20.35      
41 B 459 436 1.03      
42 B 169 160 0.26      

Unscaled Zero Point Vibrational Energy (zpe) 32620.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 30963.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.15816 0.15144 0.08528

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.120 -0.659 1.304
C2 0.120 0.659 1.304
C3 0.255 1.481 0.045
C4 -0.255 -1.481 0.045
C5 -0.255 0.723 -1.187
C6 0.255 -0.723 -1.187
H7 -0.227 -1.186 2.250
H8 0.227 1.186 2.250
H9 1.308 1.763 -0.096
H10 -1.308 -1.763 -0.096
H11 -0.293 2.424 0.161
H12 0.293 -2.424 0.161
H13 -1.352 0.715 -1.179
H14 1.352 -0.715 -1.179
H15 0.053 1.240 -2.103
H16 -0.053 -1.240 -2.103

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.33932.51081.50952.85162.52001.08852.10233.14082.14233.29232.14303.09312.88663.90353.4561
C21.33931.50952.51082.52002.85162.10231.08852.14233.14082.14303.29232.88663.09313.45613.9035
C32.51081.50953.00581.53432.52503.49402.22471.09923.60371.09673.90722.15992.74312.17053.4801
C41.50952.51083.00582.52501.53432.22473.49403.60371.09923.90721.09672.74312.15993.48012.1705
C52.85162.52001.53432.52501.53233.93183.50182.17132.91182.17163.46711.09692.15571.09532.1748
C62.52002.85162.52501.53431.53233.50183.93182.91182.17133.46712.17162.15571.09692.17481.0953
H71.08852.10233.49402.22473.93183.50182.41534.06912.64674.17152.48304.07883.80404.99084.3562
H82.10231.08852.22473.49403.50183.93182.41532.64674.06912.48304.17153.80404.07884.35624.9908
H93.14082.14231.09923.60372.17132.91184.06912.64674.39041.75084.31623.05652.70482.42353.8595
H102.14233.14083.60371.09922.91182.17132.64674.06914.39044.31621.75082.70483.05653.85952.4235
H113.29232.14301.09673.90722.17163.46714.17152.48301.75084.31624.88362.41613.78892.57824.3137
H122.14303.29233.90721.09673.46712.17162.48304.17154.31621.75084.88363.78892.41614.31372.5782
H133.09312.88662.15992.74311.09692.15574.07883.80403.05652.70482.41613.78893.05811.76102.5222
H142.88663.09312.74312.15992.15571.09693.80404.07882.70483.05653.78892.41613.05812.52221.7610
H153.90353.45612.17053.48011.09532.17484.99084.35622.42353.85952.57824.31371.76102.52222.4822
H163.45613.90353.48012.17052.17481.09534.35624.99083.85952.42354.31372.57822.52221.76102.4822

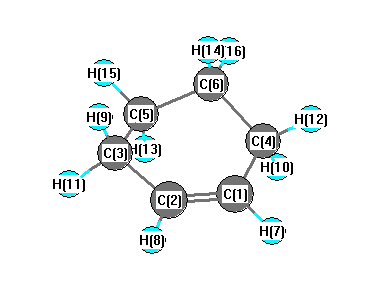
picture of cyclohexene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.502 C1 C2 H8 119.619
C1 C4 C6 111.768 C1 C4 H10 109.407
C1 C4 H12 109.609 C2 C1 C4 123.502
C2 C1 H7 119.619 C2 C3 C5 111.768
C2 C3 H9 109.407 C2 C3 H11 109.609
C3 C2 H8 116.880 C3 C5 C6 110.849
C3 C5 H13 109.218 C3 C5 H15 110.139
C4 C1 H7 116.880 C4 C6 C5 110.849
C4 C6 H14 109.218 C4 C6 H16 110.139
C5 C3 H9 109.981 C5 C3 H11 110.146
C5 C6 H14 109.025 C5 C6 H16 110.615
C6 C4 H10 109.981 C6 C4 H12 110.146
C6 C5 H13 109.025 C6 C5 H15 110.615
H9 C3 H11 105.750 H10 C4 H12 105.750
H13 C5 H15 106.894 H14 C6 H16 106.894
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.138      
2 C -0.138      
3 C -0.311      
4 C -0.311      
5 C -0.288      
6 C -0.288      
7 H 0.143      
8 H 0.143      
9 H 0.153      
10 H 0.153      
11 H 0.151      
12 H 0.151      
13 H 0.147      
14 H 0.147      
15 H 0.144      
16 H 0.144      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 6.752 0.618 0.000
y 0.618 10.347 0.000
z 0.000 0.000 8.880


<r2> (average value of r2) Å2
<r2> 154.412
(<r2>)1/2 12.426