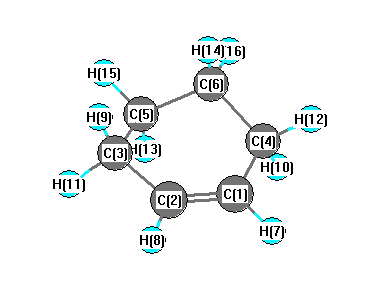
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3101 |
3050 |
48.25 |
|
|
|
| 2 |
A |
3019 |
2969 |
48.02 |
|
|
|
| 3 |
A |
2986 |
2937 |
1.61 |
|
|
|
| 4 |
A |
2966 |
2917 |
17.79 |
|
|
|
| 5 |
A |
2937 |
2889 |
0.01 |
|
|
|
| 6 |
A |
1688 |
1660 |
3.15 |
|
|
|
| 7 |
A |
1478 |
1453 |
1.49 |
|
|
|
| 8 |
A |
1450 |
1426 |
0.00 |
|
|
|
| 9 |
A |
1354 |
1332 |
0.70 |
|
|
|
| 10 |
A |
1346 |
1324 |
0.30 |
|
|
|
| 11 |
A |
1243 |
1222 |
0.69 |
|
|
|
| 12 |
A |
1222 |
1201 |
0.26 |
|
|
|
| 13 |
A |
1131 |
1113 |
0.00 |
|
|
|
| 14 |
A |
1067 |
1049 |
0.01 |
|
|
|
| 15 |
A |
1062 |
1045 |
0.09 |
|
|
|
| 16 |
A |
956 |
940 |
0.02 |
|
|
|
| 17 |
A |
905 |
890 |
0.56 |
|
|
|
| 18 |
A |
822 |
809 |
0.00 |
|
|
|
| 19 |
A |
806 |
792 |
1.39 |
|
|
|
| 20 |
A |
487 |
479 |
0.04 |
|
|
|
| 21 |
A |
387 |
381 |
0.00 |
|
|
|
| 22 |
A |
270 |
266 |
0.10 |
|
|
|
| 23 |
B |
3078 |
3027 |
9.81 |
|
|
|
| 24 |
B |
3023 |
2973 |
52.36 |
|
|
|
| 25 |
B |
2986 |
2937 |
91.07 |
|
|
|
| 26 |
B |
2970 |
2921 |
9.59 |
|
|
|
| 27 |
B |
2937 |
2889 |
65.29 |
|
|
|
| 28 |
B |
1468 |
1443 |
4.34 |
|
|
|
| 29 |
B |
1457 |
1433 |
5.78 |
|
|
|
| 30 |
B |
1391 |
1368 |
0.51 |
|
|
|
| 31 |
B |
1342 |
1320 |
2.30 |
|
|
|
| 32 |
B |
1316 |
1294 |
1.20 |
|
|
|
| 33 |
B |
1266 |
1245 |
1.56 |
|
|
|
| 34 |
B |
1134 |
1115 |
3.60 |
|
|
|
| 35 |
B |
1037 |
1020 |
4.47 |
|
|
|
| 36 |
B |
992 |
975 |
0.97 |
|
|
|
| 37 |
B |
910 |
895 |
3.18 |
|
|
|
| 38 |
B |
878 |
864 |
5.04 |
|
|
|
| 39 |
B |
710 |
698 |
6.80 |
|
|
|
| 40 |
B |
633 |
623 |
23.06 |
|
|
|
| 41 |
B |
442 |
435 |
1.50 |
|
|
|
| 42 |
B |
163 |
160 |
0.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31406.8 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 30888.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.109 |
|
|
|
| 2 |
C |
-0.109 |
|
|
|
| 3 |
C |
-0.330 |
|
|
|
| 4 |
C |
-0.330 |
|
|
|
| 5 |
C |
-0.294 |
|
|
|
| 6 |
C |
-0.294 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.126 |
|
|
|
| 9 |
H |
0.158 |
|
|
|
| 10 |
H |
0.158 |
|
|
|
| 11 |
H |
0.154 |
|
|
|
| 12 |
H |
0.154 |
|
|
|
| 13 |
H |
0.151 |
|
|
|
| 14 |
H |
0.151 |
|
|
|
| 15 |
H |
0.144 |
|
|
|
| 16 |
H |
0.144 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.262 |
0.262 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.735 |
0.493 |
0.003 |
| y |
0.493 |
-36.508 |
-0.001 |
| z |
0.003 |
-0.001 |
-37.549 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.707 |
0.493 |
0.003 |
| y |
0.493 |
1.634 |
-0.001 |
| z |
0.003 |
-0.001 |
0.072 |
|
| Polar |
|---|
| 3z2-r2 | 0.145 |
|---|
| x2-y2 | -2.227 |
|---|
| xy | 0.493 |
|---|
| xz | 0.003 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.974 |
0.659 |
0.002 |
| y |
0.659 |
10.841 |
-0.001 |
| z |
0.002 |
-0.001 |
9.338 |
<r2> (average value of r
2) Å
2
| <r2> |
155.470 |
| (<r2>)1/2 |
12.469 |