Jump to
S1C2
Vibrational Frequencies calculated at CID/6-31G*
Geometric Data calculated at CID/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -250.898998 |
| Energy at 298.15K | -250.913592 |
| Nuclear repulsion energy | 259.837441 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3639 |
3361 |
0.22 |
|
|
|
| 2 |
A' |
3217 |
2972 |
71.94 |
|
|
|
| 3 |
A' |
3205 |
2960 |
50.42 |
|
|
|
| 4 |
A' |
3198 |
2954 |
41.78 |
|
|
|
| 5 |
A' |
3161 |
2920 |
42.46 |
|
|
|
| 6 |
A' |
3150 |
2910 |
22.02 |
|
|
|
| 7 |
A' |
3147 |
2907 |
29.03 |
|
|
|
| 8 |
A' |
1610 |
1487 |
2.05 |
|
|
|
| 9 |
A' |
1594 |
1473 |
9.14 |
|
|
|
| 10 |
A' |
1589 |
1468 |
3.49 |
|
|
|
| 11 |
A' |
1493 |
1379 |
6.98 |
|
|
|
| 12 |
A' |
1479 |
1366 |
0.50 |
|
|
|
| 13 |
A' |
1421 |
1313 |
0.27 |
|
|
|
| 14 |
A' |
1361 |
1257 |
10.46 |
|
|
|
| 15 |
A' |
1272 |
1175 |
3.77 |
|
|
|
| 16 |
A' |
1115 |
1030 |
1.19 |
|
|
|
| 17 |
A' |
1081 |
998 |
11.59 |
|
|
|
| 18 |
A' |
978 |
903 |
15.85 |
|
|
|
| 19 |
A' |
923 |
852 |
44.06 |
|
|
|
| 20 |
A' |
869 |
803 |
1.79 |
|
|
|
| 21 |
A' |
834 |
770 |
97.41 |
|
|
|
| 22 |
A' |
586 |
542 |
2.03 |
|
|
|
| 23 |
A' |
457 |
423 |
1.11 |
|
|
|
| 24 |
A' |
412 |
381 |
6.35 |
|
|
|
| 25 |
A' |
255 |
236 |
4.99 |
|
|
|
| 26 |
A" |
3214 |
2969 |
33.26 |
|
|
|
| 27 |
A" |
3196 |
2953 |
70.36 |
|
|
|
| 28 |
A" |
3156 |
2916 |
16.22 |
|
|
|
| 29 |
A" |
3145 |
2905 |
25.50 |
|
|
|
| 30 |
A" |
1594 |
1473 |
7.38 |
|
|
|
| 31 |
A" |
1590 |
1469 |
3.60 |
|
|
|
| 32 |
A" |
1576 |
1456 |
8.59 |
|
|
|
| 33 |
A" |
1479 |
1367 |
0.28 |
|
|
|
| 34 |
A" |
1466 |
1354 |
2.21 |
|
|
|
| 35 |
A" |
1435 |
1326 |
1.28 |
|
|
|
| 36 |
A" |
1376 |
1271 |
0.68 |
|
|
|
| 37 |
A" |
1297 |
1198 |
14.90 |
|
|
|
| 38 |
A" |
1221 |
1128 |
17.25 |
|
|
|
| 39 |
A" |
1185 |
1095 |
1.27 |
|
|
|
| 40 |
A" |
1128 |
1042 |
2.89 |
|
|
|
| 41 |
A" |
1008 |
931 |
4.97 |
|
|
|
| 42 |
A" |
933 |
862 |
0.00 |
|
|
|
| 43 |
A" |
859 |
794 |
0.10 |
|
|
|
| 44 |
A" |
477 |
440 |
0.90 |
|
|
|
| 45 |
A" |
252 |
233 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36817.2 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 34008.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.642 |
1.317 |
0.000 |
| H2 |
0.579 |
2.405 |
0.000 |
| H3 |
1.705 |
1.066 |
0.000 |
| C4 |
-0.008 |
0.741 |
1.256 |
| C5 |
-0.008 |
0.741 |
-1.256 |
| C6 |
-0.008 |
-0.786 |
-1.208 |
| C7 |
-0.008 |
-0.786 |
1.208 |
| N8 |
-0.608 |
-1.338 |
0.000 |
| H9 |
-1.598 |
-1.133 |
0.000 |
| H10 |
0.507 |
1.090 |
2.152 |
| H11 |
0.507 |
1.090 |
-2.152 |
| H12 |
-1.039 |
1.096 |
1.326 |
| H13 |
-1.039 |
1.096 |
-1.326 |
| H14 |
1.020 |
-1.150 |
-1.258 |
| H15 |
1.020 |
-1.150 |
1.258 |
| H16 |
-0.530 |
-1.201 |
-2.070 |
| H17 |
-0.530 |
-1.201 |
2.070 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0900 | 1.0928 | 1.5269 | 1.5269 | 2.5111 | 2.5111 | 2.9344 | 3.3198 | 2.1677 | 2.1677 | 2.1529 | 2.1529 | 2.7952 | 2.7952 | 3.4635 | 3.4635 |
H2 | 1.0900 | | 1.7501 | 2.1660 | 2.1660 | 3.4624 | 3.4624 | 3.9266 | 4.1542 | 2.5229 | 2.5229 | 2.4680 | 2.4680 | 3.7972 | 3.7972 | 4.3029 | 4.3029 | H3 | 1.0928 | 1.7501 | | 2.1492 | 2.1492 | 2.7977 | 2.7977 | 3.3363 | 3.9687 | 2.4628 | 2.4628 | 3.0487 | 3.0487 | 2.6391 | 2.6391 | 3.7971 | 3.7971 | C4 | 1.5269 | 2.1660 | 2.1492 | | 2.5111 | 2.8984 | 1.5275 | 2.5013 | 2.7596 | 1.0910 | 3.4635 | 1.0930 | 2.8029 | 3.3094 | 2.1523 | 3.8858 | 2.1688 | C5 | 1.5269 | 2.1660 | 2.1492 | 2.5111 | | 1.5275 | 2.8984 | 2.5013 | 2.7596 | 3.4635 | 1.0910 | 2.8029 | 1.0930 | 2.1523 | 3.3094 | 2.1688 | 3.8858 | C6 | 2.5111 | 3.4624 | 2.7977 | 2.8984 | 1.5275 | | 2.4162 | 1.4572 | 2.0268 | 3.8822 | 2.1620 | 3.3211 | 2.1494 | 1.0919 | 2.6968 | 1.0891 | 3.3449 | C7 | 2.5111 | 3.4624 | 2.7977 | 1.5275 | 2.8984 | 2.4162 | | 1.4572 | 2.0268 | 2.1620 | 3.8822 | 2.1494 | 3.3211 | 2.6968 | 1.0919 | 3.3449 | 1.0891 | N8 | 2.9344 | 3.9266 | 3.3363 | 2.5013 | 2.5013 | 1.4572 | 1.4572 | | 1.0113 | 3.4301 | 3.4301 | 2.8053 | 2.8053 | 2.0661 | 2.0661 | 2.0759 | 2.0759 | H9 | 3.3198 | 4.1542 | 3.9687 | 2.7596 | 2.7596 | 2.0268 | 2.0268 | 1.0113 | | 3.7419 | 3.7419 | 2.6535 | 2.6535 | 2.9049 | 2.9049 | 2.3304 | 2.3304 | H10 | 2.1677 | 2.5229 | 2.4628 | 1.0910 | 3.4635 | 3.8822 | 2.1620 | 3.4301 | 3.7419 | | 4.3031 | 1.7529 | 3.8064 | 4.1118 | 2.4653 | 4.9133 | 2.5154 | H11 | 2.1677 | 2.5229 | 2.4628 | 3.4635 | 1.0910 | 2.1620 | 3.8822 | 3.4301 | 3.7419 | 4.3031 | | 3.8064 | 1.7529 | 2.4653 | 4.1118 | 2.5154 | 4.9133 | H12 | 2.1529 | 2.4680 | 3.0487 | 1.0930 | 2.8029 | 3.3211 | 2.1494 | 2.8053 | 2.6535 | 1.7529 | 3.8064 | | 2.6529 | 3.9960 | 3.0482 | 4.1316 | 2.4673 | H13 | 2.1529 | 2.4680 | 3.0487 | 2.8029 | 1.0930 | 2.1494 | 3.3211 | 2.8053 | 2.6535 | 3.8064 | 1.7529 | 2.6529 | | 3.0482 | 3.9960 | 2.4673 | 4.1316 | H14 | 2.7952 | 3.7972 | 2.6391 | 3.3094 | 2.1523 | 1.0919 | 2.6968 | 2.0661 | 2.9049 | 4.1118 | 2.4653 | 3.9960 | 3.0482 | | 2.5166 | 1.7499 | 3.6716 | H15 | 2.7952 | 3.7972 | 2.6391 | 2.1523 | 3.3094 | 2.6968 | 1.0919 | 2.0661 | 2.9049 | 2.4653 | 4.1118 | 3.0482 | 3.9960 | 2.5166 | | 3.6716 | 1.7499 | H16 | 3.4635 | 4.3029 | 3.7971 | 3.8858 | 2.1688 | 1.0891 | 3.3449 | 2.0759 | 2.3304 | 4.9133 | 2.5154 | 4.1316 | 2.4673 | 1.7499 | 3.6716 | | 4.1397 | H17 | 3.4635 | 4.3029 | 3.7971 | 2.1688 | 3.8858 | 3.3449 | 1.0891 | 2.0759 | 2.3304 | 2.5154 | 4.9133 | 2.4673 | 4.1316 | 3.6716 | 1.7499 | 4.1397 | |
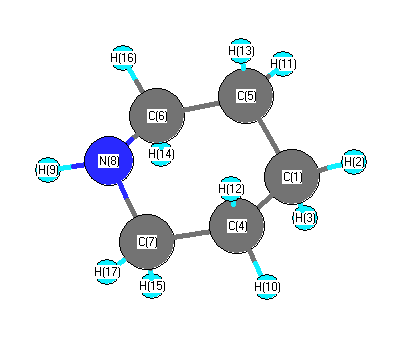 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.595 |
|
C1 |
C4 |
H10 |
110.698 |
| C1 |
C4 |
H12 |
109.409 |
|
C1 |
C5 |
C6 |
110.595 |
| C1 |
C5 |
H11 |
110.698 |
|
C1 |
C5 |
H13 |
109.409 |
| H2 |
C1 |
H3 |
106.607 |
|
H2 |
C1 |
C4 |
110.616 |
| H2 |
C1 |
C5 |
110.616 |
|
H3 |
C1 |
C4 |
109.131 |
| H3 |
C1 |
C5 |
109.131 |
|
C4 |
C1 |
C5 |
110.635 |
| C4 |
C7 |
N8 |
113.843 |
|
C4 |
C7 |
H15 |
109.376 |
| C4 |
C7 |
H17 |
110.848 |
|
C5 |
C6 |
N8 |
113.843 |
| C5 |
C6 |
H14 |
109.376 |
|
C5 |
C6 |
H16 |
110.848 |
| C6 |
C5 |
H11 |
110.198 |
|
C6 |
C5 |
H13 |
109.086 |
| C6 |
N8 |
C7 |
111.999 |
|
C6 |
N8 |
H9 |
109.043 |
| C7 |
C4 |
H10 |
110.198 |
|
C7 |
C4 |
H12 |
109.086 |
| C7 |
N8 |
H9 |
109.043 |
|
N8 |
C6 |
H14 |
107.421 |
| N8 |
C6 |
H16 |
108.352 |
|
N8 |
C7 |
H15 |
107.421 |
| N8 |
C7 |
H17 |
108.352 |
|
H10 |
C4 |
H12 |
106.763 |
| H11 |
C5 |
H13 |
106.763 |
|
H14 |
C6 |
H16 |
106.705 |
| H15 |
C7 |
H17 |
106.705 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
