Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -251.816902 |
| Energy at 298.15K | -251.831244 |
| Nuclear repulsion energy | 259.229362 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3527 |
3374 |
1.18 |
|
|
|
| 2 |
A' |
3107 |
2972 |
76.58 |
|
|
|
| 3 |
A' |
3090 |
2957 |
15.56 |
|
|
|
| 4 |
A' |
3087 |
2953 |
56.99 |
|
|
|
| 5 |
A' |
3054 |
2922 |
14.78 |
|
|
|
| 6 |
A' |
3034 |
2903 |
30.30 |
|
|
|
| 7 |
A' |
2919 |
2793 |
152.23 |
|
|
|
| 8 |
A' |
1531 |
1465 |
2.19 |
|
|
|
| 9 |
A' |
1516 |
1451 |
6.88 |
|
|
|
| 10 |
A' |
1506 |
1441 |
7.84 |
|
|
|
| 11 |
A' |
1443 |
1380 |
4.11 |
|
|
|
| 12 |
A' |
1396 |
1336 |
0.11 |
|
|
|
| 13 |
A' |
1329 |
1271 |
0.42 |
|
|
|
| 14 |
A' |
1301 |
1245 |
3.74 |
|
|
|
| 15 |
A' |
1177 |
1126 |
5.93 |
|
|
|
| 16 |
A' |
1076 |
1029 |
6.87 |
|
|
|
| 17 |
A' |
1064 |
1018 |
6.65 |
|
|
|
| 18 |
A' |
926 |
886 |
1.32 |
|
|
|
| 19 |
A' |
882 |
844 |
16.87 |
|
|
|
| 20 |
A' |
838 |
802 |
0.45 |
|
|
|
| 21 |
A' |
783 |
749 |
85.56 |
|
|
|
| 22 |
A' |
544 |
520 |
20.99 |
|
|
|
| 23 |
A' |
434 |
415 |
7.57 |
|
|
|
| 24 |
A' |
396 |
379 |
1.36 |
|
|
|
| 25 |
A' |
246 |
236 |
1.31 |
|
|
|
| 26 |
A" |
3102 |
2967 |
18.34 |
|
|
|
| 27 |
A" |
3087 |
2953 |
86.50 |
|
|
|
| 28 |
A" |
3054 |
2922 |
26.79 |
|
|
|
| 29 |
A" |
2913 |
2787 |
30.63 |
|
|
|
| 30 |
A" |
1519 |
1454 |
0.55 |
|
|
|
| 31 |
A" |
1504 |
1439 |
0.11 |
|
|
|
| 32 |
A" |
1494 |
1429 |
4.74 |
|
|
|
| 33 |
A" |
1398 |
1337 |
0.37 |
|
|
|
| 34 |
A" |
1377 |
1318 |
24.32 |
|
|
|
| 35 |
A" |
1360 |
1301 |
11.76 |
|
|
|
| 36 |
A" |
1308 |
1251 |
3.20 |
|
|
|
| 37 |
A" |
1202 |
1150 |
8.80 |
|
|
|
| 38 |
A" |
1179 |
1128 |
4.59 |
|
|
|
| 39 |
A" |
1161 |
1111 |
11.65 |
|
|
|
| 40 |
A" |
1079 |
1033 |
2.00 |
|
|
|
| 41 |
A" |
989 |
946 |
0.94 |
|
|
|
| 42 |
A" |
896 |
858 |
0.27 |
|
|
|
| 43 |
A" |
826 |
790 |
0.31 |
|
|
|
| 44 |
A" |
451 |
431 |
0.89 |
|
|
|
| 45 |
A" |
247 |
237 |
0.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35175.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 33652.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.635 |
1.328 |
0.000 |
| H2 |
0.570 |
2.423 |
0.000 |
| H3 |
1.707 |
1.077 |
0.000 |
| C4 |
-0.015 |
0.748 |
1.260 |
| C5 |
-0.015 |
0.748 |
-1.260 |
| C6 |
-0.015 |
-0.779 |
-1.208 |
| C7 |
-0.015 |
-0.779 |
1.208 |
| N8 |
-0.696 |
-1.227 |
0.000 |
| H9 |
-0.765 |
-2.241 |
0.000 |
| H10 |
0.511 |
1.092 |
2.160 |
| H11 |
0.511 |
1.092 |
-2.160 |
| H12 |
-1.054 |
1.095 |
1.333 |
| H13 |
-1.054 |
1.095 |
-1.333 |
| H14 |
1.035 |
-1.136 |
-1.262 |
| H15 |
1.035 |
-1.136 |
1.262 |
| H16 |
-0.541 |
-1.189 |
-2.078 |
| H17 |
-0.541 |
-1.189 |
2.078 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0974 | 1.1006 | 1.5314 | 1.5314 | 2.5138 | 2.5138 | 2.8807 | 3.8334 | 2.1764 | 2.1764 | 2.1640 | 2.1640 | 2.7974 | 2.7974 | 3.4697 | 3.4697 |
H2 | 1.0974 | | 1.7620 | 2.1757 | 2.1757 | 3.4718 | 3.4718 | 3.8633 | 4.8513 | 2.5381 | 2.5381 | 2.4854 | 2.4854 | 3.8054 | 3.8054 | 4.3132 | 4.3132 | H3 | 1.1006 | 1.7620 | | 2.1583 | 2.1583 | 2.8048 | 2.8048 | 3.3290 | 4.1372 | 2.4689 | 2.4689 | 3.0655 | 3.0655 | 2.6351 | 2.6351 | 3.8091 | 3.8091 | C4 | 1.5314 | 2.1757 | 2.1583 | | 2.5197 | 2.9023 | 1.5278 | 2.4399 | 3.3295 | 1.0975 | 3.4770 | 1.0979 | 2.8143 | 3.3189 | 2.1574 | 3.8955 | 2.1683 | C5 | 1.5314 | 2.1757 | 2.1583 | 2.5197 | | 1.5278 | 2.9023 | 2.4399 | 3.3295 | 3.4770 | 1.0975 | 2.8143 | 1.0979 | 2.1574 | 3.3189 | 2.1683 | 3.8955 | C6 | 2.5138 | 3.4718 | 2.8048 | 2.9023 | 1.5278 | | 2.4168 | 1.4580 | 2.0399 | 3.8885 | 2.1633 | 3.3234 | 2.1458 | 1.1104 | 2.7081 | 1.0968 | 3.3539 | C7 | 2.5138 | 3.4718 | 2.8048 | 1.5278 | 2.9023 | 2.4168 | | 1.4580 | 2.0399 | 2.1633 | 3.8885 | 2.1458 | 3.3234 | 2.7081 | 1.1104 | 3.3539 | 1.0968 | N8 | 2.8807 | 3.8633 | 3.3290 | 2.4399 | 2.4399 | 1.4580 | 1.4580 | | 1.0165 | 3.3909 | 3.3909 | 2.7004 | 2.7004 | 2.1446 | 2.1446 | 2.0845 | 2.0845 | H9 | 3.8334 | 4.8513 | 4.1372 | 3.3295 | 3.3295 | 2.0399 | 2.0399 | 1.0165 | | 4.1712 | 4.1712 | 3.6035 | 3.6035 | 2.4601 | 2.4601 | 2.3401 | 2.3401 | H10 | 2.1764 | 2.5381 | 2.4689 | 1.0975 | 3.4770 | 3.8885 | 2.1633 | 3.3909 | 4.1712 | | 4.3200 | 1.7702 | 3.8271 | 4.1171 | 2.4587 | 4.9268 | 2.5132 | H11 | 2.1764 | 2.5381 | 2.4689 | 3.4770 | 1.0975 | 2.1633 | 3.8885 | 3.3909 | 4.1712 | 4.3200 | | 3.8271 | 1.7702 | 2.4587 | 4.1171 | 2.5132 | 4.9268 | H12 | 2.1640 | 2.4854 | 3.0655 | 1.0979 | 2.8143 | 3.3234 | 2.1458 | 2.7004 | 3.6035 | 1.7702 | 3.8271 | | 2.6650 | 4.0093 | 3.0572 | 4.1368 | 2.4566 | H13 | 2.1640 | 2.4854 | 3.0655 | 2.8143 | 1.0979 | 2.1458 | 3.3234 | 2.7004 | 3.6035 | 3.8271 | 1.7702 | 2.6650 | | 3.0572 | 4.0093 | 2.4566 | 4.1368 | H14 | 2.7974 | 3.8054 | 2.6351 | 3.3189 | 2.1574 | 1.1104 | 2.7081 | 2.1446 | 2.4601 | 4.1171 | 2.4587 | 4.0093 | 3.0572 | | 2.5244 | 1.7759 | 3.6943 | H15 | 2.7974 | 3.8054 | 2.6351 | 2.1574 | 3.3189 | 2.7081 | 1.1104 | 2.1446 | 2.4601 | 2.4587 | 4.1171 | 3.0572 | 4.0093 | 2.5244 | | 3.6943 | 1.7759 | H16 | 3.4697 | 4.3132 | 3.8091 | 3.8955 | 2.1683 | 1.0968 | 3.3539 | 2.0845 | 2.3401 | 4.9268 | 2.5132 | 4.1368 | 2.4566 | 1.7759 | 3.6943 | | 4.1568 | H17 | 3.4697 | 4.3132 | 3.8091 | 2.1683 | 3.8955 | 3.3539 | 1.0968 | 2.0845 | 2.3401 | 2.5132 | 4.9268 | 2.4566 | 4.1368 | 3.6943 | 1.7759 | 4.1568 | |
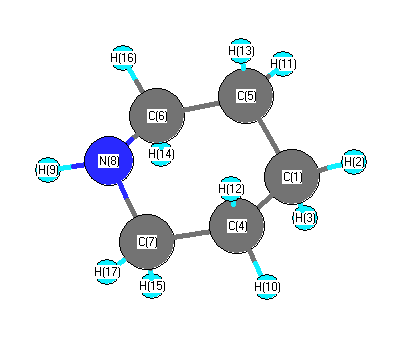 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.518 |
|
C1 |
C4 |
H10 |
110.684 |
| C1 |
C4 |
H12 |
109.679 |
|
C1 |
C5 |
C6 |
110.518 |
| C1 |
C5 |
H11 |
110.684 |
|
C1 |
C5 |
H13 |
109.679 |
| H2 |
C1 |
H3 |
106.583 |
|
H2 |
C1 |
C4 |
110.638 |
| H2 |
C1 |
C5 |
110.638 |
|
H3 |
C1 |
C4 |
109.081 |
| H3 |
C1 |
C5 |
109.081 |
|
C4 |
C1 |
C5 |
110.709 |
| C4 |
C7 |
N8 |
109.580 |
|
C4 |
C7 |
H15 |
108.689 |
| C4 |
C7 |
H17 |
110.333 |
|
C5 |
C6 |
N8 |
109.580 |
| C5 |
C6 |
H14 |
108.689 |
|
C5 |
C6 |
H16 |
110.333 |
| C6 |
C5 |
H11 |
109.895 |
|
C6 |
C5 |
H13 |
108.508 |
| C6 |
N8 |
C7 |
111.949 |
|
C6 |
N8 |
H9 |
109.761 |
| C7 |
C4 |
H10 |
109.895 |
|
C7 |
C4 |
H12 |
108.508 |
| C7 |
N8 |
H9 |
109.761 |
|
N8 |
C6 |
H14 |
112.530 |
| N8 |
C6 |
H16 |
108.535 |
|
N8 |
C7 |
H15 |
112.530 |
| N8 |
C7 |
H17 |
108.535 |
|
H10 |
C4 |
H12 |
107.481 |
| H11 |
C5 |
H13 |
107.481 |
|
H14 |
C6 |
H16 |
107.147 |
| H15 |
C7 |
H17 |
107.147 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.319 |
|
|
|
| 2 |
H |
0.161 |
|
|
|
| 3 |
H |
0.152 |
|
|
|
| 4 |
C |
-0.305 |
|
|
|
| 5 |
C |
-0.305 |
|
|
|
| 6 |
C |
-0.180 |
|
|
|
| 7 |
C |
-0.180 |
|
|
|
| 8 |
N |
-0.564 |
|
|
|
| 9 |
H |
0.314 |
|
|
|
| 10 |
H |
0.155 |
|
|
|
| 11 |
H |
0.155 |
|
|
|
| 12 |
H |
0.168 |
|
|
|
| 13 |
H |
0.168 |
|
|
|
| 14 |
H |
0.128 |
|
|
|
| 15 |
H |
0.128 |
|
|
|
| 16 |
H |
0.161 |
|
|
|
| 17 |
H |
0.161 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.705 |
-0.512 |
0.000 |
0.872 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.743 |
-0.085 |
0.000 |
| y |
-0.085 |
-35.835 |
0.000 |
| z |
0.000 |
0.000 |
-37.518 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.067 |
-0.085 |
0.000 |
| y |
-0.085 |
3.296 |
0.000 |
| z |
0.000 |
0.000 |
0.772 |
|
| Polar |
|---|
| 3z2-r2 | 1.543 |
|---|
| x2-y2 | -4.909 |
|---|
| xy | -0.085 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.834 |
0.305 |
0.000 |
| y |
0.305 |
8.770 |
0.000 |
| z |
0.000 |
0.000 |
9.226 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -251.816441 |
| Energy at 298.15K | -251.830727 |
| Nuclear repulsion energy | 258.671815 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3491 |
3339 |
2.03 |
|
|
|
| 2 |
A' |
3097 |
2963 |
75.19 |
|
|
|
| 3 |
A' |
3089 |
2955 |
45.25 |
|
|
|
| 4 |
A' |
3082 |
2949 |
45.77 |
|
|
|
| 5 |
A' |
3038 |
2906 |
66.25 |
|
|
|
| 6 |
A' |
3034 |
2903 |
19.37 |
|
|
|
| 7 |
A' |
3030 |
2899 |
17.74 |
|
|
|
| 8 |
A' |
1524 |
1458 |
2.42 |
|
|
|
| 9 |
A' |
1507 |
1442 |
12.35 |
|
|
|
| 10 |
A' |
1505 |
1440 |
1.87 |
|
|
|
| 11 |
A' |
1413 |
1352 |
5.63 |
|
|
|
| 12 |
A' |
1397 |
1336 |
0.91 |
|
|
|
| 13 |
A' |
1355 |
1296 |
0.14 |
|
|
|
| 14 |
A' |
1294 |
1238 |
9.49 |
|
|
|
| 15 |
A' |
1201 |
1149 |
3.38 |
|
|
|
| 16 |
A' |
1061 |
1015 |
0.86 |
|
|
|
| 17 |
A' |
1024 |
980 |
8.18 |
|
|
|
| 18 |
A' |
934 |
893 |
12.73 |
|
|
|
| 19 |
A' |
873 |
835 |
44.26 |
|
|
|
| 20 |
A' |
831 |
795 |
0.50 |
|
|
|
| 21 |
A' |
780 |
747 |
100.88 |
|
|
|
| 22 |
A' |
548 |
525 |
2.13 |
|
|
|
| 23 |
A' |
436 |
417 |
1.08 |
|
|
|
| 24 |
A' |
386 |
370 |
5.97 |
|
|
|
| 25 |
A' |
241 |
230 |
5.11 |
|
|
|
| 26 |
A" |
3095 |
2961 |
34.96 |
|
|
|
| 27 |
A" |
3081 |
2948 |
75.86 |
|
|
|
| 28 |
A" |
3034 |
2903 |
4.66 |
|
|
|
| 29 |
A" |
3028 |
2897 |
42.41 |
|
|
|
| 30 |
A" |
1514 |
1448 |
6.12 |
|
|
|
| 31 |
A" |
1503 |
1438 |
5.27 |
|
|
|
| 32 |
A" |
1492 |
1427 |
3.78 |
|
|
|
| 33 |
A" |
1397 |
1336 |
1.56 |
|
|
|
| 34 |
A" |
1385 |
1325 |
2.50 |
|
|
|
| 35 |
A" |
1360 |
1301 |
0.86 |
|
|
|
| 36 |
A" |
1312 |
1256 |
0.88 |
|
|
|
| 37 |
A" |
1232 |
1178 |
15.19 |
|
|
|
| 38 |
A" |
1147 |
1097 |
18.55 |
|
|
|
| 39 |
A" |
1131 |
1082 |
0.28 |
|
|
|
| 40 |
A" |
1078 |
1032 |
2.55 |
|
|
|
| 41 |
A" |
956 |
915 |
5.21 |
|
|
|
| 42 |
A" |
888 |
849 |
0.12 |
|
|
|
| 43 |
A" |
811 |
776 |
0.05 |
|
|
|
| 44 |
A" |
455 |
435 |
0.89 |
|
|
|
| 45 |
A" |
233 |
223 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35150.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 33628.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.639 |
1.325 |
0.000 |
| H2 |
0.575 |
2.420 |
0.000 |
| H3 |
1.711 |
1.076 |
0.000 |
| C4 |
-0.008 |
0.744 |
1.261 |
| C5 |
-0.008 |
0.744 |
-1.261 |
| C6 |
-0.008 |
-0.789 |
-1.210 |
| C7 |
-0.008 |
-0.789 |
1.210 |
| N8 |
-0.606 |
-1.349 |
0.000 |
| H9 |
-1.604 |
-1.138 |
0.000 |
| H10 |
0.513 |
1.095 |
2.162 |
| H11 |
0.513 |
1.095 |
-2.162 |
| H12 |
-1.047 |
1.102 |
1.338 |
| H13 |
-1.047 |
1.102 |
-1.338 |
| H14 |
1.029 |
-1.153 |
-1.262 |
| H15 |
1.029 |
-1.153 |
1.262 |
| H16 |
-0.531 |
-1.208 |
-2.079 |
| H17 |
-0.531 |
-1.208 |
2.079 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0974 | 1.1005 | 1.5319 | 1.5319 | 2.5204 | 2.5204 | 2.9496 | 3.3314 | 2.1778 | 2.1778 | 2.1640 | 2.1640 | 2.8077 | 2.8077 | 3.4796 | 3.4796 |
H2 | 1.0974 | | 1.7608 | 2.1768 | 2.1768 | 3.4790 | 3.4790 | 3.9497 | 4.1723 | 2.5365 | 2.5365 | 2.4814 | 2.4814 | 3.8167 | 3.8167 | 4.3255 | 4.3255 | H3 | 1.1005 | 1.7608 | | 2.1582 | 2.1582 | 2.8106 | 2.8106 | 3.3540 | 3.9864 | 2.4722 | 2.4722 | 3.0656 | 3.0656 | 2.6505 | 2.6505 | 3.8164 | 3.8164 | C4 | 1.5319 | 2.1768 | 2.1582 | | 2.5218 | 2.9082 | 1.5341 | 2.5157 | 2.7711 | 1.0984 | 3.4801 | 1.1007 | 2.8213 | 3.3225 | 2.1623 | 3.9038 | 2.1805 | C5 | 1.5319 | 2.1768 | 2.1582 | 2.5218 | | 1.5341 | 2.9082 | 2.5157 | 2.7711 | 3.4801 | 1.0984 | 2.8213 | 1.1007 | 2.1623 | 3.3225 | 2.1805 | 3.9038 | C6 | 2.5204 | 3.4790 | 2.8106 | 2.9082 | 1.5341 | | 2.4204 | 1.4613 | 2.0325 | 3.8979 | 2.1744 | 3.3384 | 2.1607 | 1.1003 | 2.7052 | 1.0969 | 3.3566 | C7 | 2.5204 | 3.4790 | 2.8106 | 1.5341 | 2.9082 | 2.4204 | | 1.4613 | 2.0325 | 2.1744 | 3.8979 | 2.1607 | 3.3384 | 2.7052 | 1.1003 | 3.3566 | 1.0969 | N8 | 2.9496 | 3.9497 | 3.3540 | 2.5157 | 2.5157 | 1.4613 | 1.4613 | | 1.0193 | 3.4496 | 3.4496 | 2.8265 | 2.8265 | 2.0743 | 2.0743 | 2.0850 | 2.0850 | H9 | 3.3314 | 4.1723 | 3.9864 | 2.7711 | 2.7711 | 2.0325 | 2.0325 | 1.0193 | | 3.7605 | 3.7605 | 2.6679 | 2.6679 | 2.9189 | 2.9189 | 2.3404 | 2.3404 | H10 | 2.1778 | 2.5365 | 2.4722 | 1.0984 | 3.4801 | 3.8979 | 2.1744 | 3.4496 | 3.7605 | | 4.3240 | 1.7637 | 3.8316 | 4.1283 | 2.4763 | 4.9376 | 2.5302 | H11 | 2.1778 | 2.5365 | 2.4722 | 3.4801 | 1.0984 | 2.1744 | 3.8979 | 3.4496 | 3.7605 | 4.3240 | | 3.8316 | 1.7637 | 2.4763 | 4.1283 | 2.5302 | 4.9376 | H12 | 2.1640 | 2.4814 | 3.0656 | 1.1007 | 2.8213 | 3.3384 | 2.1607 | 2.8265 | 2.6679 | 1.7637 | 3.8316 | | 2.6759 | 4.0185 | 3.0653 | 4.1565 | 2.4802 | H13 | 2.1640 | 2.4814 | 3.0656 | 2.8213 | 1.1007 | 2.1607 | 3.3384 | 2.8265 | 2.6679 | 3.8316 | 1.7637 | 2.6759 | | 3.0653 | 4.0185 | 2.4802 | 4.1565 | H14 | 2.8077 | 3.8167 | 2.6505 | 3.3225 | 2.1623 | 1.1003 | 2.7052 | 2.0743 | 2.9189 | 4.1283 | 2.4763 | 4.0185 | 3.0653 | | 2.5234 | 1.7612 | 3.6869 | H15 | 2.8077 | 3.8167 | 2.6505 | 2.1623 | 3.3225 | 2.7052 | 1.1003 | 2.0743 | 2.9189 | 2.4763 | 4.1283 | 3.0653 | 4.0185 | 2.5234 | | 3.6869 | 1.7612 | H16 | 3.4796 | 4.3255 | 3.8164 | 3.9038 | 2.1805 | 1.0969 | 3.3566 | 2.0850 | 2.3404 | 4.9376 | 2.5302 | 4.1565 | 2.4802 | 1.7612 | 3.6869 | | 4.1577 | H17 | 3.4796 | 4.3255 | 3.8164 | 2.1805 | 3.9038 | 3.3566 | 1.0969 | 2.0850 | 2.3404 | 2.5302 | 4.9376 | 2.4802 | 4.1565 | 3.6869 | 1.7612 | 4.1577 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.579 |
|
C1 |
C4 |
H10 |
110.702 |
| C1 |
C4 |
H12 |
109.486 |
|
C1 |
C5 |
C6 |
110.579 |
| C1 |
C5 |
H11 |
110.702 |
|
C1 |
C5 |
H13 |
109.486 |
| H2 |
C1 |
H3 |
106.479 |
|
H2 |
C1 |
C4 |
110.684 |
| H2 |
C1 |
C5 |
110.684 |
|
H3 |
C1 |
C4 |
109.038 |
| H3 |
C1 |
C5 |
109.038 |
|
C4 |
C1 |
C5 |
110.795 |
| C4 |
C7 |
N8 |
114.224 |
|
C4 |
C7 |
H15 |
109.220 |
| C4 |
C7 |
H17 |
110.850 |
|
C5 |
C6 |
N8 |
114.224 |
| C5 |
C6 |
H14 |
109.220 |
|
C5 |
C6 |
H16 |
110.850 |
| C6 |
C5 |
H11 |
110.270 |
|
C6 |
C5 |
H13 |
109.071 |
| C6 |
N8 |
C7 |
111.823 |
|
C6 |
N8 |
H9 |
108.733 |
| C7 |
C4 |
H10 |
110.270 |
|
C7 |
C4 |
H12 |
109.071 |
| C7 |
N8 |
H9 |
108.733 |
|
N8 |
C6 |
H14 |
107.309 |
| N8 |
C6 |
H16 |
108.339 |
|
N8 |
C7 |
H15 |
107.309 |
| N8 |
C7 |
H17 |
108.339 |
|
H10 |
C4 |
H12 |
106.639 |
| H11 |
C5 |
H13 |
106.639 |
|
H14 |
C6 |
H16 |
106.565 |
| H15 |
C7 |
H17 |
106.565 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.311 |
|
|
|
| 2 |
H |
0.161 |
|
|
|
| 3 |
H |
0.155 |
|
|
|
| 4 |
C |
-0.318 |
|
|
|
| 5 |
C |
-0.318 |
|
|
|
| 6 |
C |
-0.198 |
|
|
|
| 7 |
C |
-0.198 |
|
|
|
| 8 |
N |
-0.541 |
|
|
|
| 9 |
H |
0.304 |
|
|
|
| 10 |
H |
0.157 |
|
|
|
| 11 |
H |
0.157 |
|
|
|
| 12 |
H |
0.149 |
|
|
|
| 13 |
H |
0.149 |
|
|
|
| 14 |
H |
0.162 |
|
|
|
| 15 |
H |
0.162 |
|
|
|
| 16 |
H |
0.164 |
|
|
|
| 17 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.290 |
1.175 |
0.000 |
1.210 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.418 |
-0.298 |
0.000 |
| y |
-0.298 |
-43.089 |
0.000 |
| z |
0.000 |
0.000 |
-37.489 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.871 |
-0.298 |
0.000 |
| y |
-0.298 |
-6.136 |
0.000 |
| z |
0.000 |
0.000 |
2.265 |
|
| Polar |
|---|
| 3z2-r2 | 4.529 |
|---|
| x2-y2 | 6.671 |
|---|
| xy | -0.298 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.111 |
0.253 |
0.000 |
| y |
0.253 |
8.319 |
0.000 |
| z |
0.000 |
0.000 |
9.243 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
