Jump to
S1C2
Vibrational Frequencies calculated at QCISD/6-31G*
Geometric Data calculated at QCISD/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -251.067883 |
| Energy at 298.15K | -251.082313 |
| Nuclear repulsion energy | 258.295419 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3473 |
3308 |
2.14 |
|
|
|
| 2 |
A' |
3107 |
2959 |
68.69 |
|
|
|
| 3 |
A' |
3096 |
2949 |
45.19 |
|
|
|
| 4 |
A' |
3090 |
2942 |
40.09 |
|
|
|
| 5 |
A' |
3052 |
2907 |
42.52 |
|
|
|
| 6 |
A' |
3043 |
2898 |
22.19 |
|
|
|
| 7 |
A' |
3040 |
2895 |
24.35 |
|
|
|
| 8 |
A' |
1557 |
1483 |
2.21 |
|
|
|
| 9 |
A' |
1543 |
1469 |
10.75 |
|
|
|
| 10 |
A' |
1538 |
1465 |
1.66 |
|
|
|
| 11 |
A' |
1435 |
1367 |
5.23 |
|
|
|
| 12 |
A' |
1425 |
1357 |
0.47 |
|
|
|
| 13 |
A' |
1371 |
1306 |
0.51 |
|
|
|
| 14 |
A' |
1316 |
1253 |
10.05 |
|
|
|
| 15 |
A' |
1233 |
1174 |
4.48 |
|
|
|
| 16 |
A' |
1081 |
1030 |
0.78 |
|
|
|
| 17 |
A' |
1047 |
997 |
9.73 |
|
|
|
| 18 |
A' |
956 |
910 |
25.03 |
|
|
|
| 19 |
A' |
891 |
848 |
38.05 |
|
|
|
| 20 |
A' |
844 |
804 |
3.78 |
|
|
|
| 21 |
A' |
814 |
776 |
80.97 |
|
|
|
| 22 |
A' |
568 |
541 |
2.01 |
|
|
|
| 23 |
A' |
444 |
423 |
1.01 |
|
|
|
| 24 |
A' |
404 |
385 |
5.82 |
|
|
|
| 25 |
A' |
249 |
237 |
4.73 |
|
|
|
| 26 |
A" |
3105 |
2956 |
30.64 |
|
|
|
| 27 |
A" |
3088 |
2941 |
67.28 |
|
|
|
| 28 |
A" |
3048 |
2903 |
14.93 |
|
|
|
| 29 |
A" |
3038 |
2893 |
25.58 |
|
|
|
| 30 |
A" |
1541 |
1467 |
4.24 |
|
|
|
| 31 |
A" |
1538 |
1464 |
3.64 |
|
|
|
| 32 |
A" |
1521 |
1448 |
7.72 |
|
|
|
| 33 |
A" |
1425 |
1357 |
0.37 |
|
|
|
| 34 |
A" |
1412 |
1345 |
2.16 |
|
|
|
| 35 |
A" |
1384 |
1318 |
1.32 |
|
|
|
| 36 |
A" |
1330 |
1266 |
0.76 |
|
|
|
| 37 |
A" |
1250 |
1190 |
11.98 |
|
|
|
| 38 |
A" |
1173 |
1117 |
15.12 |
|
|
|
| 39 |
A" |
1145 |
1091 |
1.73 |
|
|
|
| 40 |
A" |
1095 |
1043 |
3.44 |
|
|
|
| 41 |
A" |
975 |
928 |
4.84 |
|
|
|
| 42 |
A" |
904 |
861 |
0.00 |
|
|
|
| 43 |
A" |
830 |
790 |
0.08 |
|
|
|
| 44 |
A" |
461 |
439 |
0.95 |
|
|
|
| 45 |
A" |
246 |
235 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35561.2 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 33864.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.646 |
1.323 |
0.000 |
| H2 |
0.586 |
2.420 |
0.000 |
| H3 |
1.718 |
1.066 |
0.000 |
| C4 |
-0.008 |
0.745 |
1.261 |
| C5 |
-0.008 |
0.745 |
-1.261 |
| C6 |
-0.008 |
-0.789 |
-1.214 |
| C7 |
-0.008 |
-0.789 |
1.214 |
| N8 |
-0.616 |
-1.350 |
0.000 |
| H9 |
-1.611 |
-1.112 |
0.000 |
| H10 |
0.510 |
1.098 |
2.165 |
| H11 |
0.510 |
1.098 |
-2.165 |
| H12 |
-1.048 |
1.102 |
1.330 |
| H13 |
-1.048 |
1.102 |
-1.330 |
| H14 |
1.030 |
-1.156 |
-1.257 |
| H15 |
1.030 |
-1.156 |
1.257 |
| H16 |
-0.530 |
-1.208 |
-2.084 |
| H17 |
-0.530 |
-1.208 |
2.084 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0990 | 1.1019 | 1.5335 | 1.5335 | 2.5219 | 2.5219 | 2.9562 | 3.3207 | 2.1811 | 2.1811 | 2.1654 | 2.1654 | 2.8057 | 2.8057 | 3.4831 | 3.4831 |
H2 | 1.0990 | | 1.7650 | 2.1791 | 2.1791 | 3.4817 | 3.4817 | 3.9573 | 4.1599 | 2.5382 | 2.5382 | 2.4852 | 2.4852 | 3.8166 | 3.8166 | 4.3302 | 4.3302 | H3 | 1.1019 | 1.7650 | | 2.1611 | 2.1611 | 2.8092 | 2.8092 | 3.3597 | 3.9786 | 2.4795 | 2.4795 | 3.0693 | 3.0693 | 2.6443 | 2.6443 | 3.8167 | 3.8167 | C4 | 1.5335 | 2.1791 | 2.1611 | | 2.5216 | 2.9111 | 1.5343 | 2.5197 | 2.7587 | 1.1001 | 3.4827 | 1.1021 | 2.8147 | 3.3209 | 2.1654 | 3.9083 | 2.1825 | C5 | 1.5335 | 2.1791 | 2.1611 | 2.5216 | | 1.5343 | 2.9111 | 2.5197 | 2.7587 | 3.4827 | 1.1001 | 2.8147 | 1.1021 | 2.1654 | 3.3209 | 2.1825 | 3.9083 | C6 | 2.5219 | 3.4817 | 2.8092 | 2.9111 | 1.5343 | | 2.4273 | 1.4692 | 2.0370 | 3.9041 | 2.1752 | 3.3358 | 2.1610 | 1.1011 | 2.7046 | 1.0983 | 3.3652 | C7 | 2.5219 | 3.4817 | 2.8092 | 1.5343 | 2.9111 | 2.4273 | | 1.4692 | 2.0370 | 2.1752 | 3.9041 | 2.1610 | 3.3358 | 2.7046 | 1.1011 | 3.3652 | 1.0983 | N8 | 2.9562 | 3.9573 | 3.3597 | 2.5197 | 2.5197 | 1.4692 | 1.4692 | | 1.0230 | 3.4565 | 3.4565 | 2.8227 | 2.8227 | 2.0801 | 2.0801 | 2.0910 | 2.0910 | H9 | 3.3207 | 4.1599 | 3.9786 | 2.7587 | 2.7587 | 2.0370 | 2.0370 | 1.0230 | | 3.7511 | 3.7511 | 2.6436 | 2.6436 | 2.9251 | 2.9251 | 2.3501 | 2.3501 | H10 | 2.1811 | 2.5382 | 2.4795 | 1.1001 | 3.4827 | 3.9041 | 2.1752 | 3.4565 | 3.7511 | | 4.3302 | 1.7676 | 3.8267 | 4.1304 | 2.4846 | 4.9450 | 2.5303 | H11 | 2.1811 | 2.5382 | 2.4795 | 3.4827 | 1.1001 | 2.1752 | 3.9041 | 3.4565 | 3.7511 | 4.3302 | | 3.8267 | 1.7676 | 2.4846 | 4.1304 | 2.5303 | 4.9450 | H12 | 2.1654 | 2.4852 | 3.0693 | 1.1021 | 2.8147 | 3.3358 | 2.1610 | 2.8227 | 2.6436 | 1.7676 | 3.8267 | | 2.6603 | 4.0134 | 3.0690 | 4.1546 | 2.4842 | H13 | 2.1654 | 2.4852 | 3.0693 | 2.8147 | 1.1021 | 2.1610 | 3.3358 | 2.8227 | 2.6436 | 3.8267 | 1.7676 | 2.6603 | | 3.0690 | 4.0134 | 2.4842 | 4.1546 | H14 | 2.8057 | 3.8166 | 2.6443 | 3.3209 | 2.1654 | 1.1011 | 2.7046 | 2.0801 | 2.9251 | 4.1304 | 2.4846 | 4.0134 | 3.0690 | | 2.5142 | 1.7658 | 3.6877 | H15 | 2.8057 | 3.8166 | 2.6443 | 2.1654 | 3.3209 | 2.7046 | 1.1011 | 2.0801 | 2.9251 | 2.4846 | 4.1304 | 3.0690 | 4.0134 | 2.5142 | | 3.6877 | 1.7658 | H16 | 3.4831 | 4.3302 | 3.8167 | 3.9083 | 2.1825 | 1.0983 | 3.3652 | 2.0910 | 2.3501 | 4.9450 | 2.5303 | 4.1546 | 2.4842 | 1.7658 | 3.6877 | | 4.1686 | H17 | 3.4831 | 4.3302 | 3.8167 | 2.1825 | 3.9083 | 3.3652 | 1.0983 | 2.0910 | 2.3501 | 2.5303 | 4.9450 | 2.4842 | 4.1546 | 3.6877 | 1.7658 | 4.1686 | |
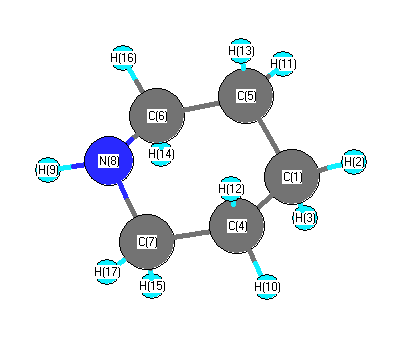 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.580 |
|
C1 |
C4 |
H10 |
110.752 |
| C1 |
C4 |
H12 |
109.404 |
|
C1 |
C5 |
C6 |
110.580 |
| C1 |
C5 |
H11 |
110.752 |
|
C1 |
C5 |
H13 |
109.404 |
| H2 |
C1 |
H3 |
106.635 |
|
H2 |
C1 |
C4 |
110.662 |
| H2 |
C1 |
C5 |
110.662 |
|
H3 |
C1 |
C4 |
109.083 |
| H3 |
C1 |
C5 |
109.083 |
|
C4 |
C1 |
C5 |
110.610 |
| C4 |
C7 |
N8 |
114.035 |
|
C4 |
C7 |
H15 |
109.400 |
| C4 |
C7 |
H17 |
110.914 |
|
C5 |
C6 |
N8 |
114.035 |
| C5 |
C6 |
H14 |
109.400 |
|
C5 |
C6 |
H16 |
110.914 |
| C6 |
C5 |
H11 |
110.232 |
|
C6 |
C5 |
H13 |
109.007 |
| C6 |
N8 |
C7 |
111.391 |
|
C6 |
N8 |
H9 |
108.309 |
| C7 |
C4 |
H10 |
110.232 |
|
C7 |
C4 |
H12 |
109.007 |
| C7 |
N8 |
H9 |
108.309 |
|
N8 |
C6 |
H14 |
107.183 |
| N8 |
C6 |
H16 |
108.187 |
|
N8 |
C7 |
H15 |
107.183 |
| N8 |
C7 |
H17 |
108.187 |
|
H10 |
C4 |
H12 |
106.773 |
| H11 |
C5 |
H13 |
106.773 |
|
H14 |
C6 |
H16 |
106.812 |
| H15 |
C7 |
H17 |
106.812 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
