Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -251.585435 |
| Energy at 298.15K | |
| HF Energy | -251.317175 |
| Nuclear repulsion energy | 259.029303 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3519 |
3340 |
1.40 |
|
|
|
| 2 |
A' |
3127 |
2968 |
80.28 |
|
|
|
| 3 |
A' |
3112 |
2954 |
9.88 |
|
|
|
| 4 |
A' |
3108 |
2950 |
58.43 |
|
|
|
| 5 |
A' |
3076 |
2919 |
12.59 |
|
|
|
| 6 |
A' |
3055 |
2900 |
28.93 |
|
|
|
| 7 |
A' |
2956 |
2805 |
139.21 |
|
|
|
| 8 |
A' |
1558 |
1479 |
1.79 |
|
|
|
| 9 |
A' |
1542 |
1464 |
5.05 |
|
|
|
| 10 |
A' |
1533 |
1455 |
5.98 |
|
|
|
| 11 |
A' |
1462 |
1388 |
5.43 |
|
|
|
| 12 |
A' |
1417 |
1345 |
0.31 |
|
|
|
| 13 |
A' |
1341 |
1273 |
0.29 |
|
|
|
| 14 |
A' |
1317 |
1250 |
3.69 |
|
|
|
| 15 |
A' |
1198 |
1137 |
6.09 |
|
|
|
| 16 |
A' |
1088 |
1033 |
11.08 |
|
|
|
| 17 |
A' |
1071 |
1017 |
2.10 |
|
|
|
| 18 |
A' |
931 |
884 |
4.25 |
|
|
|
| 19 |
A' |
893 |
848 |
14.14 |
|
|
|
| 20 |
A' |
841 |
798 |
1.09 |
|
|
|
| 21 |
A' |
800 |
759 |
81.64 |
|
|
|
| 22 |
A' |
555 |
527 |
19.11 |
|
|
|
| 23 |
A' |
438 |
416 |
7.53 |
|
|
|
| 24 |
A' |
407 |
387 |
1.33 |
|
|
|
| 25 |
A' |
251 |
238 |
1.51 |
|
|
|
| 26 |
A" |
3121 |
2963 |
11.65 |
|
|
|
| 27 |
A" |
3111 |
2953 |
85.59 |
|
|
|
| 28 |
A" |
3076 |
2919 |
30.14 |
|
|
|
| 29 |
A" |
2950 |
2800 |
25.21 |
|
|
|
| 30 |
A" |
1546 |
1468 |
0.53 |
|
|
|
| 31 |
A" |
1526 |
1448 |
0.91 |
|
|
|
| 32 |
A" |
1512 |
1436 |
2.25 |
|
|
|
| 33 |
A" |
1418 |
1346 |
0.52 |
|
|
|
| 34 |
A" |
1396 |
1325 |
18.67 |
|
|
|
| 35 |
A" |
1378 |
1308 |
17.46 |
|
|
|
| 36 |
A" |
1322 |
1255 |
2.48 |
|
|
|
| 37 |
A" |
1207 |
1146 |
5.56 |
|
|
|
| 38 |
A" |
1193 |
1132 |
4.86 |
|
|
|
| 39 |
A" |
1163 |
1104 |
14.14 |
|
|
|
| 40 |
A" |
1086 |
1031 |
2.28 |
|
|
|
| 41 |
A" |
1003 |
952 |
0.82 |
|
|
|
| 42 |
A" |
900 |
854 |
0.08 |
|
|
|
| 43 |
A" |
838 |
796 |
0.45 |
|
|
|
| 44 |
A" |
455 |
432 |
0.91 |
|
|
|
| 45 |
A" |
253 |
240 |
0.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35524.3 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 33719.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.638 |
1.328 |
0.000 |
| H2 |
0.573 |
2.421 |
0.000 |
| H3 |
1.707 |
1.076 |
0.000 |
| C4 |
-0.013 |
0.749 |
1.261 |
| C5 |
-0.013 |
0.749 |
-1.261 |
| C6 |
-0.013 |
-0.779 |
-1.211 |
| C7 |
-0.013 |
-0.779 |
1.211 |
| N8 |
-0.704 |
-1.225 |
0.000 |
| H9 |
-0.767 |
-2.240 |
0.000 |
| H10 |
0.512 |
1.092 |
2.159 |
| H11 |
0.512 |
1.092 |
-2.159 |
| H12 |
-1.051 |
1.094 |
1.333 |
| H13 |
-1.051 |
1.094 |
-1.333 |
| H14 |
1.033 |
-1.138 |
-1.257 |
| H15 |
1.033 |
-1.138 |
1.257 |
| H16 |
-0.537 |
-1.190 |
-2.080 |
| H17 |
-0.537 |
-1.190 |
2.080 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0955 | 1.0985 | 1.5331 | 1.5331 | 2.5159 | 2.5159 | 2.8842 | 3.8346 | 2.1759 | 2.1759 | 2.1642 | 2.1642 | 2.7959 | 2.7959 | 3.4707 | 3.4707 |
H2 | 1.0955 | | 1.7599 | 2.1753 | 2.1753 | 3.4717 | 3.4717 | 3.8636 | 4.8503 | 2.5364 | 2.5364 | 2.4850 | 2.4850 | 3.8029 | 3.8029 | 4.3127 | 4.3127 | H3 | 1.0985 | 1.7599 | | 2.1584 | 2.1584 | 2.8049 | 2.8049 | 3.3330 | 4.1372 | 2.4683 | 2.4683 | 3.0634 | 3.0634 | 2.6337 | 2.6337 | 3.8075 | 3.8075 | C4 | 1.5331 | 2.1753 | 2.1584 | | 2.5226 | 2.9065 | 1.5287 | 2.4422 | 3.3307 | 1.0957 | 3.4779 | 1.0958 | 2.8152 | 3.3163 | 2.1577 | 3.8983 | 2.1687 | C5 | 1.5331 | 2.1753 | 2.1584 | 2.5226 | | 1.5287 | 2.9065 | 2.4422 | 3.3307 | 3.4779 | 1.0957 | 2.8152 | 1.0958 | 2.1577 | 3.3163 | 2.1687 | 3.8983 | C6 | 2.5159 | 3.4717 | 2.8049 | 2.9065 | 1.5287 | | 2.4223 | 1.4640 | 2.0420 | 3.8909 | 2.1626 | 3.3252 | 2.1448 | 1.1071 | 2.7048 | 1.0945 | 3.3577 | C7 | 2.5159 | 3.4717 | 2.8049 | 1.5287 | 2.9065 | 2.4223 | | 1.4640 | 2.0420 | 2.1626 | 3.8909 | 2.1448 | 3.3252 | 2.7048 | 1.1071 | 3.3577 | 1.0945 | N8 | 2.8842 | 3.8636 | 3.3330 | 2.4422 | 2.4422 | 1.4640 | 1.4640 | | 1.0173 | 3.3930 | 3.3930 | 2.6972 | 2.6972 | 2.1460 | 2.1460 | 2.0868 | 2.0868 | H9 | 3.8346 | 4.8503 | 4.1372 | 3.3307 | 3.3307 | 2.0420 | 2.0420 | 1.0173 | | 4.1719 | 4.1719 | 3.6023 | 3.6023 | 2.4562 | 2.4562 | 2.3413 | 2.3413 | H10 | 2.1759 | 2.5364 | 2.4683 | 1.0957 | 3.4779 | 3.8909 | 2.1626 | 3.3930 | 4.1719 | | 4.3190 | 1.7679 | 3.8260 | 4.1133 | 2.4619 | 4.9276 | 2.5131 | H11 | 2.1759 | 2.5364 | 2.4683 | 3.4779 | 1.0957 | 2.1626 | 3.8909 | 3.3930 | 4.1719 | 4.3190 | | 3.8260 | 1.7679 | 2.4619 | 4.1133 | 2.5131 | 4.9276 | H12 | 2.1642 | 2.4850 | 3.0634 | 1.0958 | 2.8152 | 3.3252 | 2.1448 | 2.6972 | 3.6023 | 1.7679 | 3.8260 | | 2.6657 | 4.0041 | 3.0547 | 4.1385 | 2.4574 | H13 | 2.1642 | 2.4850 | 3.0634 | 2.8152 | 1.0958 | 2.1448 | 3.3252 | 2.6972 | 3.6023 | 3.8260 | 1.7679 | 2.6657 | | 3.0547 | 4.0041 | 2.4574 | 4.1385 | H14 | 2.7959 | 3.8029 | 2.6337 | 3.3163 | 2.1577 | 1.1071 | 2.7048 | 2.1460 | 2.4562 | 4.1133 | 2.4619 | 4.0041 | 3.0547 | | 2.5141 | 1.7735 | 3.6882 | H15 | 2.7959 | 3.8029 | 2.6337 | 2.1577 | 3.3163 | 2.7048 | 1.1071 | 2.1460 | 2.4562 | 2.4619 | 4.1133 | 3.0547 | 4.0041 | 2.5141 | | 3.6882 | 1.7735 | H16 | 3.4707 | 4.3127 | 3.8075 | 3.8983 | 2.1687 | 1.0945 | 3.3577 | 2.0868 | 2.3413 | 4.9276 | 2.5131 | 4.1385 | 2.4574 | 1.7735 | 3.6882 | | 4.1596 | H17 | 3.4707 | 4.3127 | 3.8075 | 2.1687 | 3.8983 | 3.3577 | 1.0945 | 2.0868 | 2.3413 | 2.5131 | 4.9276 | 2.4574 | 4.1385 | 3.6882 | 1.7735 | 4.1596 | |
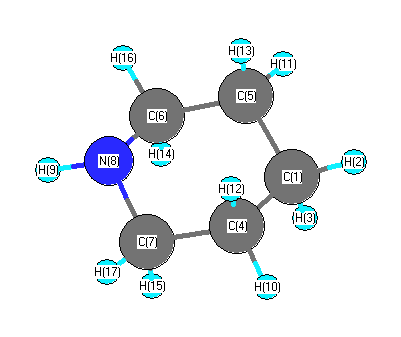 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.511 |
|
C1 |
C4 |
H10 |
110.629 |
| C1 |
C4 |
H12 |
109.697 |
|
C1 |
C5 |
C6 |
110.511 |
| C1 |
C5 |
H11 |
110.629 |
|
C1 |
C5 |
H13 |
109.697 |
| H2 |
C1 |
H3 |
106.666 |
|
H2 |
C1 |
C4 |
110.593 |
| H2 |
C1 |
C5 |
110.593 |
|
H3 |
C1 |
C4 |
109.084 |
| H3 |
C1 |
C5 |
109.084 |
|
C4 |
C1 |
C5 |
110.715 |
| C4 |
C7 |
N8 |
109.365 |
|
C4 |
C7 |
H15 |
108.843 |
| C4 |
C7 |
H17 |
110.438 |
|
C5 |
C6 |
N8 |
109.365 |
| C5 |
C6 |
H14 |
108.843 |
|
C5 |
C6 |
H16 |
110.438 |
| C6 |
C5 |
H11 |
109.886 |
|
C6 |
C5 |
H13 |
108.489 |
| C6 |
N8 |
C7 |
111.647 |
|
C6 |
N8 |
H9 |
109.448 |
| C7 |
C4 |
H10 |
109.886 |
|
C7 |
C4 |
H12 |
108.489 |
| C7 |
N8 |
H9 |
109.448 |
|
N8 |
C6 |
H14 |
112.423 |
| N8 |
C6 |
H16 |
108.435 |
|
N8 |
C7 |
H15 |
112.423 |
| N8 |
C7 |
H17 |
108.435 |
|
H10 |
C4 |
H12 |
107.556 |
| H11 |
C5 |
H13 |
107.556 |
|
H14 |
C6 |
H16 |
107.321 |
| H15 |
C7 |
H17 |
107.321 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.291 |
|
|
|
| 2 |
H |
0.146 |
|
|
|
| 3 |
H |
0.137 |
|
|
|
| 4 |
C |
-0.275 |
|
|
|
| 5 |
C |
-0.275 |
|
|
|
| 6 |
C |
-0.131 |
|
|
|
| 7 |
C |
-0.131 |
|
|
|
| 8 |
N |
-0.609 |
|
|
|
| 9 |
H |
0.311 |
|
|
|
| 10 |
H |
0.141 |
|
|
|
| 11 |
H |
0.141 |
|
|
|
| 12 |
H |
0.153 |
|
|
|
| 13 |
H |
0.153 |
|
|
|
| 14 |
H |
0.117 |
|
|
|
| 15 |
H |
0.117 |
|
|
|
| 16 |
H |
0.148 |
|
|
|
| 17 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.707 |
0.312 |
0.000 |
| y |
0.312 |
8.638 |
0.000 |
| z |
0.000 |
0.000 |
9.080 |
<r2> (average value of r
2) Å
2
| <r2> |
157.992 |
| (<r2>)1/2 |
12.569 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -251.584772 |
| Energy at 298.15K | -251.599140 |
| HF Energy | -251.316305 |
| Nuclear repulsion energy | 258.483609 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3489 |
3312 |
2.11 |
|
|
|
| 2 |
A' |
3121 |
2963 |
70.89 |
|
|
|
| 3 |
A' |
3110 |
2952 |
51.27 |
|
|
|
| 4 |
A' |
3102 |
2945 |
44.41 |
|
|
|
| 5 |
A' |
3064 |
2908 |
49.68 |
|
|
|
| 6 |
A' |
3057 |
2902 |
21.68 |
|
|
|
| 7 |
A' |
3053 |
2898 |
22.21 |
|
|
|
| 8 |
A' |
1551 |
1472 |
1.87 |
|
|
|
| 9 |
A' |
1535 |
1457 |
9.90 |
|
|
|
| 10 |
A' |
1532 |
1454 |
1.15 |
|
|
|
| 11 |
A' |
1431 |
1358 |
6.78 |
|
|
|
| 12 |
A' |
1416 |
1344 |
0.52 |
|
|
|
| 13 |
A' |
1369 |
1299 |
0.37 |
|
|
|
| 14 |
A' |
1310 |
1243 |
9.88 |
|
|
|
| 15 |
A' |
1222 |
1160 |
3.89 |
|
|
|
| 16 |
A' |
1070 |
1016 |
0.93 |
|
|
|
| 17 |
A' |
1035 |
982 |
8.05 |
|
|
|
| 18 |
A' |
944 |
896 |
22.84 |
|
|
|
| 19 |
A' |
881 |
836 |
33.46 |
|
|
|
| 20 |
A' |
835 |
792 |
4.25 |
|
|
|
| 21 |
A' |
796 |
756 |
92.16 |
|
|
|
| 22 |
A' |
561 |
532 |
2.12 |
|
|
|
| 23 |
A' |
441 |
419 |
1.01 |
|
|
|
| 24 |
A' |
396 |
376 |
5.69 |
|
|
|
| 25 |
A' |
245 |
233 |
4.76 |
|
|
|
| 26 |
A" |
3119 |
2961 |
34.34 |
|
|
|
| 27 |
A" |
3101 |
2944 |
71.21 |
|
|
|
| 28 |
A" |
3060 |
2905 |
11.51 |
|
|
|
| 29 |
A" |
3051 |
2896 |
32.79 |
|
|
|
| 30 |
A" |
1534 |
1456 |
2.91 |
|
|
|
| 31 |
A" |
1530 |
1453 |
3.50 |
|
|
|
| 32 |
A" |
1512 |
1435 |
6.93 |
|
|
|
| 33 |
A" |
1416 |
1344 |
1.69 |
|
|
|
| 34 |
A" |
1404 |
1333 |
2.80 |
|
|
|
| 35 |
A" |
1380 |
1310 |
0.86 |
|
|
|
| 36 |
A" |
1328 |
1260 |
0.87 |
|
|
|
| 37 |
A" |
1240 |
1177 |
11.99 |
|
|
|
| 38 |
A" |
1148 |
1090 |
20.36 |
|
|
|
| 39 |
A" |
1142 |
1084 |
0.20 |
|
|
|
| 40 |
A" |
1085 |
1030 |
3.31 |
|
|
|
| 41 |
A" |
970 |
921 |
4.78 |
|
|
|
| 42 |
A" |
892 |
847 |
0.00 |
|
|
|
| 43 |
A" |
824 |
782 |
0.13 |
|
|
|
| 44 |
A" |
459 |
436 |
0.88 |
|
|
|
| 45 |
A" |
240 |
228 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35500.8 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 33697.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.642 |
1.326 |
0.000 |
| H2 |
0.575 |
2.419 |
0.000 |
| H3 |
1.711 |
1.075 |
0.000 |
| C4 |
-0.009 |
0.746 |
1.263 |
| C5 |
-0.009 |
0.746 |
-1.263 |
| C6 |
-0.009 |
-0.790 |
-1.215 |
| C7 |
-0.009 |
-0.790 |
1.215 |
| N8 |
-0.603 |
-1.350 |
0.000 |
| H9 |
-1.604 |
-1.166 |
0.000 |
| H10 |
0.511 |
1.099 |
2.162 |
| H11 |
0.511 |
1.099 |
-2.162 |
| H12 |
-1.046 |
1.102 |
1.335 |
| H13 |
-1.046 |
1.102 |
-1.335 |
| H14 |
1.026 |
-1.151 |
-1.271 |
| H15 |
1.026 |
-1.151 |
1.271 |
| H16 |
-0.537 |
-1.205 |
-2.080 |
| H17 |
-0.537 |
-1.205 |
2.080 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0955 | 1.0982 | 1.5348 | 1.5348 | 2.5248 | 2.5248 | 2.9512 | 3.3547 | 2.1782 | 2.1782 | 2.1639 | 2.1639 | 2.8107 | 2.8107 | 3.4812 | 3.4812 |
H2 | 1.0955 | | 1.7597 | 2.1762 | 2.1762 | 3.4805 | 3.4805 | 3.9490 | 4.1957 | 2.5344 | 2.5344 | 2.4793 | 2.4793 | 3.8169 | 3.8169 | 4.3240 | 4.3240 | H3 | 1.0982 | 1.7597 | | 2.1596 | 2.1596 | 2.8131 | 2.8131 | 3.3523 | 4.0019 | 2.4733 | 2.4733 | 3.0636 | 3.0636 | 2.6541 | 2.6541 | 3.8181 | 3.8181 | C4 | 1.5348 | 2.1762 | 2.1596 | | 2.5265 | 2.9154 | 1.5369 | 2.5185 | 2.7923 | 1.0969 | 3.4828 | 1.0983 | 2.8204 | 3.3313 | 2.1617 | 3.9065 | 2.1801 | C5 | 1.5348 | 2.1762 | 2.1596 | 2.5265 | | 1.5369 | 2.9154 | 2.5185 | 2.7923 | 3.4828 | 1.0969 | 2.8204 | 1.0983 | 2.1617 | 3.3313 | 2.1801 | 3.9065 | C6 | 2.5248 | 3.4805 | 2.8131 | 2.9154 | 1.5369 | | 2.4291 | 1.4635 | 2.0398 | 3.9040 | 2.1762 | 3.3399 | 2.1604 | 1.0980 | 2.7171 | 1.0950 | 3.3618 | C7 | 2.5248 | 3.4805 | 2.8131 | 1.5369 | 2.9154 | 2.4291 | | 1.4635 | 2.0398 | 2.1762 | 3.9040 | 2.1604 | 3.3399 | 2.7171 | 1.0980 | 3.3618 | 1.0950 | N8 | 2.9512 | 3.9490 | 3.3523 | 2.5185 | 2.5185 | 1.4635 | 1.4635 | | 1.0178 | 3.4516 | 3.4516 | 2.8267 | 2.8267 | 2.0761 | 2.0761 | 2.0857 | 2.0857 | H9 | 3.3547 | 4.1957 | 4.0019 | 2.7923 | 2.7923 | 2.0398 | 2.0398 | 1.0178 | | 3.7787 | 3.7787 | 2.6904 | 2.6904 | 2.9214 | 2.9214 | 2.3379 | 2.3379 | H10 | 2.1782 | 2.5344 | 2.4733 | 1.0969 | 3.4828 | 3.9040 | 2.1762 | 3.4516 | 3.7787 | | 4.3247 | 1.7628 | 3.8286 | 4.1376 | 2.4743 | 4.9396 | 2.5323 | H11 | 2.1782 | 2.5344 | 2.4733 | 3.4828 | 1.0969 | 2.1762 | 3.9040 | 3.4516 | 3.7787 | 4.3247 | | 3.8286 | 1.7628 | 2.4743 | 4.1376 | 2.5323 | 4.9396 | H12 | 2.1639 | 2.4793 | 3.0636 | 1.0983 | 2.8204 | 3.3399 | 2.1604 | 2.8267 | 2.6904 | 1.7628 | 3.8286 | | 2.6709 | 4.0206 | 3.0616 | 4.1526 | 2.4769 | H13 | 2.1639 | 2.4793 | 3.0636 | 2.8204 | 1.0983 | 2.1604 | 3.3399 | 2.8267 | 2.6904 | 3.8286 | 1.7628 | 2.6709 | | 3.0616 | 4.0206 | 2.4769 | 4.1526 | H14 | 2.8107 | 3.8169 | 2.6541 | 3.3313 | 2.1617 | 1.0980 | 2.7171 | 2.0761 | 2.9214 | 4.1376 | 2.4743 | 4.0206 | 3.0616 | | 2.5429 | 1.7600 | 3.6978 | H15 | 2.8107 | 3.8169 | 2.6541 | 2.1617 | 3.3313 | 2.7171 | 1.0980 | 2.0761 | 2.9214 | 2.4743 | 4.1376 | 3.0616 | 4.0206 | 2.5429 | | 3.6978 | 1.7600 | H16 | 3.4812 | 4.3240 | 3.8181 | 3.9065 | 2.1801 | 1.0950 | 3.3618 | 2.0857 | 2.3379 | 4.9396 | 2.5323 | 4.1526 | 2.4769 | 1.7600 | 3.6978 | | 4.1592 | H17 | 3.4812 | 4.3240 | 3.8181 | 2.1801 | 3.9065 | 3.3618 | 1.0950 | 2.0857 | 2.3379 | 2.5323 | 4.9396 | 2.4769 | 4.1526 | 3.6978 | 1.7600 | 4.1592 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.562 |
|
C1 |
C4 |
H10 |
110.623 |
| C1 |
C4 |
H12 |
109.412 |
|
C1 |
C5 |
C6 |
110.562 |
| C1 |
C5 |
H11 |
110.623 |
|
C1 |
C5 |
H13 |
109.412 |
| H2 |
C1 |
H3 |
106.668 |
|
H2 |
C1 |
C4 |
110.549 |
| H2 |
C1 |
C5 |
110.549 |
|
H3 |
C1 |
C4 |
109.085 |
| H3 |
C1 |
C5 |
109.085 |
|
C4 |
C1 |
C5 |
110.795 |
| C4 |
C7 |
N8 |
114.134 |
|
C4 |
C7 |
H15 |
109.116 |
| C4 |
C7 |
H17 |
110.740 |
|
C5 |
C6 |
N8 |
114.134 |
| C5 |
C6 |
H14 |
109.116 |
|
C5 |
C6 |
H16 |
110.740 |
| C6 |
C5 |
H11 |
110.313 |
|
C6 |
C5 |
H13 |
108.998 |
| C6 |
N8 |
C7 |
112.181 |
|
C6 |
N8 |
H9 |
109.270 |
| C7 |
C4 |
H10 |
110.313 |
|
C7 |
C4 |
H12 |
108.998 |
| C7 |
N8 |
H9 |
109.270 |
|
N8 |
C6 |
H14 |
107.435 |
| N8 |
C6 |
H16 |
108.358 |
|
N8 |
C7 |
H15 |
107.435 |
| N8 |
C7 |
H17 |
108.358 |
|
H10 |
C4 |
H12 |
106.842 |
| H11 |
C5 |
H13 |
106.842 |
|
H14 |
C6 |
H16 |
106.752 |
| H15 |
C7 |
H17 |
106.752 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.283 |
|
|
|
| 2 |
H |
0.147 |
|
|
|
| 3 |
H |
0.140 |
|
|
|
| 4 |
C |
-0.288 |
|
|
|
| 5 |
C |
-0.288 |
|
|
|
| 6 |
C |
-0.145 |
|
|
|
| 7 |
C |
-0.145 |
|
|
|
| 8 |
N |
-0.589 |
|
|
|
| 9 |
H |
0.301 |
|
|
|
| 10 |
H |
0.143 |
|
|
|
| 11 |
H |
0.143 |
|
|
|
| 12 |
H |
0.134 |
|
|
|
| 13 |
H |
0.134 |
|
|
|
| 14 |
H |
0.148 |
|
|
|
| 15 |
H |
0.148 |
|
|
|
| 16 |
H |
0.151 |
|
|
|
| 17 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.008 |
0.245 |
0.000 |
| y |
0.245 |
8.170 |
0.000 |
| z |
0.000 |
0.000 |
9.095 |
<r2> (average value of r
2) Å
2
| <r2> |
158.627 |
| (<r2>)1/2 |
12.595 |
