Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -251.620060 |
| Energy at 298.15K | -251.634437 |
| Nuclear repulsion energy | 259.723993 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3547 |
3374 |
0.82 |
|
|
|
| 2 |
A' |
3122 |
2969 |
69.96 |
|
|
|
| 3 |
A' |
3107 |
2954 |
14.93 |
|
|
|
| 4 |
A' |
3103 |
2951 |
55.51 |
|
|
|
| 5 |
A' |
3068 |
2918 |
15.30 |
|
|
|
| 6 |
A' |
3047 |
2898 |
29.25 |
|
|
|
| 7 |
A' |
2934 |
2791 |
146.48 |
|
|
|
| 8 |
A' |
1533 |
1458 |
2.36 |
|
|
|
| 9 |
A' |
1518 |
1443 |
7.55 |
|
|
|
| 10 |
A' |
1507 |
1433 |
8.39 |
|
|
|
| 11 |
A' |
1448 |
1377 |
3.73 |
|
|
|
| 12 |
A' |
1401 |
1332 |
0.11 |
|
|
|
| 13 |
A' |
1334 |
1269 |
0.50 |
|
|
|
| 14 |
A' |
1305 |
1241 |
3.76 |
|
|
|
| 15 |
A' |
1182 |
1124 |
6.14 |
|
|
|
| 16 |
A' |
1082 |
1029 |
5.96 |
|
|
|
| 17 |
A' |
1071 |
1018 |
8.01 |
|
|
|
| 18 |
A' |
933 |
887 |
1.04 |
|
|
|
| 19 |
A' |
886 |
842 |
18.75 |
|
|
|
| 20 |
A' |
847 |
805 |
0.43 |
|
|
|
| 21 |
A' |
790 |
751 |
84.62 |
|
|
|
| 22 |
A' |
546 |
519 |
20.55 |
|
|
|
| 23 |
A' |
437 |
415 |
7.42 |
|
|
|
| 24 |
A' |
401 |
381 |
1.36 |
|
|
|
| 25 |
A' |
248 |
236 |
1.32 |
|
|
|
| 26 |
A" |
3118 |
2965 |
19.51 |
|
|
|
| 27 |
A" |
3103 |
2951 |
80.81 |
|
|
|
| 28 |
A" |
3068 |
2918 |
25.33 |
|
|
|
| 29 |
A" |
2928 |
2785 |
29.09 |
|
|
|
| 30 |
A" |
1522 |
1448 |
0.72 |
|
|
|
| 31 |
A" |
1508 |
1434 |
0.48 |
|
|
|
| 32 |
A" |
1496 |
1422 |
4.98 |
|
|
|
| 33 |
A" |
1404 |
1335 |
0.27 |
|
|
|
| 34 |
A" |
1383 |
1315 |
26.74 |
|
|
|
| 35 |
A" |
1364 |
1297 |
9.87 |
|
|
|
| 36 |
A" |
1312 |
1248 |
3.21 |
|
|
|
| 37 |
A" |
1209 |
1150 |
10.72 |
|
|
|
| 38 |
A" |
1182 |
1124 |
5.37 |
|
|
|
| 39 |
A" |
1171 |
1114 |
7.54 |
|
|
|
| 40 |
A" |
1086 |
1032 |
2.15 |
|
|
|
| 41 |
A" |
991 |
943 |
0.86 |
|
|
|
| 42 |
A" |
904 |
860 |
0.29 |
|
|
|
| 43 |
A" |
829 |
788 |
0.28 |
|
|
|
| 44 |
A" |
453 |
431 |
0.87 |
|
|
|
| 45 |
A" |
250 |
237 |
0.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35337.0 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 33605.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.636 |
1.324 |
0.000 |
| H2 |
0.575 |
2.419 |
0.000 |
| H3 |
1.706 |
1.068 |
0.000 |
| C4 |
-0.015 |
0.747 |
1.257 |
| C5 |
-0.015 |
0.747 |
-1.257 |
| C6 |
-0.015 |
-0.777 |
-1.205 |
| C7 |
-0.015 |
-0.777 |
1.205 |
| N8 |
-0.697 |
-1.223 |
0.000 |
| H9 |
-0.766 |
-2.236 |
0.000 |
| H10 |
0.508 |
1.091 |
2.157 |
| H11 |
0.508 |
1.091 |
-2.157 |
| H12 |
-1.054 |
1.093 |
1.326 |
| H13 |
-1.054 |
1.093 |
-1.326 |
| H14 |
1.035 |
-1.134 |
-1.255 |
| H15 |
1.035 |
-1.134 |
1.255 |
| H16 |
-0.539 |
-1.188 |
-2.076 |
| H17 |
-0.539 |
-1.188 |
2.076 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0968 | 1.1002 | 1.5282 | 1.5282 | 2.5077 | 2.5077 | 2.8750 | 3.8264 | 2.1737 | 2.1737 | 2.1607 | 2.1607 | 2.7883 | 2.7883 | 3.4637 | 3.4637 |
H2 | 1.0968 | | 1.7618 | 2.1733 | 2.1733 | 3.4661 | 3.4661 | 3.8582 | 4.8448 | 2.5346 | 2.5346 | 2.4845 | 2.4845 | 3.7959 | 3.7959 | 4.3080 | 4.3080 | H3 | 1.1002 | 1.7618 | | 2.1548 | 2.1548 | 2.7958 | 2.7958 | 3.3205 | 4.1268 | 2.4676 | 2.4676 | 3.0623 | 3.0623 | 2.6218 | 2.6218 | 3.7995 | 3.7995 | C4 | 1.5282 | 2.1733 | 2.1548 | | 2.5133 | 2.8951 | 1.5247 | 2.4345 | 3.3233 | 1.0969 | 3.4709 | 1.0976 | 2.8054 | 3.3088 | 2.1539 | 3.8887 | 2.1654 | C5 | 1.5282 | 2.1733 | 2.1548 | 2.5133 | | 1.5247 | 2.8951 | 2.4345 | 3.3233 | 3.4709 | 1.0969 | 2.8054 | 1.0976 | 2.1539 | 3.3088 | 2.1654 | 3.8887 | C6 | 2.5077 | 3.4661 | 2.7958 | 2.8951 | 1.5247 | | 2.4099 | 1.4549 | 2.0363 | 3.8815 | 2.1606 | 3.3138 | 2.1425 | 1.1099 | 2.6983 | 1.0962 | 3.3475 | C7 | 2.5077 | 3.4661 | 2.7958 | 1.5247 | 2.8951 | 2.4099 | | 1.4549 | 2.0363 | 2.1606 | 3.8815 | 2.1425 | 3.3138 | 2.6983 | 1.1099 | 3.3475 | 1.0962 | N8 | 2.8750 | 3.8582 | 3.3205 | 2.4345 | 2.4345 | 1.4549 | 1.4549 | | 1.0154 | 3.3855 | 3.3855 | 2.6925 | 2.6925 | 2.1409 | 2.1409 | 2.0820 | 2.0820 | H9 | 3.8264 | 4.8448 | 4.1268 | 3.3233 | 3.3233 | 2.0363 | 2.0363 | 1.0154 | | 4.1650 | 4.1650 | 3.5950 | 3.5950 | 2.4566 | 2.4566 | 2.3366 | 2.3366 | H10 | 2.1737 | 2.5346 | 2.4676 | 1.0969 | 3.4709 | 3.8815 | 2.1606 | 3.3855 | 4.1650 | | 4.3148 | 1.7700 | 3.8179 | 4.1073 | 2.4574 | 4.9200 | 2.5088 | H11 | 2.1737 | 2.5346 | 2.4676 | 3.4709 | 1.0969 | 2.1606 | 3.8815 | 3.3855 | 4.1650 | 4.3148 | | 3.8179 | 1.7700 | 2.4574 | 4.1073 | 2.5088 | 4.9200 | H12 | 2.1607 | 2.4845 | 3.0623 | 1.0976 | 2.8054 | 3.3138 | 2.1425 | 2.6925 | 3.5950 | 1.7700 | 3.8179 | | 2.6522 | 3.9980 | 3.0539 | 4.1277 | 2.4552 | H13 | 2.1607 | 2.4845 | 3.0623 | 2.8054 | 1.0976 | 2.1425 | 3.3138 | 2.6925 | 3.5950 | 3.8179 | 1.7700 | 2.6522 | | 3.0539 | 3.9980 | 2.4552 | 4.1277 | H14 | 2.7883 | 3.7959 | 2.6218 | 3.3088 | 2.1539 | 1.1099 | 2.6983 | 2.1409 | 2.4566 | 4.1073 | 2.4574 | 3.9980 | 3.0539 | | 2.5102 | 1.7757 | 3.6842 | H15 | 2.7883 | 3.7959 | 2.6218 | 2.1539 | 3.3088 | 2.6983 | 1.1099 | 2.1409 | 2.4566 | 2.4574 | 4.1073 | 3.0539 | 3.9980 | 2.5102 | | 3.6842 | 1.7757 | H16 | 3.4637 | 4.3080 | 3.7995 | 3.8887 | 2.1654 | 1.0962 | 3.3475 | 2.0820 | 2.3366 | 4.9200 | 2.5088 | 4.1277 | 2.4552 | 1.7757 | 3.6842 | | 4.1513 | H17 | 3.4637 | 4.3080 | 3.7995 | 2.1654 | 3.8887 | 3.3475 | 1.0962 | 2.0820 | 2.3366 | 2.5088 | 4.9200 | 2.4552 | 4.1277 | 3.6842 | 1.7757 | 4.1513 | |
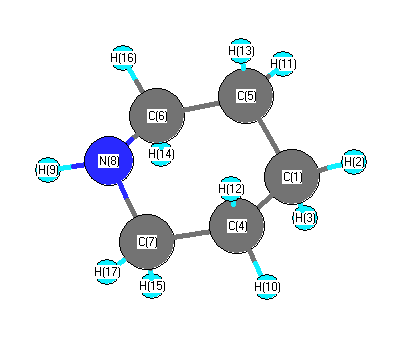 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.454 |
|
C1 |
C4 |
H10 |
110.728 |
| C1 |
C4 |
H12 |
109.656 |
|
C1 |
C5 |
C6 |
110.454 |
| C1 |
C5 |
H11 |
110.728 |
|
C1 |
C5 |
H13 |
109.656 |
| H2 |
C1 |
H3 |
106.624 |
|
H2 |
C1 |
C4 |
110.694 |
| H2 |
C1 |
C5 |
110.694 |
|
H3 |
C1 |
C4 |
109.046 |
| H3 |
C1 |
C5 |
109.046 |
|
C4 |
C1 |
C5 |
110.628 |
| C4 |
C7 |
N8 |
109.558 |
|
C4 |
C7 |
H15 |
108.654 |
| C4 |
C7 |
H17 |
110.347 |
|
C5 |
C6 |
N8 |
109.558 |
| C5 |
C6 |
H14 |
108.654 |
|
C5 |
C6 |
H16 |
110.347 |
| C6 |
C5 |
H11 |
109.927 |
|
C6 |
C5 |
H13 |
108.475 |
| C6 |
N8 |
C7 |
111.824 |
|
C6 |
N8 |
H9 |
109.755 |
| C7 |
C4 |
H10 |
109.927 |
|
C7 |
C4 |
H12 |
108.475 |
| C7 |
N8 |
H9 |
109.755 |
|
N8 |
C6 |
H14 |
112.477 |
| N8 |
C6 |
H16 |
108.579 |
|
N8 |
C7 |
H15 |
112.477 |
| N8 |
C7 |
H17 |
108.579 |
|
H10 |
C4 |
H12 |
107.525 |
| H11 |
C5 |
H13 |
107.525 |
|
H14 |
C6 |
H16 |
107.202 |
| H15 |
C7 |
H17 |
107.202 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.324 |
|
|
|
| 2 |
H |
0.164 |
|
|
|
| 3 |
H |
0.154 |
|
|
|
| 4 |
C |
-0.311 |
|
|
|
| 5 |
C |
-0.311 |
|
|
|
| 6 |
C |
-0.184 |
|
|
|
| 7 |
C |
-0.184 |
|
|
|
| 8 |
N |
-0.567 |
|
|
|
| 9 |
H |
0.317 |
|
|
|
| 10 |
H |
0.158 |
|
|
|
| 11 |
H |
0.158 |
|
|
|
| 12 |
H |
0.170 |
|
|
|
| 13 |
H |
0.170 |
|
|
|
| 14 |
H |
0.131 |
|
|
|
| 15 |
H |
0.131 |
|
|
|
| 16 |
H |
0.164 |
|
|
|
| 17 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.703 |
-0.519 |
0.000 |
0.874 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.852 |
-0.071 |
0.000 |
| y |
-0.071 |
-35.929 |
0.000 |
| z |
0.000 |
0.000 |
-37.615 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.079 |
-0.071 |
0.000 |
| y |
-0.071 |
3.304 |
0.000 |
| z |
0.000 |
0.000 |
0.776 |
|
| Polar |
|---|
| 3z2-r2 | 1.551 |
|---|
| x2-y2 | -4.922 |
|---|
| xy | -0.071 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.802 |
0.304 |
0.000 |
| y |
0.304 |
8.730 |
0.000 |
| z |
0.000 |
0.000 |
9.177 |
<r2> (average value of r
2) Å
2
| <r2> |
156.990 |
| (<r2>)1/2 |
12.530 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -251.619518 |
| Energy at 298.15K | -251.633839 |
| Nuclear repulsion energy | 259.192707 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3509 |
3337 |
1.58 |
|
|
|
| 2 |
A' |
3112 |
2960 |
67.37 |
|
|
|
| 3 |
A' |
3105 |
2953 |
43.26 |
|
|
|
| 4 |
A' |
3099 |
2947 |
43.69 |
|
|
|
| 5 |
A' |
3050 |
2901 |
72.78 |
|
|
|
| 6 |
A' |
3046 |
2897 |
16.63 |
|
|
|
| 7 |
A' |
3042 |
2893 |
14.11 |
|
|
|
| 8 |
A' |
1526 |
1451 |
2.58 |
|
|
|
| 9 |
A' |
1509 |
1435 |
13.29 |
|
|
|
| 10 |
A' |
1506 |
1433 |
2.11 |
|
|
|
| 11 |
A' |
1418 |
1348 |
5.40 |
|
|
|
| 12 |
A' |
1402 |
1333 |
1.09 |
|
|
|
| 13 |
A' |
1360 |
1293 |
0.10 |
|
|
|
| 14 |
A' |
1298 |
1235 |
9.79 |
|
|
|
| 15 |
A' |
1206 |
1146 |
3.49 |
|
|
|
| 16 |
A' |
1069 |
1016 |
0.85 |
|
|
|
| 17 |
A' |
1031 |
981 |
8.96 |
|
|
|
| 18 |
A' |
940 |
894 |
10.04 |
|
|
|
| 19 |
A' |
878 |
835 |
48.93 |
|
|
|
| 20 |
A' |
838 |
797 |
0.11 |
|
|
|
| 21 |
A' |
787 |
748 |
99.14 |
|
|
|
| 22 |
A' |
550 |
523 |
2.26 |
|
|
|
| 23 |
A' |
438 |
416 |
1.07 |
|
|
|
| 24 |
A' |
391 |
372 |
6.10 |
|
|
|
| 25 |
A' |
243 |
231 |
5.16 |
|
|
|
| 26 |
A" |
3110 |
2958 |
34.89 |
|
|
|
| 27 |
A" |
3098 |
2946 |
72.89 |
|
|
|
| 28 |
A" |
3047 |
2897 |
2.51 |
|
|
|
| 29 |
A" |
3040 |
2891 |
40.96 |
|
|
|
| 30 |
A" |
1519 |
1445 |
7.74 |
|
|
|
| 31 |
A" |
1504 |
1430 |
5.52 |
|
|
|
| 32 |
A" |
1494 |
1421 |
3.25 |
|
|
|
| 33 |
A" |
1402 |
1334 |
1.14 |
|
|
|
| 34 |
A" |
1390 |
1322 |
2.67 |
|
|
|
| 35 |
A" |
1363 |
1296 |
1.03 |
|
|
|
| 36 |
A" |
1316 |
1252 |
0.81 |
|
|
|
| 37 |
A" |
1240 |
1179 |
16.57 |
|
|
|
| 38 |
A" |
1161 |
1104 |
16.16 |
|
|
|
| 39 |
A" |
1133 |
1077 |
0.86 |
|
|
|
| 40 |
A" |
1084 |
1031 |
2.55 |
|
|
|
| 41 |
A" |
959 |
912 |
5.01 |
|
|
|
| 42 |
A" |
896 |
852 |
0.15 |
|
|
|
| 43 |
A" |
815 |
775 |
0.03 |
|
|
|
| 44 |
A" |
457 |
434 |
0.90 |
|
|
|
| 45 |
A" |
235 |
224 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35306.3 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 33576.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.641 |
1.321 |
0.000 |
| H2 |
0.575 |
2.416 |
0.000 |
| H3 |
1.711 |
1.069 |
0.000 |
| C4 |
-0.009 |
0.744 |
1.259 |
| C5 |
-0.009 |
0.744 |
-1.259 |
| C6 |
-0.009 |
-0.788 |
-1.208 |
| C7 |
-0.009 |
-0.788 |
1.208 |
| N8 |
-0.602 |
-1.345 |
0.000 |
| H9 |
-1.601 |
-1.159 |
0.000 |
| H10 |
0.511 |
1.096 |
2.159 |
| H11 |
0.511 |
1.096 |
-2.159 |
| H12 |
-1.047 |
1.101 |
1.331 |
| H13 |
-1.047 |
1.101 |
-1.331 |
| H14 |
1.027 |
-1.150 |
-1.264 |
| H15 |
1.027 |
-1.150 |
1.264 |
| H16 |
-0.536 |
-1.205 |
-2.075 |
| H17 |
-0.536 |
-1.205 |
2.075 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0968 | 1.0998 | 1.5298 | 1.5298 | 2.5160 | 2.5160 | 2.9411 | 3.3436 | 2.1745 | 2.1745 | 2.1609 | 2.1609 | 2.8029 | 2.8029 | 3.4741 | 3.4741 |
H2 | 1.0968 | | 1.7623 | 2.1731 | 2.1731 | 3.4736 | 3.4736 | 3.9406 | 4.1858 | 2.5312 | 2.5312 | 2.4768 | 2.4768 | 3.8107 | 3.8107 | 4.3185 | 4.3185 | H3 | 1.0998 | 1.7623 | | 2.1564 | 2.1564 | 2.8048 | 2.8048 | 3.3426 | 3.9921 | 2.4701 | 2.4701 | 3.0630 | 3.0630 | 2.6439 | 2.6439 | 3.8109 | 3.8109 | C4 | 1.5298 | 2.1731 | 2.1564 | | 2.5173 | 2.9037 | 1.5324 | 2.5093 | 2.7821 | 1.0981 | 3.4749 | 1.0999 | 2.8125 | 3.3208 | 2.1593 | 3.8968 | 2.1772 | C5 | 1.5298 | 2.1731 | 2.1564 | 2.5173 | | 1.5324 | 2.9037 | 2.5093 | 2.7821 | 3.4749 | 1.0981 | 2.8125 | 1.0999 | 2.1593 | 3.3208 | 2.1772 | 3.8968 | C6 | 2.5160 | 3.4736 | 2.8048 | 2.9037 | 1.5324 | | 2.4165 | 1.4563 | 2.0327 | 3.8933 | 2.1735 | 3.3301 | 2.1582 | 1.0997 | 2.7055 | 1.0963 | 3.3509 | C7 | 2.5160 | 3.4736 | 2.8048 | 1.5324 | 2.9037 | 2.4165 | | 1.4563 | 2.0327 | 2.1735 | 3.8933 | 2.1582 | 3.3301 | 2.7055 | 1.0997 | 3.3509 | 1.0963 | N8 | 2.9411 | 3.9406 | 3.3426 | 2.5093 | 2.5093 | 1.4563 | 1.4563 | | 1.0167 | 3.4434 | 3.4434 | 2.8194 | 2.8194 | 2.0710 | 2.0710 | 2.0803 | 2.0803 | H9 | 3.3436 | 4.1858 | 3.9921 | 2.7821 | 2.7821 | 2.0327 | 2.0327 | 1.0167 | | 3.7698 | 3.7698 | 2.6805 | 2.6805 | 2.9168 | 2.9168 | 2.3324 | 2.3324 | H10 | 2.1745 | 2.5312 | 2.4701 | 1.0981 | 3.4749 | 3.8933 | 2.1735 | 3.4434 | 3.7698 | | 4.3178 | 1.7650 | 3.8220 | 4.1269 | 2.4725 | 4.9309 | 2.5296 | H11 | 2.1745 | 2.5312 | 2.4701 | 3.4749 | 1.0981 | 2.1735 | 3.8933 | 3.4434 | 3.7698 | 4.3178 | | 3.8220 | 1.7650 | 2.4725 | 4.1269 | 2.5296 | 4.9309 | H12 | 2.1609 | 2.4768 | 3.0630 | 1.0999 | 2.8125 | 3.3301 | 2.1582 | 2.8194 | 2.6805 | 1.7650 | 3.8220 | | 2.6617 | 4.0131 | 3.0618 | 4.1439 | 2.4756 | H13 | 2.1609 | 2.4768 | 3.0630 | 2.8125 | 1.0999 | 2.1582 | 3.3301 | 2.8194 | 2.6805 | 3.8220 | 1.7650 | 2.6617 | | 3.0618 | 4.0131 | 2.4756 | 4.1439 | H14 | 2.8029 | 3.8107 | 2.6439 | 3.3208 | 2.1593 | 1.0997 | 2.7055 | 2.0710 | 2.9168 | 4.1269 | 2.4725 | 4.0131 | 3.0618 | | 2.5286 | 1.7618 | 3.6872 | H15 | 2.8029 | 3.8107 | 2.6439 | 2.1593 | 3.3208 | 2.7055 | 1.0997 | 2.0710 | 2.9168 | 2.4725 | 4.1269 | 3.0618 | 4.0131 | 2.5286 | | 3.6872 | 1.7618 | H16 | 3.4741 | 4.3185 | 3.8109 | 3.8968 | 2.1772 | 1.0963 | 3.3509 | 2.0803 | 2.3324 | 4.9309 | 2.5296 | 4.1439 | 2.4756 | 1.7618 | 3.6872 | | 4.1491 | H17 | 3.4741 | 4.3185 | 3.8109 | 2.1772 | 3.8968 | 3.3509 | 1.0963 | 2.0803 | 2.3324 | 2.5296 | 4.9309 | 2.4756 | 4.1439 | 3.6872 | 1.7618 | 4.1491 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.497 |
|
C1 |
C4 |
H10 |
110.603 |
| C1 |
C4 |
H12 |
109.430 |
|
C1 |
C5 |
C6 |
110.497 |
| C1 |
C5 |
H11 |
110.603 |
|
C1 |
C5 |
H13 |
109.430 |
| H2 |
C1 |
H3 |
106.694 |
|
H2 |
C1 |
C4 |
110.573 |
| H2 |
C1 |
C5 |
110.573 |
|
H3 |
C1 |
C4 |
109.085 |
| H3 |
C1 |
C5 |
109.085 |
|
C4 |
C1 |
C5 |
110.725 |
| C4 |
C7 |
N8 |
114.178 |
|
C4 |
C7 |
H15 |
109.138 |
| C4 |
C7 |
H17 |
110.744 |
|
C5 |
C6 |
N8 |
114.178 |
| C5 |
C6 |
H14 |
109.138 |
|
C5 |
C6 |
H16 |
110.744 |
| C6 |
C5 |
H11 |
110.344 |
|
C6 |
C5 |
H13 |
109.041 |
| C6 |
N8 |
C7 |
112.134 |
|
C6 |
N8 |
H9 |
109.273 |
| C7 |
C4 |
H10 |
110.344 |
|
C7 |
C4 |
H12 |
109.041 |
| C7 |
N8 |
H9 |
109.273 |
|
N8 |
C6 |
H14 |
107.426 |
| N8 |
C6 |
H16 |
108.346 |
|
N8 |
C7 |
H15 |
107.426 |
| N8 |
C7 |
H17 |
108.346 |
|
H10 |
C4 |
H12 |
106.839 |
| H11 |
C5 |
H13 |
106.839 |
|
H14 |
C6 |
H16 |
106.696 |
| H15 |
C7 |
H17 |
106.696 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.317 |
|
|
|
| 2 |
H |
0.164 |
|
|
|
| 3 |
H |
0.157 |
|
|
|
| 4 |
C |
-0.323 |
|
|
|
| 5 |
C |
-0.323 |
|
|
|
| 6 |
C |
-0.202 |
|
|
|
| 7 |
C |
-0.202 |
|
|
|
| 8 |
N |
-0.544 |
|
|
|
| 9 |
H |
0.306 |
|
|
|
| 10 |
H |
0.160 |
|
|
|
| 11 |
H |
0.160 |
|
|
|
| 12 |
H |
0.151 |
|
|
|
| 13 |
H |
0.151 |
|
|
|
| 14 |
H |
0.164 |
|
|
|
| 15 |
H |
0.164 |
|
|
|
| 16 |
H |
0.167 |
|
|
|
| 17 |
H |
0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.287 |
1.177 |
0.000 |
1.212 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.538 |
-0.307 |
0.000 |
| y |
-0.307 |
-43.184 |
0.000 |
| z |
0.000 |
0.000 |
-37.586 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.847 |
-0.307 |
0.000 |
| y |
-0.307 |
-6.122 |
0.000 |
| z |
0.000 |
0.000 |
2.275 |
|
| Polar |
|---|
| 3z2-r2 | 4.550 |
|---|
| x2-y2 | 6.646 |
|---|
| xy | -0.307 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.082 |
0.247 |
0.000 |
| y |
0.247 |
8.280 |
0.000 |
| z |
0.000 |
0.000 |
9.196 |
<r2> (average value of r
2) Å
2
| <r2> |
157.580 |
| (<r2>)1/2 |
12.553 |
