Jump to
S1C2
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -251.053937 |
| Energy at 298.15K | -251.068438 |
| Nuclear repulsion energy | 259.065746 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3555 |
3336 |
0.60 |
|
|
|
| 2 |
A' |
3152 |
2958 |
67.96 |
|
|
|
| 3 |
A' |
3137 |
2945 |
6.68 |
|
|
|
| 4 |
A' |
3134 |
2942 |
52.18 |
|
|
|
| 5 |
A' |
3100 |
2909 |
11.96 |
|
|
|
| 6 |
A' |
3080 |
2891 |
25.66 |
|
|
|
| 7 |
A' |
2994 |
2810 |
115.46 |
|
|
|
| 8 |
A' |
1576 |
1480 |
2.85 |
|
|
|
| 9 |
A' |
1558 |
1463 |
5.72 |
|
|
|
| 10 |
A' |
1549 |
1454 |
6.77 |
|
|
|
| 11 |
A' |
1479 |
1388 |
3.63 |
|
|
|
| 12 |
A' |
1432 |
1344 |
0.03 |
|
|
|
| 13 |
A' |
1352 |
1269 |
0.29 |
|
|
|
| 14 |
A' |
1328 |
1246 |
3.53 |
|
|
|
| 15 |
A' |
1215 |
1141 |
6.70 |
|
|
|
| 16 |
A' |
1102 |
1035 |
12.75 |
|
|
|
| 17 |
A' |
1087 |
1021 |
3.48 |
|
|
|
| 18 |
A' |
945 |
887 |
4.38 |
|
|
|
| 19 |
A' |
908 |
852 |
18.46 |
|
|
|
| 20 |
A' |
857 |
804 |
1.79 |
|
|
|
| 21 |
A' |
821 |
771 |
73.33 |
|
|
|
| 22 |
A' |
562 |
528 |
19.83 |
|
|
|
| 23 |
A' |
441 |
414 |
7.90 |
|
|
|
| 24 |
A' |
416 |
391 |
1.45 |
|
|
|
| 25 |
A' |
255 |
239 |
1.88 |
|
|
|
| 26 |
A" |
3147 |
2954 |
10.90 |
|
|
|
| 27 |
A" |
3135 |
2942 |
73.07 |
|
|
|
| 28 |
A" |
3099 |
2909 |
26.66 |
|
|
|
| 29 |
A" |
2991 |
2807 |
19.56 |
|
|
|
| 30 |
A" |
1565 |
1469 |
0.41 |
|
|
|
| 31 |
A" |
1541 |
1447 |
0.97 |
|
|
|
| 32 |
A" |
1528 |
1435 |
5.32 |
|
|
|
| 33 |
A" |
1436 |
1348 |
0.31 |
|
|
|
| 34 |
A" |
1412 |
1326 |
23.36 |
|
|
|
| 35 |
A" |
1393 |
1307 |
10.05 |
|
|
|
| 36 |
A" |
1333 |
1251 |
2.23 |
|
|
|
| 37 |
A" |
1222 |
1147 |
7.06 |
|
|
|
| 38 |
A" |
1206 |
1131 |
7.44 |
|
|
|
| 39 |
A" |
1189 |
1116 |
4.81 |
|
|
|
| 40 |
A" |
1102 |
1035 |
2.33 |
|
|
|
| 41 |
A" |
1012 |
950 |
0.72 |
|
|
|
| 42 |
A" |
916 |
860 |
0.19 |
|
|
|
| 43 |
A" |
848 |
795 |
0.27 |
|
|
|
| 44 |
A" |
458 |
430 |
0.93 |
|
|
|
| 45 |
A" |
259 |
243 |
0.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35914.1 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 33709.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.640 |
1.327 |
0.000 |
| H2 |
0.574 |
2.421 |
0.000 |
| H3 |
1.709 |
1.074 |
0.000 |
| C4 |
-0.012 |
0.748 |
1.260 |
| C5 |
-0.012 |
0.748 |
-1.260 |
| C6 |
-0.012 |
-0.779 |
-1.210 |
| C7 |
-0.012 |
-0.779 |
1.210 |
| N8 |
-0.710 |
-1.223 |
0.000 |
| H9 |
-0.767 |
-2.239 |
0.000 |
| H10 |
0.511 |
1.093 |
2.160 |
| H11 |
0.511 |
1.093 |
-2.160 |
| H12 |
-1.051 |
1.092 |
1.329 |
| H13 |
-1.051 |
1.092 |
-1.329 |
| H14 |
1.033 |
-1.141 |
-1.249 |
| H15 |
1.033 |
-1.141 |
1.249 |
| H16 |
-0.535 |
-1.189 |
-2.081 |
| H17 |
-0.535 |
-1.189 |
2.081 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0962 | 1.0992 | 1.5325 | 1.5325 | 2.5147 | 2.5147 | 2.8851 | 3.8331 | 2.1763 | 2.1763 | 2.1634 | 2.1634 | 2.7936 | 2.7936 | 3.4698 | 3.4698 |
H2 | 1.0962 | | 1.7621 | 2.1750 | 2.1750 | 3.4710 | 3.4710 | 3.8636 | 4.8489 | 2.5362 | 2.5362 | 2.4847 | 2.4847 | 3.8023 | 3.8023 | 4.3119 | 4.3119 | H3 | 1.0992 | 1.7621 | | 2.1584 | 2.1584 | 2.8036 | 2.8036 | 3.3359 | 4.1356 | 2.4701 | 2.4701 | 3.0639 | 3.0639 | 2.6308 | 2.6308 | 3.8062 | 3.8062 | C4 | 1.5325 | 2.1750 | 2.1584 | | 2.5206 | 2.9042 | 1.5279 | 2.4414 | 3.3286 | 1.0965 | 3.4772 | 1.0966 | 2.8108 | 3.3105 | 2.1588 | 3.8974 | 2.1678 | C5 | 1.5325 | 2.1750 | 2.1584 | 2.5206 | | 1.5279 | 2.9042 | 2.4414 | 3.3286 | 3.4772 | 1.0965 | 2.8108 | 1.0966 | 2.1588 | 3.3105 | 2.1678 | 3.8974 | C6 | 2.5147 | 3.4710 | 2.8036 | 2.9042 | 1.5279 | | 2.4201 | 1.4657 | 2.0408 | 3.8904 | 2.1636 | 3.3202 | 2.1432 | 1.1070 | 2.6968 | 1.0954 | 3.3573 | C7 | 2.5147 | 3.4710 | 2.8036 | 1.5279 | 2.9042 | 2.4201 | | 1.4657 | 2.0408 | 2.1636 | 3.8904 | 2.1432 | 3.3202 | 2.6968 | 1.1070 | 3.3573 | 1.0954 | N8 | 2.8851 | 3.8636 | 3.3359 | 2.4414 | 2.4414 | 1.4657 | 1.4657 | | 1.0175 | 3.3943 | 3.3943 | 2.6906 | 2.6906 | 2.1463 | 2.1463 | 2.0886 | 2.0886 | H9 | 3.8331 | 4.8489 | 4.1356 | 3.3286 | 3.3286 | 2.0408 | 2.0408 | 1.0175 | | 4.1714 | 4.1714 | 3.5971 | 3.5971 | 2.4508 | 2.4508 | 2.3422 | 2.3422 | H10 | 2.1763 | 2.5362 | 2.4701 | 1.0965 | 3.4772 | 3.8904 | 2.1636 | 3.3943 | 4.1714 | | 4.3198 | 1.7699 | 3.8225 | 4.1093 | 2.4682 | 4.9283 | 2.5118 | H11 | 2.1763 | 2.5362 | 2.4701 | 3.4772 | 1.0965 | 2.1636 | 3.8904 | 3.3943 | 4.1714 | 4.3198 | | 3.8225 | 1.7699 | 2.4682 | 4.1093 | 2.5118 | 4.9283 | H12 | 2.1634 | 2.4847 | 3.0639 | 1.0966 | 2.8108 | 3.3202 | 2.1432 | 2.6906 | 3.5971 | 1.7699 | 3.8225 | | 2.6573 | 3.9969 | 3.0554 | 4.1345 | 2.4566 | H13 | 2.1634 | 2.4847 | 3.0639 | 2.8108 | 1.0966 | 2.1432 | 3.3202 | 2.6906 | 3.5971 | 3.8225 | 1.7699 | 2.6573 | | 3.0554 | 3.9969 | 2.4566 | 4.1345 | H14 | 2.7936 | 3.8023 | 2.6308 | 3.3105 | 2.1588 | 1.1070 | 2.6968 | 2.1463 | 2.4508 | 4.1093 | 2.4682 | 3.9969 | 3.0554 | | 2.4988 | 1.7756 | 3.6813 | H15 | 2.7936 | 3.8023 | 2.6308 | 2.1588 | 3.3105 | 2.6968 | 1.1070 | 2.1463 | 2.4508 | 2.4682 | 4.1093 | 3.0554 | 3.9969 | 2.4988 | | 3.6813 | 1.7756 | H16 | 3.4698 | 4.3119 | 3.8062 | 3.8974 | 2.1678 | 1.0954 | 3.3573 | 2.0886 | 2.3422 | 4.9283 | 2.5118 | 4.1345 | 2.4566 | 1.7756 | 3.6813 | | 4.1618 | H17 | 3.4698 | 4.3119 | 3.8062 | 2.1678 | 3.8974 | 3.3573 | 1.0954 | 2.0886 | 2.3422 | 2.5118 | 4.9283 | 2.4566 | 4.1345 | 3.6813 | 1.7756 | 4.1618 | |
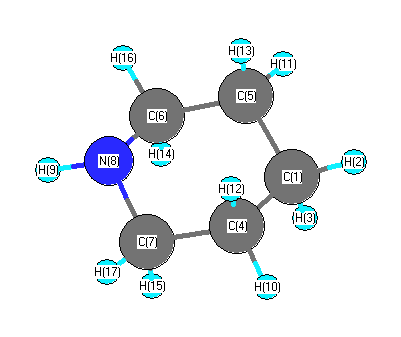 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.510 |
|
C1 |
C4 |
H10 |
110.654 |
| C1 |
C4 |
H12 |
109.639 |
|
C1 |
C5 |
C6 |
110.510 |
| C1 |
C5 |
H11 |
110.654 |
|
C1 |
C5 |
H13 |
109.639 |
| H2 |
C1 |
H3 |
106.761 |
|
H2 |
C1 |
C4 |
110.575 |
| H2 |
C1 |
C5 |
110.575 |
|
H3 |
C1 |
C4 |
109.089 |
| H3 |
C1 |
C5 |
109.089 |
|
C4 |
C1 |
C5 |
110.652 |
| C4 |
C7 |
N8 |
109.265 |
|
C4 |
C7 |
H15 |
108.993 |
| C4 |
C7 |
H17 |
110.368 |
|
C5 |
C6 |
N8 |
109.265 |
| C5 |
C6 |
H14 |
108.993 |
|
C5 |
C6 |
H16 |
110.368 |
| C6 |
C5 |
H11 |
109.971 |
|
C6 |
C5 |
H13 |
108.378 |
| C6 |
N8 |
C7 |
111.291 |
|
C6 |
N8 |
H9 |
109.205 |
| C7 |
C4 |
H10 |
109.971 |
|
C7 |
C4 |
H12 |
108.378 |
| C7 |
N8 |
H9 |
109.205 |
|
N8 |
C6 |
H14 |
112.335 |
| N8 |
C6 |
H16 |
108.408 |
|
N8 |
C7 |
H15 |
112.335 |
| N8 |
C7 |
H17 |
108.408 |
|
H10 |
C4 |
H12 |
107.614 |
| H11 |
C5 |
H13 |
107.614 |
|
H14 |
C6 |
H16 |
107.457 |
| H15 |
C7 |
H17 |
107.457 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -251.053251 |
| Energy at 298.15K | -251.067712 |
| Nuclear repulsion energy | 258.589914 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3519 |
3303 |
1.08 |
|
|
|
| 2 |
A' |
3146 |
2953 |
62.80 |
|
|
|
| 3 |
A' |
3137 |
2944 |
39.58 |
|
|
|
| 4 |
A' |
3130 |
2938 |
35.71 |
|
|
|
| 5 |
A' |
3089 |
2899 |
45.79 |
|
|
|
| 6 |
A' |
3081 |
2892 |
22.44 |
|
|
|
| 7 |
A' |
3077 |
2888 |
18.50 |
|
|
|
| 8 |
A' |
1568 |
1472 |
2.48 |
|
|
|
| 9 |
A' |
1554 |
1458 |
11.73 |
|
|
|
| 10 |
A' |
1549 |
1454 |
2.11 |
|
|
|
| 11 |
A' |
1444 |
1356 |
4.99 |
|
|
|
| 12 |
A' |
1433 |
1345 |
0.54 |
|
|
|
| 13 |
A' |
1380 |
1295 |
0.45 |
|
|
|
| 14 |
A' |
1323 |
1242 |
10.37 |
|
|
|
| 15 |
A' |
1239 |
1163 |
4.42 |
|
|
|
| 16 |
A' |
1088 |
1021 |
0.84 |
|
|
|
| 17 |
A' |
1053 |
988 |
10.00 |
|
|
|
| 18 |
A' |
961 |
902 |
23.28 |
|
|
|
| 19 |
A' |
897 |
842 |
40.97 |
|
|
|
| 20 |
A' |
850 |
798 |
2.29 |
|
|
|
| 21 |
A' |
820 |
769 |
82.58 |
|
|
|
| 22 |
A' |
571 |
536 |
1.95 |
|
|
|
| 23 |
A' |
447 |
419 |
1.01 |
|
|
|
| 24 |
A' |
405 |
380 |
5.99 |
|
|
|
| 25 |
A' |
250 |
235 |
4.86 |
|
|
|
| 26 |
A" |
3144 |
2951 |
27.28 |
|
|
|
| 27 |
A" |
3129 |
2936 |
61.51 |
|
|
|
| 28 |
A" |
3085 |
2896 |
12.73 |
|
|
|
| 29 |
A" |
3076 |
2887 |
25.06 |
|
|
|
| 30 |
A" |
1551 |
1456 |
4.25 |
|
|
|
| 31 |
A" |
1548 |
1453 |
4.19 |
|
|
|
| 32 |
A" |
1531 |
1437 |
8.73 |
|
|
|
| 33 |
A" |
1434 |
1346 |
0.25 |
|
|
|
| 34 |
A" |
1420 |
1333 |
2.00 |
|
|
|
| 35 |
A" |
1391 |
1305 |
1.20 |
|
|
|
| 36 |
A" |
1337 |
1255 |
0.73 |
|
|
|
| 37 |
A" |
1258 |
1181 |
12.47 |
|
|
|
| 38 |
A" |
1182 |
1110 |
14.45 |
|
|
|
| 39 |
A" |
1152 |
1081 |
1.81 |
|
|
|
| 40 |
A" |
1102 |
1034 |
3.27 |
|
|
|
| 41 |
A" |
980 |
920 |
5.14 |
|
|
|
| 42 |
A" |
910 |
854 |
0.00 |
|
|
|
| 43 |
A" |
835 |
783 |
0.07 |
|
|
|
| 44 |
A" |
463 |
435 |
0.93 |
|
|
|
| 45 |
A" |
247 |
232 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35891.6 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 33687.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.646 |
1.322 |
0.000 |
| H2 |
0.585 |
2.417 |
0.000 |
| H3 |
1.715 |
1.066 |
0.000 |
| C4 |
-0.008 |
0.744 |
1.260 |
| C5 |
-0.008 |
0.744 |
-1.260 |
| C6 |
-0.008 |
-0.789 |
-1.213 |
| C7 |
-0.008 |
-0.789 |
1.213 |
| N8 |
-0.616 |
-1.348 |
0.000 |
| H9 |
-1.608 |
-1.111 |
0.000 |
| H10 |
0.509 |
1.095 |
2.162 |
| H11 |
0.509 |
1.095 |
-2.162 |
| H12 |
-1.046 |
1.100 |
1.328 |
| H13 |
-1.046 |
1.100 |
-1.328 |
| H14 |
1.027 |
-1.155 |
-1.256 |
| H15 |
1.027 |
-1.155 |
1.256 |
| H16 |
-0.528 |
-1.206 |
-2.081 |
| H17 |
-0.528 |
-1.206 |
2.081 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0962 | 1.0992 | 1.5327 | 1.5327 | 2.5206 | 2.5206 | 2.9539 | 3.3175 | 2.1782 | 2.1782 | 2.1623 | 2.1623 | 2.8030 | 2.8030 | 3.4792 | 3.4792 |
H2 | 1.0962 | | 1.7615 | 2.1762 | 2.1762 | 3.4778 | 3.4778 | 3.9519 | 4.1541 | 2.5349 | 2.5349 | 2.4810 | 2.4810 | 3.8115 | 3.8115 | 4.3239 | 4.3239 | H3 | 1.0992 | 1.7615 | | 2.1582 | 2.1582 | 2.8064 | 2.8064 | 3.3558 | 3.9729 | 2.4757 | 2.4757 | 3.0636 | 3.0636 | 2.6419 | 2.6419 | 3.8113 | 3.8113 | C4 | 1.5327 | 2.1762 | 2.1582 | | 2.5200 | 2.9091 | 1.5335 | 2.5172 | 2.7556 | 1.0972 | 3.4786 | 1.0994 | 2.8113 | 3.3173 | 2.1625 | 3.9037 | 2.1794 | C5 | 1.5327 | 2.1762 | 2.1582 | 2.5200 | | 1.5335 | 2.9091 | 2.5172 | 2.7556 | 3.4786 | 1.0972 | 2.8113 | 1.0994 | 2.1625 | 3.3173 | 2.1794 | 3.9037 | C6 | 2.5206 | 3.4778 | 2.8064 | 2.9091 | 1.5335 | | 2.4251 | 1.4676 | 2.0340 | 3.8992 | 2.1721 | 3.3317 | 2.1581 | 1.0985 | 2.7012 | 1.0956 | 3.3607 | C7 | 2.5206 | 3.4778 | 2.8064 | 1.5335 | 2.9091 | 2.4251 | | 1.4676 | 2.0340 | 2.1721 | 3.8992 | 2.1581 | 3.3317 | 2.7012 | 1.0985 | 3.3607 | 1.0956 | N8 | 2.9539 | 3.9519 | 3.3558 | 2.5172 | 2.5172 | 1.4676 | 1.4676 | | 1.0202 | 3.4514 | 3.4514 | 2.8183 | 2.8183 | 2.0771 | 2.0771 | 2.0879 | 2.0879 | H9 | 3.3175 | 4.1541 | 3.9729 | 2.7556 | 2.7556 | 2.0340 | 2.0340 | 1.0202 | | 3.7454 | 3.7454 | 2.6403 | 2.6403 | 2.9198 | 2.9198 | 2.3466 | 2.3466 | H10 | 2.1782 | 2.5349 | 2.4757 | 1.0972 | 3.4786 | 3.8992 | 2.1721 | 3.4514 | 3.7454 | | 4.3240 | 1.7641 | 3.8208 | 4.1244 | 2.4806 | 4.9375 | 2.5261 | H11 | 2.1782 | 2.5349 | 2.4757 | 3.4786 | 1.0972 | 2.1721 | 3.8992 | 3.4514 | 3.7454 | 4.3240 | | 3.8208 | 1.7641 | 2.4806 | 4.1244 | 2.5261 | 4.9375 | H12 | 2.1623 | 2.4810 | 3.0636 | 1.0994 | 2.8113 | 3.3317 | 2.1581 | 2.8183 | 2.6403 | 1.7641 | 3.8208 | | 2.6566 | 4.0070 | 3.0636 | 4.1486 | 2.4805 | H13 | 2.1623 | 2.4810 | 3.0636 | 2.8113 | 1.0994 | 2.1581 | 3.3317 | 2.8183 | 2.6403 | 3.8208 | 1.7641 | 2.6566 | | 3.0636 | 4.0070 | 2.4805 | 4.1486 | H14 | 2.8030 | 3.8115 | 2.6419 | 3.3173 | 2.1625 | 1.0985 | 2.7012 | 2.0771 | 2.9198 | 4.1244 | 2.4806 | 4.0070 | 3.0636 | | 2.5110 | 1.7620 | 3.6819 | H15 | 2.8030 | 3.8115 | 2.6419 | 2.1625 | 3.3173 | 2.7012 | 1.0985 | 2.0771 | 2.9198 | 2.4806 | 4.1244 | 3.0636 | 4.0070 | 2.5110 | | 3.6819 | 1.7620 | H16 | 3.4792 | 4.3239 | 3.8113 | 3.9037 | 2.1794 | 1.0956 | 3.3607 | 2.0879 | 2.3466 | 4.9375 | 2.5261 | 4.1486 | 2.4805 | 1.7620 | 3.6819 | | 4.1623 | H17 | 3.4792 | 4.3239 | 3.8113 | 2.1794 | 3.9037 | 3.3607 | 1.0956 | 2.0879 | 2.3466 | 2.5261 | 4.9375 | 2.4805 | 4.1486 | 3.6819 | 1.7620 | 4.1623 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.584 |
|
C1 |
C4 |
H10 |
110.746 |
| C1 |
C4 |
H12 |
109.367 |
|
C1 |
C5 |
C6 |
110.584 |
| C1 |
C5 |
H11 |
110.746 |
|
C1 |
C5 |
H13 |
109.367 |
| H2 |
C1 |
H3 |
106.720 |
|
H2 |
C1 |
C4 |
110.649 |
| H2 |
C1 |
C5 |
110.649 |
|
H3 |
C1 |
C4 |
109.066 |
| H3 |
C1 |
C5 |
109.066 |
|
C4 |
C1 |
C5 |
110.589 |
| C4 |
C7 |
N8 |
114.004 |
|
C4 |
C7 |
H15 |
109.381 |
| C4 |
C7 |
H17 |
110.887 |
|
C5 |
C6 |
N8 |
114.004 |
| C5 |
C6 |
H14 |
109.381 |
|
C5 |
C6 |
H16 |
110.887 |
| C6 |
C5 |
H11 |
110.211 |
|
C6 |
C5 |
H13 |
108.989 |
| C6 |
N8 |
C7 |
111.422 |
|
C6 |
N8 |
H9 |
108.343 |
| C7 |
C4 |
H10 |
110.211 |
|
C7 |
C4 |
H12 |
108.989 |
| C7 |
N8 |
H9 |
108.343 |
|
N8 |
C6 |
H14 |
107.206 |
| N8 |
C6 |
H16 |
108.210 |
|
N8 |
C7 |
H15 |
107.206 |
| N8 |
C7 |
H17 |
108.210 |
|
H10 |
C4 |
H12 |
106.852 |
| H11 |
C5 |
H13 |
106.852 |
|
H14 |
C6 |
H16 |
106.849 |
| H15 |
C7 |
H17 |
106.849 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
