Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -251.554190 |
| Energy at 298.15K | -251.568393 |
| HF Energy | -251.554190 |
| Nuclear repulsion energy | 257.840731 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3409 |
3353 |
3.19 |
|
|
|
| 2 |
A' |
3030 |
2980 |
68.72 |
|
|
|
| 3 |
A' |
3014 |
2964 |
25.12 |
|
|
|
| 4 |
A' |
3010 |
2960 |
46.50 |
|
|
|
| 5 |
A' |
2978 |
2929 |
15.70 |
|
|
|
| 6 |
A' |
2956 |
2908 |
29.10 |
|
|
|
| 7 |
A' |
2832 |
2785 |
155.35 |
|
|
|
| 8 |
A' |
1485 |
1461 |
2.15 |
|
|
|
| 9 |
A' |
1470 |
1446 |
6.93 |
|
|
|
| 10 |
A' |
1460 |
1436 |
8.26 |
|
|
|
| 11 |
A' |
1396 |
1373 |
4.45 |
|
|
|
| 12 |
A' |
1350 |
1328 |
0.26 |
|
|
|
| 13 |
A' |
1288 |
1266 |
0.15 |
|
|
|
| 14 |
A' |
1263 |
1242 |
3.37 |
|
|
|
| 15 |
A' |
1139 |
1120 |
5.15 |
|
|
|
| 16 |
A' |
1049 |
1032 |
6.62 |
|
|
|
| 17 |
A' |
1033 |
1016 |
5.95 |
|
|
|
| 18 |
A' |
905 |
890 |
1.73 |
|
|
|
| 19 |
A' |
856 |
842 |
13.74 |
|
|
|
| 20 |
A' |
819 |
805 |
0.87 |
|
|
|
| 21 |
A' |
759 |
747 |
84.07 |
|
|
|
| 22 |
A' |
532 |
523 |
18.67 |
|
|
|
| 23 |
A' |
424 |
417 |
7.23 |
|
|
|
| 24 |
A' |
396 |
389 |
1.25 |
|
|
|
| 25 |
A' |
243 |
239 |
1.08 |
|
|
|
| 26 |
A" |
3026 |
2976 |
20.97 |
|
|
|
| 27 |
A" |
3009 |
2960 |
80.19 |
|
|
|
| 28 |
A" |
2978 |
2929 |
23.51 |
|
|
|
| 29 |
A" |
2825 |
2778 |
33.07 |
|
|
|
| 30 |
A" |
1472 |
1448 |
0.17 |
|
|
|
| 31 |
A" |
1456 |
1432 |
0.10 |
|
|
|
| 32 |
A" |
1447 |
1423 |
3.29 |
|
|
|
| 33 |
A" |
1351 |
1329 |
0.14 |
|
|
|
| 34 |
A" |
1331 |
1309 |
19.86 |
|
|
|
| 35 |
A" |
1313 |
1292 |
14.91 |
|
|
|
| 36 |
A" |
1268 |
1247 |
3.23 |
|
|
|
| 37 |
A" |
1162 |
1143 |
7.62 |
|
|
|
| 38 |
A" |
1142 |
1123 |
4.36 |
|
|
|
| 39 |
A" |
1117 |
1098 |
13.88 |
|
|
|
| 40 |
A" |
1050 |
1032 |
2.60 |
|
|
|
| 41 |
A" |
961 |
945 |
0.85 |
|
|
|
| 42 |
A" |
873 |
858 |
0.23 |
|
|
|
| 43 |
A" |
802 |
789 |
0.38 |
|
|
|
| 44 |
A" |
440 |
433 |
0.83 |
|
|
|
| 45 |
A" |
248 |
244 |
0.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34181.9 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 33617.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.636 |
1.336 |
0.000 |
| H2 |
0.563 |
2.438 |
0.000 |
| H3 |
1.716 |
1.089 |
0.000 |
| C4 |
-0.015 |
0.752 |
1.267 |
| C5 |
-0.015 |
0.752 |
-1.267 |
| C6 |
-0.015 |
-0.783 |
-1.216 |
| C7 |
-0.015 |
-0.783 |
1.216 |
| N8 |
-0.695 |
-1.235 |
0.000 |
| H9 |
-0.775 |
-2.255 |
0.000 |
| H10 |
0.517 |
1.097 |
2.172 |
| H11 |
0.517 |
1.097 |
-2.172 |
| H12 |
-1.061 |
1.100 |
1.342 |
| H13 |
-1.061 |
1.100 |
-1.342 |
| H14 |
1.043 |
-1.141 |
-1.281 |
| H15 |
1.043 |
-1.141 |
1.281 |
| H16 |
-0.552 |
-1.193 |
-2.090 |
| H17 |
-0.552 |
-1.193 |
2.090 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.1046 | 1.1077 | 1.5398 | 1.5398 | 2.5285 | 2.5285 | 2.8954 | 3.8587 | 2.1885 | 2.1885 | 2.1763 | 2.1763 | 2.8182 | 2.8182 | 3.4893 | 3.4893 |
H2 | 1.1046 | | 1.7748 | 2.1873 | 2.1873 | 3.4914 | 3.4914 | 3.8830 | 4.8808 | 2.5533 | 2.5533 | 2.4958 | 2.4958 | 3.8317 | 3.8317 | 4.3357 | 4.3357 | H3 | 1.1077 | 1.7748 | | 2.1712 | 2.1712 | 2.8243 | 2.8243 | 3.3485 | 4.1697 | 2.4810 | 2.4810 | 3.0839 | 3.0839 | 2.6580 | 2.6580 | 3.8360 | 3.8360 | C4 | 1.5398 | 2.1873 | 2.1712 | | 2.5348 | 2.9197 | 1.5357 | 2.4531 | 3.3508 | 1.1048 | 3.4975 | 1.1052 | 2.8328 | 3.3459 | 2.1683 | 3.9169 | 2.1790 | C5 | 1.5398 | 2.1873 | 2.1712 | 2.5348 | | 1.5357 | 2.9197 | 2.4531 | 3.3508 | 3.4975 | 1.1048 | 2.8328 | 1.1052 | 2.1683 | 3.3459 | 2.1790 | 3.9169 | C6 | 2.5285 | 3.4914 | 2.8243 | 2.9197 | 1.5357 | | 2.4327 | 1.4652 | 2.0556 | 3.9115 | 2.1751 | 3.3446 | 2.1579 | 1.1184 | 2.7355 | 1.1043 | 3.3745 | C7 | 2.5285 | 3.4914 | 2.8243 | 1.5357 | 2.9197 | 2.4327 | | 1.4652 | 2.0556 | 2.1751 | 3.9115 | 2.1579 | 3.3446 | 2.7355 | 1.1184 | 3.3745 | 1.1043 | N8 | 2.8954 | 3.8830 | 3.3485 | 2.4531 | 2.4531 | 1.4652 | 1.4652 | | 1.0233 | 3.4099 | 3.4099 | 2.7184 | 2.7184 | 2.1610 | 2.1610 | 2.0951 | 2.0951 | H9 | 3.8587 | 4.8808 | 4.1697 | 3.3508 | 3.3508 | 2.0556 | 2.0556 | 1.0233 | | 4.1985 | 4.1985 | 3.6254 | 3.6254 | 2.4874 | 2.4874 | 2.3549 | 2.3549 | H10 | 2.1885 | 2.5533 | 2.4810 | 1.1048 | 3.4975 | 3.9115 | 2.1751 | 3.4099 | 4.1985 | | 4.3443 | 1.7829 | 3.8522 | 4.1484 | 2.4658 | 4.9550 | 2.5287 | H11 | 2.1885 | 2.5533 | 2.4810 | 3.4975 | 1.1048 | 2.1751 | 3.9115 | 3.4099 | 4.1985 | 4.3443 | | 3.8522 | 1.7829 | 2.4658 | 4.1484 | 2.5287 | 4.9550 | H12 | 2.1763 | 2.4958 | 3.0839 | 1.1052 | 2.8328 | 3.3446 | 2.1579 | 2.7184 | 3.6254 | 1.7829 | 3.8522 | | 2.6841 | 4.0409 | 3.0745 | 4.1589 | 2.4654 | H13 | 2.1763 | 2.4958 | 3.0839 | 2.8328 | 1.1052 | 2.1579 | 3.3446 | 2.7184 | 3.6254 | 3.8522 | 1.7829 | 2.6841 | | 3.0745 | 4.0409 | 2.4654 | 4.1589 | H14 | 2.8182 | 3.8317 | 2.6580 | 3.3459 | 2.1683 | 1.1184 | 2.7355 | 2.1610 | 2.4874 | 4.1484 | 2.4658 | 4.0409 | 3.0745 | | 2.5618 | 1.7887 | 3.7292 | H15 | 2.8182 | 3.8317 | 2.6580 | 2.1683 | 3.3459 | 2.7355 | 1.1184 | 2.1610 | 2.4874 | 2.4658 | 4.1484 | 3.0745 | 4.0409 | 2.5618 | | 3.7292 | 1.7887 | H16 | 3.4893 | 4.3357 | 3.8360 | 3.9169 | 2.1790 | 1.1043 | 3.3745 | 2.0951 | 2.3549 | 4.9550 | 2.5287 | 4.1589 | 2.4654 | 1.7887 | 3.7292 | | 4.1796 | H17 | 3.4893 | 4.3357 | 3.8360 | 2.1790 | 3.9169 | 3.3745 | 1.1043 | 2.0951 | 2.3549 | 2.5287 | 4.9550 | 2.4654 | 4.1589 | 3.7292 | 1.7887 | 4.1796 | |
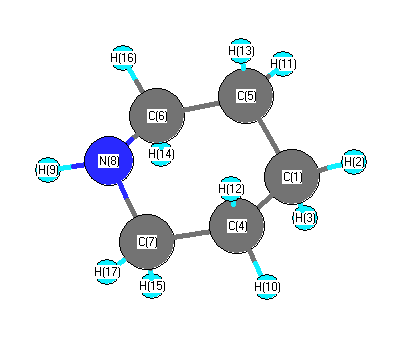 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.601 |
|
C1 |
C4 |
H10 |
110.615 |
| C1 |
C4 |
H12 |
109.638 |
|
C1 |
C5 |
C6 |
110.601 |
| C1 |
C5 |
H11 |
110.615 |
|
C1 |
C5 |
H13 |
109.638 |
| H2 |
C1 |
H3 |
106.690 |
|
H2 |
C1 |
C4 |
110.530 |
| H2 |
C1 |
C5 |
110.530 |
|
H3 |
C1 |
C4 |
109.099 |
| H3 |
C1 |
C5 |
109.099 |
|
C4 |
C1 |
C5 |
110.785 |
| C4 |
C7 |
N8 |
109.639 |
|
C4 |
C7 |
H15 |
108.544 |
| C4 |
C7 |
H17 |
110.182 |
|
C5 |
C6 |
N8 |
109.639 |
| C5 |
C6 |
H14 |
108.544 |
|
C5 |
C6 |
H16 |
110.182 |
| C6 |
C5 |
H11 |
109.848 |
|
C6 |
C5 |
H13 |
108.500 |
| C6 |
N8 |
C7 |
112.230 |
|
C6 |
N8 |
H9 |
110.123 |
| C7 |
C4 |
H10 |
109.848 |
|
C7 |
C4 |
H12 |
108.500 |
| C7 |
N8 |
H9 |
110.123 |
|
N8 |
C6 |
H14 |
112.836 |
| N8 |
C6 |
H16 |
108.432 |
|
N8 |
C7 |
H15 |
112.836 |
| N8 |
C7 |
H17 |
108.432 |
|
H10 |
C4 |
H12 |
107.563 |
| H11 |
C5 |
H13 |
107.563 |
|
H14 |
C6 |
H16 |
107.171 |
| H15 |
C7 |
H17 |
107.171 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.293 |
|
|
|
| 2 |
H |
0.147 |
|
|
|
| 3 |
H |
0.140 |
|
|
|
| 4 |
C |
-0.279 |
|
|
|
| 5 |
C |
-0.279 |
|
|
|
| 6 |
C |
-0.172 |
|
|
|
| 7 |
C |
-0.172 |
|
|
|
| 8 |
N |
-0.512 |
|
|
|
| 9 |
H |
0.297 |
|
|
|
| 10 |
H |
0.142 |
|
|
|
| 11 |
H |
0.142 |
|
|
|
| 12 |
H |
0.155 |
|
|
|
| 13 |
H |
0.155 |
|
|
|
| 14 |
H |
0.117 |
|
|
|
| 15 |
H |
0.117 |
|
|
|
| 16 |
H |
0.147 |
|
|
|
| 17 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.657 |
-0.512 |
0.000 |
0.833 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.995 |
-0.036 |
0.003 |
| y |
-0.036 |
-36.083 |
-0.001 |
| z |
0.003 |
-0.001 |
-37.824 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.042 |
-0.036 |
0.003 |
| y |
-0.036 |
3.326 |
-0.001 |
| z |
0.003 |
-0.001 |
0.716 |
|
| Polar |
|---|
| 3z2-r2 | 1.431 |
|---|
| x2-y2 | -4.912 |
|---|
| xy | -0.036 |
|---|
| xz | 0.003 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.047 |
0.337 |
0.000 |
| y |
0.337 |
9.087 |
0.001 |
| z |
0.000 |
0.001 |
9.561 |
<r2> (average value of r
2) Å
2
| <r2> |
158.980 |
| (<r2>)1/2 |
12.609 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -251.554048 |
| Energy at 298.15K | -251.568201 |
| Nuclear repulsion energy | 257.218809 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3368 |
3312 |
4.69 |
|
|
|
| 2 |
A' |
3018 |
2968 |
70.06 |
|
|
|
| 3 |
A' |
3012 |
2962 |
42.73 |
|
|
|
| 4 |
A' |
3004 |
2955 |
46.14 |
|
|
|
| 5 |
A' |
2958 |
2910 |
70.93 |
|
|
|
| 6 |
A' |
2955 |
2906 |
16.97 |
|
|
|
| 7 |
A' |
2952 |
2903 |
12.66 |
|
|
|
| 8 |
A' |
1478 |
1454 |
1.92 |
|
|
|
| 9 |
A' |
1463 |
1439 |
11.66 |
|
|
|
| 10 |
A' |
1459 |
1435 |
2.49 |
|
|
|
| 11 |
A' |
1368 |
1345 |
5.53 |
|
|
|
| 12 |
A' |
1353 |
1331 |
0.54 |
|
|
|
| 13 |
A' |
1313 |
1291 |
0.49 |
|
|
|
| 14 |
A' |
1256 |
1235 |
9.26 |
|
|
|
| 15 |
A' |
1165 |
1146 |
3.92 |
|
|
|
| 16 |
A' |
1031 |
1014 |
0.52 |
|
|
|
| 17 |
A' |
995 |
979 |
6.43 |
|
|
|
| 18 |
A' |
915 |
900 |
18.17 |
|
|
|
| 19 |
A' |
848 |
834 |
39.96 |
|
|
|
| 20 |
A' |
810 |
797 |
0.93 |
|
|
|
| 21 |
A' |
764 |
752 |
91.50 |
|
|
|
| 22 |
A' |
538 |
529 |
1.82 |
|
|
|
| 23 |
A' |
427 |
420 |
1.05 |
|
|
|
| 24 |
A' |
385 |
379 |
5.43 |
|
|
|
| 25 |
A' |
238 |
234 |
4.57 |
|
|
|
| 26 |
A" |
3016 |
2967 |
33.56 |
|
|
|
| 27 |
A" |
3003 |
2953 |
75.64 |
|
|
|
| 28 |
A" |
2955 |
2907 |
0.40 |
|
|
|
| 29 |
A" |
2949 |
2901 |
45.99 |
|
|
|
| 30 |
A" |
1467 |
1443 |
3.32 |
|
|
|
| 31 |
A" |
1457 |
1433 |
6.00 |
|
|
|
| 32 |
A" |
1445 |
1421 |
2.82 |
|
|
|
| 33 |
A" |
1350 |
1328 |
2.43 |
|
|
|
| 34 |
A" |
1340 |
1318 |
2.69 |
|
|
|
| 35 |
A" |
1316 |
1294 |
0.78 |
|
|
|
| 36 |
A" |
1274 |
1253 |
1.06 |
|
|
|
| 37 |
A" |
1189 |
1170 |
13.06 |
|
|
|
| 38 |
A" |
1102 |
1084 |
18.52 |
|
|
|
| 39 |
A" |
1098 |
1079 |
0.01 |
|
|
|
| 40 |
A" |
1048 |
1031 |
3.19 |
|
|
|
| 41 |
A" |
928 |
912 |
4.82 |
|
|
|
| 42 |
A" |
863 |
849 |
0.15 |
|
|
|
| 43 |
A" |
790 |
776 |
0.05 |
|
|
|
| 44 |
A" |
445 |
438 |
0.90 |
|
|
|
| 45 |
A" |
235 |
231 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34170.3 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 33606.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.644 |
1.331 |
0.000 |
| H2 |
0.585 |
2.434 |
0.000 |
| H3 |
1.723 |
1.074 |
0.000 |
| C4 |
-0.009 |
0.749 |
1.266 |
| C5 |
-0.009 |
0.749 |
-1.266 |
| C6 |
-0.009 |
-0.791 |
-1.216 |
| C7 |
-0.009 |
-0.791 |
1.216 |
| N8 |
-0.610 |
-1.362 |
0.000 |
| H9 |
-1.613 |
-1.132 |
0.000 |
| H10 |
0.513 |
1.104 |
2.175 |
| H11 |
0.513 |
1.104 |
-2.175 |
| H12 |
-1.056 |
1.108 |
1.340 |
| H13 |
-1.056 |
1.108 |
-1.340 |
| H14 |
1.037 |
-1.156 |
-1.265 |
| H15 |
1.037 |
-1.156 |
1.265 |
| H16 |
-0.533 |
-1.214 |
-2.092 |
| H17 |
-0.533 |
-1.214 |
2.092 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.1053 | 1.1089 | 1.5385 | 1.5385 | 2.5313 | 2.5313 | 2.9704 | 3.3406 | 2.1908 | 2.1908 | 2.1757 | 2.1757 | 2.8177 | 2.8177 | 3.4983 | 3.4983 |
H2 | 1.1053 | | 1.7739 | 2.1896 | 2.1896 | 3.4980 | 3.4980 | 3.9801 | 4.1893 | 2.5507 | 2.5507 | 2.4989 | 2.4989 | 3.8337 | 3.8337 | 4.3517 | 4.3517 | H3 | 1.1089 | 1.7739 | | 2.1694 | 2.1694 | 2.8204 | 2.8204 | 3.3724 | 3.9990 | 2.4891 | 2.4891 | 3.0847 | 3.0847 | 2.6539 | 2.6539 | 3.8340 | 3.8340 | C4 | 1.5385 | 2.1896 | 2.1694 | | 2.5318 | 2.9209 | 1.5415 | 2.5342 | 2.7777 | 1.1066 | 3.4983 | 1.1089 | 2.8310 | 3.3362 | 2.1738 | 3.9248 | 2.1937 | C5 | 1.5385 | 2.1896 | 2.1694 | 2.5318 | | 1.5415 | 2.9209 | 2.5342 | 2.7777 | 3.4983 | 1.1066 | 2.8310 | 1.1089 | 2.1738 | 3.3362 | 2.1937 | 3.9248 | C6 | 2.5313 | 3.4980 | 2.8204 | 2.9209 | 1.5415 | | 2.4313 | 1.4712 | 2.0412 | 3.9194 | 2.1874 | 3.3518 | 2.1724 | 1.1088 | 2.7166 | 1.1051 | 3.3752 | C7 | 2.5313 | 3.4980 | 2.8204 | 1.5415 | 2.9209 | 2.4313 | | 1.4712 | 2.0412 | 2.1874 | 3.9194 | 2.1724 | 3.3518 | 2.7166 | 1.1088 | 3.3752 | 1.1051 | N8 | 2.9704 | 3.9801 | 3.3724 | 2.5342 | 2.5342 | 1.4712 | 1.4712 | | 1.0293 | 3.4746 | 3.4746 | 2.8456 | 2.8456 | 2.0864 | 2.0864 | 2.0983 | 2.0983 | H9 | 3.3406 | 4.1893 | 3.9990 | 2.7777 | 2.7777 | 2.0412 | 2.0412 | 1.0293 | | 3.7749 | 3.7749 | 2.6694 | 2.6694 | 2.9363 | 2.9363 | 2.3553 | 2.3553 | H10 | 2.1908 | 2.5507 | 2.4891 | 1.1066 | 3.4983 | 3.9194 | 2.1874 | 3.4746 | 3.7749 | | 4.3501 | 1.7769 | 3.8490 | 4.1492 | 2.4924 | 4.9672 | 2.5446 | H11 | 2.1908 | 2.5507 | 2.4891 | 3.4983 | 1.1066 | 2.1874 | 3.9194 | 3.4746 | 3.7749 | 4.3501 | | 3.8490 | 1.7769 | 2.4924 | 4.1492 | 2.5446 | 4.9672 | H12 | 2.1757 | 2.4989 | 3.0847 | 1.1089 | 2.8310 | 3.3518 | 2.1724 | 2.8456 | 2.6694 | 1.7769 | 3.8490 | | 2.6798 | 4.0363 | 3.0843 | 4.1764 | 2.4963 | H13 | 2.1757 | 2.4989 | 3.0847 | 2.8310 | 1.1089 | 2.1724 | 3.3518 | 2.8456 | 2.6694 | 3.8490 | 1.7769 | 2.6798 | | 3.0843 | 4.0363 | 2.4963 | 4.1764 | H14 | 2.8177 | 3.8337 | 2.6539 | 3.3362 | 2.1738 | 1.1088 | 2.7166 | 2.0864 | 2.9363 | 4.1492 | 2.4924 | 4.0363 | 3.0843 | | 2.5296 | 1.7756 | 3.7061 | H15 | 2.8177 | 3.8337 | 2.6539 | 2.1738 | 3.3362 | 2.7166 | 1.1088 | 2.0864 | 2.9363 | 2.4924 | 4.1492 | 3.0843 | 4.0363 | 2.5296 | | 3.7061 | 1.7756 | H16 | 3.4983 | 4.3517 | 3.8340 | 3.9248 | 2.1937 | 1.1051 | 3.3752 | 2.0983 | 2.3553 | 4.9672 | 2.5446 | 4.1764 | 2.4963 | 1.7756 | 3.7061 | | 4.1834 | H17 | 3.4983 | 4.3517 | 3.8340 | 2.1937 | 3.9248 | 3.3752 | 1.1051 | 2.0983 | 2.3553 | 2.5446 | 4.9672 | 2.4963 | 4.1764 | 3.7061 | 1.7756 | 4.1834 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.540 |
|
C1 |
C4 |
H10 |
110.780 |
| C1 |
C4 |
H12 |
109.463 |
|
C1 |
C5 |
C6 |
110.540 |
| C1 |
C5 |
H11 |
110.780 |
|
C1 |
C5 |
H13 |
109.463 |
| H2 |
C1 |
H3 |
106.476 |
|
H2 |
C1 |
C4 |
110.768 |
| H2 |
C1 |
C5 |
110.768 |
|
H3 |
C1 |
C4 |
108.983 |
| H3 |
C1 |
C5 |
108.983 |
|
C4 |
C1 |
C5 |
110.739 |
| C4 |
C7 |
N8 |
114.504 |
|
C4 |
C7 |
H15 |
109.120 |
| C4 |
C7 |
H17 |
110.892 |
|
C5 |
C6 |
N8 |
114.504 |
| C5 |
C6 |
H14 |
109.120 |
|
C5 |
C6 |
H16 |
110.892 |
| C6 |
C5 |
H11 |
110.308 |
|
C6 |
C5 |
H13 |
109.006 |
| C6 |
N8 |
C7 |
111.439 |
|
C6 |
N8 |
H9 |
108.134 |
| C7 |
C4 |
H10 |
110.308 |
|
C7 |
C4 |
H12 |
109.006 |
| C7 |
N8 |
H9 |
108.134 |
|
N8 |
C6 |
H14 |
107.096 |
| N8 |
C6 |
H16 |
108.229 |
|
N8 |
C7 |
H15 |
107.096 |
| N8 |
C7 |
H17 |
108.229 |
|
H10 |
C4 |
H12 |
106.647 |
| H11 |
C5 |
H13 |
106.647 |
|
H14 |
C6 |
H16 |
106.648 |
| H15 |
C7 |
H17 |
106.648 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.284 |
|
|
|
| 2 |
H |
0.147 |
|
|
|
| 3 |
H |
0.143 |
|
|
|
| 4 |
C |
-0.291 |
|
|
|
| 5 |
C |
-0.291 |
|
|
|
| 6 |
C |
-0.191 |
|
|
|
| 7 |
C |
-0.191 |
|
|
|
| 8 |
N |
-0.491 |
|
|
|
| 9 |
H |
0.287 |
|
|
|
| 10 |
H |
0.143 |
|
|
|
| 11 |
H |
0.143 |
|
|
|
| 12 |
H |
0.137 |
|
|
|
| 13 |
H |
0.137 |
|
|
|
| 14 |
H |
0.150 |
|
|
|
| 15 |
H |
0.150 |
|
|
|
| 16 |
H |
0.150 |
|
|
|
| 17 |
H |
0.150 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.289 |
1.132 |
0.000 |
1.169 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.736 |
-0.287 |
0.000 |
| y |
-0.287 |
-43.197 |
0.000 |
| z |
0.000 |
0.000 |
-37.811 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.768 |
-0.287 |
0.000 |
| y |
-0.287 |
-5.923 |
0.000 |
| z |
0.000 |
0.000 |
2.156 |
|
| Polar |
|---|
| 3z2-r2 | 4.311 |
|---|
| x2-y2 | 6.460 |
|---|
| xy | -0.287 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.327 |
0.294 |
0.000 |
| y |
0.294 |
8.621 |
0.000 |
| z |
0.000 |
0.000 |
9.591 |
<r2> (average value of r
2) Å
2
| <r2> |
158.980 |
| (<r2>)1/2 |
12.609 |
