Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -311.035093 |
| Energy at 298.15K | -311.050931 |
| HF Energy | -310.213966 |
| Nuclear repulsion energy | 316.287113 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3253 |
3005 |
49.52 |
|
|
|
| 2 |
A1 |
3189 |
2946 |
40.53 |
|
|
|
| 3 |
A1 |
3171 |
2929 |
22.49 |
|
|
|
| 4 |
A1 |
3119 |
2881 |
78.39 |
|
|
|
| 5 |
A1 |
1647 |
1521 |
0.24 |
|
|
|
| 6 |
A1 |
1615 |
1491 |
7.38 |
|
|
|
| 7 |
A1 |
1600 |
1478 |
0.01 |
|
|
|
| 8 |
A1 |
1566 |
1446 |
0.34 |
|
|
|
| 9 |
A1 |
1524 |
1408 |
1.62 |
|
|
|
| 10 |
A1 |
1432 |
1323 |
2.99 |
|
|
|
| 11 |
A1 |
1254 |
1158 |
4.49 |
|
|
|
| 12 |
A1 |
1141 |
1054 |
9.94 |
|
|
|
| 13 |
A1 |
1083 |
1000 |
17.03 |
|
|
|
| 14 |
A1 |
981 |
906 |
6.42 |
|
|
|
| 15 |
A1 |
461 |
426 |
0.96 |
|
|
|
| 16 |
A1 |
335 |
310 |
0.24 |
|
|
|
| 17 |
A1 |
110 |
102 |
0.18 |
|
|
|
| 18 |
A2 |
3248 |
3001 |
0.00 |
|
|
|
| 19 |
A2 |
3226 |
2980 |
0.00 |
|
|
|
| 20 |
A2 |
3145 |
2905 |
0.00 |
|
|
|
| 21 |
A2 |
1605 |
1483 |
0.00 |
|
|
|
| 22 |
A2 |
1399 |
1292 |
0.00 |
|
|
|
| 23 |
A2 |
1351 |
1248 |
0.00 |
|
|
|
| 24 |
A2 |
1260 |
1164 |
0.00 |
|
|
|
| 25 |
A2 |
951 |
879 |
0.00 |
|
|
|
| 26 |
A2 |
806 |
744 |
0.00 |
|
|
|
| 27 |
A2 |
249 |
230 |
0.00 |
|
|
|
| 28 |
A2 |
143 |
132 |
0.00 |
|
|
|
| 29 |
A2 |
75 |
69 |
0.00 |
|
|
|
| 30 |
B1 |
3249 |
3001 |
137.96 |
|
|
|
| 31 |
B1 |
3226 |
2980 |
1.96 |
|
|
|
| 32 |
B1 |
3147 |
2907 |
103.48 |
|
|
|
| 33 |
B1 |
1605 |
1483 |
13.66 |
|
|
|
| 34 |
B1 |
1403 |
1296 |
0.56 |
|
|
|
| 35 |
B1 |
1360 |
1256 |
3.36 |
|
|
|
| 36 |
B1 |
1285 |
1187 |
6.09 |
|
|
|
| 37 |
B1 |
965 |
892 |
3.54 |
|
|
|
| 38 |
B1 |
809 |
747 |
2.14 |
|
|
|
| 39 |
B1 |
253 |
233 |
0.46 |
|
|
|
| 40 |
B1 |
171 |
158 |
6.69 |
|
|
|
| 41 |
B1 |
58 |
54 |
0.31 |
|
|
|
| 42 |
B2 |
3253 |
3005 |
21.34 |
|
|
|
| 43 |
B2 |
3188 |
2945 |
1.88 |
|
|
|
| 44 |
B2 |
3170 |
2929 |
39.41 |
|
|
|
| 45 |
B2 |
3107 |
2870 |
8.44 |
|
|
|
| 46 |
B2 |
1629 |
1504 |
9.68 |
|
|
|
| 47 |
B2 |
1613 |
1490 |
1.89 |
|
|
|
| 48 |
B2 |
1599 |
1477 |
1.70 |
|
|
|
| 49 |
B2 |
1527 |
1410 |
5.46 |
|
|
|
| 50 |
B2 |
1516 |
1401 |
54.35 |
|
|
|
| 51 |
B2 |
1415 |
1307 |
21.49 |
|
|
|
| 52 |
B2 |
1270 |
1173 |
233.90 |
|
|
|
| 53 |
B2 |
1200 |
1108 |
2.04 |
|
|
|
| 54 |
B2 |
1119 |
1034 |
0.37 |
|
|
|
| 55 |
B2 |
955 |
882 |
1.36 |
|
|
|
| 56 |
B2 |
542 |
500 |
5.32 |
|
|
|
| 57 |
B2 |
268 |
248 |
0.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 44918.1 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 41490.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.367 |
| C2 |
0.000 |
1.174 |
-0.402 |
| C3 |
0.000 |
-1.174 |
-0.402 |
| C4 |
0.000 |
2.363 |
0.534 |
| C5 |
0.000 |
-2.363 |
0.534 |
| C6 |
0.000 |
3.689 |
-0.215 |
| C7 |
0.000 |
-3.689 |
-0.215 |
| H8 |
0.881 |
1.200 |
-1.052 |
| H9 |
-0.881 |
1.200 |
-1.052 |
| H10 |
-0.881 |
-1.200 |
-1.052 |
| H11 |
0.881 |
-1.200 |
-1.052 |
| H12 |
0.874 |
2.292 |
1.179 |
| H13 |
-0.874 |
2.292 |
1.179 |
| H14 |
-0.874 |
-2.292 |
1.179 |
| H15 |
0.874 |
-2.292 |
1.179 |
| H16 |
0.000 |
4.528 |
0.477 |
| H17 |
-0.880 |
3.784 |
-0.849 |
| H18 |
0.880 |
3.784 |
-0.849 |
| H19 |
0.000 |
-4.528 |
0.477 |
| H20 |
0.880 |
-3.784 |
-0.849 |
| H21 |
-0.880 |
-3.784 |
-0.849 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
H21 |
| O1 | | 1.4033 | 1.4033 | 2.3689 | 2.3689 | 3.7345 | 3.7345 | 2.0568 | 2.0568 | 2.0568 | 2.0568 | 2.5842 | 2.5842 | 2.5842 | 2.5842 | 4.5290 | 4.0712 | 4.0712 | 4.5290 | 4.0712 | 4.0712 |
C2 | 1.4033 | | 2.3475 | 1.5135 | 3.6586 | 2.5222 | 4.8664 | 1.0950 | 1.0950 | 2.6143 | 2.6143 | 2.1251 | 2.1251 | 3.9087 | 3.9087 | 3.4674 | 2.7909 | 2.7909 | 5.7689 | 5.0554 | 5.0554 | C3 | 1.4033 | 2.3475 | | 3.6586 | 1.5135 | 4.8664 | 2.5222 | 2.6143 | 2.6143 | 1.0950 | 1.0950 | 3.9087 | 3.9087 | 2.1251 | 2.1251 | 5.7689 | 5.0554 | 5.0554 | 3.4674 | 2.7909 | 2.7909 | C4 | 2.3689 | 1.5135 | 3.6586 | | 4.7260 | 1.5226 | 6.0981 | 2.1547 | 2.1547 | 3.9986 | 3.9986 | 1.0886 | 1.0886 | 4.7803 | 4.7803 | 2.1654 | 2.1696 | 2.1696 | 6.8909 | 6.3622 | 6.3622 | C5 | 2.3689 | 3.6586 | 1.5135 | 4.7260 | | 6.0981 | 1.5226 | 3.9986 | 3.9986 | 2.1547 | 2.1547 | 4.7803 | 4.7803 | 1.0886 | 1.0886 | 6.8909 | 6.3622 | 6.3622 | 2.1654 | 2.1696 | 2.1696 | C6 | 3.7345 | 2.5222 | 4.8664 | 1.5226 | 6.0981 | | 7.3780 | 2.7697 | 2.7697 | 5.0382 | 5.0382 | 2.1581 | 2.1581 | 6.2034 | 6.2034 | 1.0873 | 1.0889 | 1.0889 | 8.2458 | 7.5517 | 7.5517 | C7 | 3.7345 | 4.8664 | 2.5222 | 6.0981 | 1.5226 | 7.3780 | | 5.0382 | 5.0382 | 2.7697 | 2.7697 | 6.2034 | 6.2034 | 2.1581 | 2.1581 | 8.2458 | 7.5517 | 7.5517 | 1.0873 | 1.0889 | 1.0889 | H8 | 2.0568 | 1.0950 | 2.6143 | 2.1547 | 3.9986 | 2.7697 | 5.0382 | | 1.7621 | 2.9779 | 2.4007 | 2.4843 | 3.0415 | 4.5007 | 4.1446 | 3.7665 | 3.1334 | 2.5920 | 5.9938 | 4.9889 | 5.2904 | H9 | 2.0568 | 1.0950 | 2.6143 | 2.1547 | 3.9986 | 2.7697 | 5.0382 | 1.7621 | | 2.4007 | 2.9779 | 3.0415 | 2.4843 | 4.1446 | 4.5007 | 3.7665 | 2.5920 | 3.1334 | 5.9938 | 5.2904 | 4.9889 | H10 | 2.0568 | 2.6143 | 1.0950 | 3.9986 | 2.1547 | 5.0382 | 2.7697 | 2.9779 | 2.4007 | | 1.7621 | 4.5007 | 4.1446 | 2.4843 | 3.0415 | 5.9938 | 4.9889 | 5.2904 | 3.7665 | 3.1334 | 2.5920 | H11 | 2.0568 | 2.6143 | 1.0950 | 3.9986 | 2.1547 | 5.0382 | 2.7697 | 2.4007 | 2.9779 | 1.7621 | | 4.1446 | 4.5007 | 3.0415 | 2.4843 | 5.9938 | 5.2904 | 4.9889 | 3.7665 | 2.5920 | 3.1334 | H12 | 2.5842 | 2.1251 | 3.9087 | 1.0886 | 4.7803 | 2.1581 | 6.2034 | 2.4843 | 3.0415 | 4.5007 | 4.1446 | | 1.7473 | 4.9062 | 4.5845 | 2.5006 | 3.0686 | 2.5184 | 6.9114 | 6.4064 | 6.6419 | H13 | 2.5842 | 2.1251 | 3.9087 | 1.0886 | 4.7803 | 2.1581 | 6.2034 | 3.0415 | 2.4843 | 4.1446 | 4.5007 | 1.7473 | | 4.5845 | 4.9062 | 2.5006 | 2.5184 | 3.0686 | 6.9114 | 6.6419 | 6.4064 | H14 | 2.5842 | 3.9087 | 2.1251 | 4.7803 | 1.0886 | 6.2034 | 2.1581 | 4.5007 | 4.1446 | 2.4843 | 3.0415 | 4.9062 | 4.5845 | | 1.7473 | 6.9114 | 6.4064 | 6.6419 | 2.5006 | 3.0686 | 2.5184 | H15 | 2.5842 | 3.9087 | 2.1251 | 4.7803 | 1.0886 | 6.2034 | 2.1581 | 4.1446 | 4.5007 | 3.0415 | 2.4843 | 4.5845 | 4.9062 | 1.7473 | | 6.9114 | 6.6419 | 6.4064 | 2.5006 | 2.5184 | 3.0686 | H16 | 4.5290 | 3.4674 | 5.7689 | 2.1654 | 6.8909 | 1.0873 | 8.2458 | 3.7665 | 3.7665 | 5.9938 | 5.9938 | 2.5006 | 2.5006 | 6.9114 | 6.9114 | | 1.7569 | 1.7569 | 9.0553 | 8.4631 | 8.4631 | H17 | 4.0712 | 2.7909 | 5.0554 | 2.1696 | 6.3622 | 1.0889 | 7.5517 | 3.1334 | 2.5920 | 4.9889 | 5.2904 | 3.0686 | 2.5184 | 6.4064 | 6.6419 | 1.7569 | | 1.7591 | 8.4631 | 7.7705 | 7.5688 | H18 | 4.0712 | 2.7909 | 5.0554 | 2.1696 | 6.3622 | 1.0889 | 7.5517 | 2.5920 | 3.1334 | 5.2904 | 4.9889 | 2.5184 | 3.0686 | 6.6419 | 6.4064 | 1.7569 | 1.7591 | | 8.4631 | 7.5688 | 7.7705 | H19 | 4.5290 | 5.7689 | 3.4674 | 6.8909 | 2.1654 | 8.2458 | 1.0873 | 5.9938 | 5.9938 | 3.7665 | 3.7665 | 6.9114 | 6.9114 | 2.5006 | 2.5006 | 9.0553 | 8.4631 | 8.4631 | | 1.7569 | 1.7569 | H20 | 4.0712 | 5.0554 | 2.7909 | 6.3622 | 2.1696 | 7.5517 | 1.0889 | 4.9889 | 5.2904 | 3.1334 | 2.5920 | 6.4064 | 6.6419 | 3.0686 | 2.5184 | 8.4631 | 7.7705 | 7.5688 | 1.7569 | | 1.7591 | H21 | 4.0712 | 5.0554 | 2.7909 | 6.3622 | 2.1696 | 7.5517 | 1.0889 | 5.2904 | 4.9889 | 2.5920 | 3.1334 | 6.6419 | 6.4064 | 2.5184 | 3.0686 | 8.4631 | 7.5688 | 7.7705 | 1.7569 | 1.7591 | |
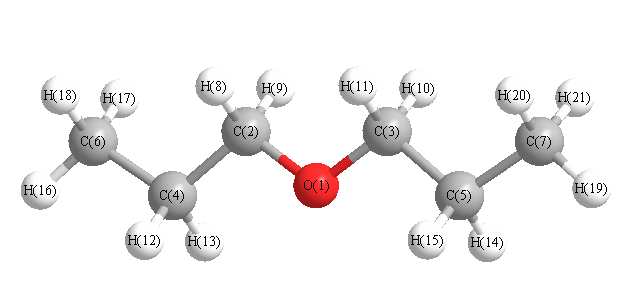 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
108.556 |
|
O1 |
C2 |
H8 |
110.214 |
| O1 |
C2 |
H9 |
110.214 |
|
O1 |
C3 |
C5 |
108.556 |
| O1 |
C3 |
H10 |
110.214 |
|
O1 |
C3 |
H11 |
110.214 |
| C2 |
O1 |
C3 |
113.525 |
|
C2 |
C4 |
C6 |
112.354 |
| C2 |
C4 |
H12 |
108.404 |
|
C2 |
C4 |
H13 |
108.404 |
| C3 |
C5 |
C7 |
112.354 |
|
C3 |
C5 |
H14 |
108.404 |
| C3 |
C5 |
H15 |
108.404 |
|
C4 |
C2 |
H8 |
110.361 |
| C4 |
C2 |
H9 |
110.361 |
|
C4 |
C6 |
H16 |
111.030 |
| C4 |
C6 |
H17 |
111.274 |
|
C4 |
C6 |
H18 |
111.274 |
| C5 |
C3 |
H10 |
110.361 |
|
C5 |
C3 |
H11 |
110.361 |
| C5 |
C7 |
H19 |
111.030 |
|
C5 |
C7 |
H20 |
111.274 |
| C5 |
C7 |
H21 |
111.274 |
|
C6 |
C4 |
H12 |
110.372 |
| C6 |
C4 |
H13 |
110.372 |
|
C7 |
C5 |
H14 |
110.372 |
| C7 |
C5 |
H15 |
110.372 |
|
H8 |
C2 |
H9 |
107.138 |
| H10 |
C3 |
H11 |
107.138 |
|
H12 |
C4 |
H13 |
106.745 |
| H14 |
C5 |
H15 |
106.745 |
|
H16 |
C6 |
H17 |
107.667 |
| H16 |
C6 |
H18 |
107.667 |
|
H17 |
C6 |
H18 |
107.754 |
| H19 |
C7 |
H20 |
107.667 |
|
H19 |
C7 |
H21 |
107.667 |
| H20 |
C7 |
H21 |
107.754 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -311.036184 |
| Energy at 298.15K | -311.052246 |
| HF Energy | -310.214376 |
| Nuclear repulsion energy | 327.032877 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3275 |
3025 |
8.99 |
|
|
|
| 2 |
A |
3245 |
2997 |
60.29 |
|
|
|
| 3 |
A |
3219 |
2973 |
9.84 |
|
|
|
| 4 |
A |
3179 |
2936 |
2.17 |
|
|
|
| 5 |
A |
3174 |
2932 |
13.76 |
|
|
|
| 6 |
A |
3150 |
2910 |
2.25 |
|
|
|
| 7 |
A |
3121 |
2883 |
85.98 |
|
|
|
| 8 |
A |
1644 |
1518 |
0.70 |
|
|
|
| 9 |
A |
1614 |
1491 |
4.95 |
|
|
|
| 10 |
A |
1600 |
1478 |
5.79 |
|
|
|
| 11 |
A |
1589 |
1468 |
0.57 |
|
|
|
| 12 |
A |
1556 |
1437 |
2.18 |
|
|
|
| 13 |
A |
1524 |
1407 |
1.60 |
|
|
|
| 14 |
A |
1476 |
1364 |
0.20 |
|
|
|
| 15 |
A |
1397 |
1290 |
3.09 |
|
|
|
| 16 |
A |
1351 |
1248 |
0.03 |
|
|
|
| 17 |
A |
1266 |
1169 |
9.95 |
|
|
|
| 18 |
A |
1225 |
1132 |
11.49 |
|
|
|
| 19 |
A |
1159 |
1071 |
5.79 |
|
|
|
| 20 |
A |
1025 |
946 |
8.54 |
|
|
|
| 21 |
A |
980 |
905 |
6.19 |
|
|
|
| 22 |
A |
958 |
885 |
0.64 |
|
|
|
| 23 |
A |
802 |
741 |
0.30 |
|
|
|
| 24 |
A |
508 |
470 |
0.01 |
|
|
|
| 25 |
A |
352 |
326 |
0.45 |
|
|
|
| 26 |
A |
287 |
265 |
0.50 |
|
|
|
| 27 |
A |
172 |
159 |
0.08 |
|
|
|
| 28 |
A |
120 |
110 |
0.03 |
|
|
|
| 29 |
A |
47 |
44 |
0.00 |
|
|
|
| 30 |
B |
3275 |
3025 |
49.21 |
|
|
|
| 31 |
B |
3245 |
2997 |
38.68 |
|
|
|
| 32 |
B |
3219 |
2973 |
34.53 |
|
|
|
| 33 |
B |
3178 |
2935 |
50.42 |
|
|
|
| 34 |
B |
3174 |
2932 |
88.90 |
|
|
|
| 35 |
B |
3151 |
2911 |
82.63 |
|
|
|
| 36 |
B |
3110 |
2872 |
6.56 |
|
|
|
| 37 |
B |
1625 |
1501 |
4.52 |
|
|
|
| 38 |
B |
1613 |
1489 |
11.26 |
|
|
|
| 39 |
B |
1600 |
1478 |
7.02 |
|
|
|
| 40 |
B |
1589 |
1468 |
2.44 |
|
|
|
| 41 |
B |
1524 |
1407 |
10.95 |
|
|
|
| 42 |
B |
1502 |
1388 |
54.06 |
|
|
|
| 43 |
B |
1469 |
1357 |
25.83 |
|
|
|
| 44 |
B |
1389 |
1283 |
2.92 |
|
|
|
| 45 |
B |
1363 |
1259 |
6.49 |
|
|
|
| 46 |
B |
1275 |
1177 |
124.37 |
|
|
|
| 47 |
B |
1261 |
1165 |
60.96 |
|
|
|
| 48 |
B |
1184 |
1094 |
16.47 |
|
|
|
| 49 |
B |
1094 |
1010 |
24.95 |
|
|
|
| 50 |
B |
995 |
919 |
1.37 |
|
|
|
| 51 |
B |
934 |
863 |
0.14 |
|
|
|
| 52 |
B |
836 |
772 |
1.83 |
|
|
|
| 53 |
B |
528 |
488 |
1.45 |
|
|
|
| 54 |
B |
340 |
314 |
0.57 |
|
|
|
| 55 |
B |
241 |
223 |
2.78 |
|
|
|
| 56 |
B |
177 |
164 |
5.59 |
|
|
|
| 57 |
B |
81 |
75 |
1.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 44990.2 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 41557.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.128 |
| C2 |
0.018 |
1.175 |
0.897 |
| C3 |
-0.018 |
-1.175 |
0.897 |
| C4 |
0.000 |
2.367 |
-0.037 |
| C5 |
0.000 |
-2.367 |
-0.037 |
| C6 |
-1.259 |
2.431 |
-0.890 |
| C7 |
1.259 |
-2.431 |
-0.890 |
| H8 |
-0.851 |
1.202 |
1.563 |
| H9 |
0.913 |
1.197 |
1.527 |
| H10 |
0.851 |
-1.202 |
1.563 |
| H11 |
-0.913 |
-1.197 |
1.527 |
| H12 |
0.881 |
2.316 |
-0.675 |
| H13 |
0.094 |
3.273 |
0.563 |
| H14 |
-0.881 |
-2.316 |
-0.675 |
| H15 |
-0.094 |
-3.273 |
0.563 |
| H16 |
-1.241 |
3.298 |
-1.547 |
| H17 |
-1.352 |
1.540 |
-1.502 |
| H18 |
-2.148 |
2.505 |
-0.265 |
| H19 |
1.241 |
-3.298 |
-1.547 |
| H20 |
1.352 |
-1.540 |
-1.502 |
| H21 |
2.148 |
-2.505 |
-0.265 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
H21 |
| O1 | | 1.4041 | 1.4041 | 2.3728 | 2.3728 | 2.9211 | 2.9211 | 2.0555 | 2.0545 | 2.0555 | 2.0545 | 2.6053 | 3.3029 | 2.6053 | 3.3029 | 3.9020 | 2.6186 | 3.3232 | 3.9020 | 2.6186 | 3.3232 |
C2 | 1.4041 | | 2.3504 | 1.5142 | 3.6631 | 2.5302 | 4.2115 | 1.0952 | 1.0942 | 2.6047 | 2.6247 | 2.1255 | 2.1255 | 3.9332 | 4.4618 | 3.4739 | 2.7868 | 2.7951 | 5.2419 | 3.8605 | 4.4078 | C3 | 1.4041 | 2.3504 | | 3.6631 | 1.5142 | 4.2115 | 2.5302 | 2.6047 | 2.6247 | 1.0952 | 1.0942 | 3.9332 | 4.4618 | 2.1255 | 2.1255 | 5.2419 | 3.8605 | 4.4078 | 3.4739 | 2.7868 | 2.7951 | C4 | 2.3728 | 1.5142 | 3.6631 | | 4.7341 | 1.5221 | 5.0335 | 2.1542 | 2.1557 | 4.0021 | 3.9973 | 1.0891 | 1.0905 | 4.8082 | 5.6725 | 2.1656 | 2.1587 | 2.1647 | 5.9929 | 4.3860 | 5.3293 | C5 | 2.3728 | 3.6631 | 1.5142 | 4.7341 | | 5.0335 | 1.5221 | 4.0021 | 3.9973 | 2.1542 | 2.1557 | 4.8082 | 5.6725 | 1.0891 | 1.0905 | 5.9929 | 4.3860 | 5.3293 | 2.1656 | 2.1587 | 2.1647 | C6 | 2.9211 | 2.5302 | 4.2115 | 1.5221 | 5.0335 | | 5.4757 | 2.7739 | 3.4757 | 4.8645 | 4.3729 | 2.1536 | 2.1564 | 4.7676 | 6.0004 | 1.0880 | 1.0857 | 1.0892 | 6.2855 | 4.7915 | 6.0302 | C7 | 2.9211 | 4.2115 | 2.5302 | 5.0335 | 1.5221 | 5.4757 | | 4.8645 | 4.3729 | 2.7739 | 3.4757 | 4.7676 | 6.0004 | 2.1536 | 2.1564 | 6.2855 | 4.7915 | 6.0302 | 1.0880 | 1.0857 | 1.0892 | H8 | 2.0555 | 1.0952 | 2.6047 | 2.1542 | 4.0021 | 2.7739 | 4.8645 | | 1.7639 | 2.9444 | 2.3994 | 3.0413 | 2.4864 | 4.1696 | 4.6467 | 3.7710 | 3.1242 | 2.5931 | 5.8563 | 4.6647 | 5.1061 | H9 | 2.0545 | 1.0942 | 2.6247 | 2.1557 | 3.9973 | 3.4757 | 4.3729 | 1.7639 | | 2.3994 | 3.0103 | 2.4704 | 2.4308 | 4.5175 | 4.6819 | 4.3018 | 3.7975 | 3.7805 | 5.4553 | 4.1055 | 4.2941 | H10 | 2.0555 | 2.6047 | 1.0952 | 4.0021 | 2.1542 | 4.8645 | 2.7739 | 2.9444 | 2.3994 | | 1.7639 | 4.1696 | 4.6467 | 3.0413 | 2.4864 | 5.8563 | 4.6647 | 5.1061 | 3.7710 | 3.1242 | 2.5931 | H11 | 2.0545 | 2.6247 | 1.0942 | 3.9973 | 2.1557 | 4.3729 | 3.4757 | 2.3994 | 3.0103 | 1.7639 | | 4.5175 | 4.6819 | 2.4704 | 2.4308 | 5.4553 | 4.1055 | 4.2941 | 4.3018 | 3.7975 | 3.7805 | H12 | 2.6053 | 2.1255 | 3.9332 | 1.0891 | 4.8082 | 2.1536 | 4.7676 | 3.0413 | 2.4704 | 4.1696 | 4.5175 | | 1.7513 | 4.9566 | 5.8072 | 2.4957 | 2.5046 | 3.0625 | 5.6932 | 3.9717 | 5.0018 | H13 | 3.3029 | 2.1255 | 4.4618 | 1.0905 | 5.6725 | 2.1564 | 6.0004 | 2.4864 | 2.4308 | 4.6467 | 4.6819 | 1.7513 | | 5.8072 | 6.5484 | 2.4977 | 3.0596 | 2.5108 | 6.9961 | 5.3858 | 6.1876 | H14 | 2.6053 | 3.9332 | 2.1255 | 4.8082 | 1.0891 | 4.7676 | 2.1536 | 4.1696 | 4.5175 | 3.0413 | 2.4704 | 4.9566 | 5.8072 | | 1.7513 | 5.6932 | 3.9717 | 5.0018 | 2.4957 | 2.5046 | 3.0625 | H15 | 3.3029 | 4.4618 | 2.1255 | 5.6725 | 1.0905 | 6.0004 | 2.1564 | 4.6467 | 4.6819 | 2.4864 | 2.4308 | 5.8072 | 6.5484 | 1.7513 | | 6.9961 | 5.3858 | 6.1876 | 2.4977 | 3.0596 | 2.5108 | H16 | 3.9020 | 3.4739 | 5.2419 | 2.1656 | 5.9929 | 1.0880 | 6.2855 | 3.7710 | 4.3018 | 5.8563 | 5.4553 | 2.4957 | 2.4977 | 5.6932 | 6.9961 | | 1.7625 | 1.7592 | 7.0478 | 5.4890 | 6.8415 | H17 | 2.6186 | 2.7868 | 3.8605 | 2.1587 | 4.3860 | 1.0857 | 4.7915 | 3.1242 | 3.7975 | 4.6647 | 4.1055 | 2.5046 | 3.0596 | 3.9717 | 5.3858 | 1.7625 | | 1.7595 | 5.4890 | 4.0977 | 5.4898 | H18 | 3.3232 | 2.7951 | 4.4078 | 2.1647 | 5.3293 | 1.0892 | 6.0302 | 2.5931 | 3.7805 | 5.1061 | 4.2941 | 3.0625 | 2.5108 | 5.0018 | 6.1876 | 1.7592 | 1.7595 | | 6.8415 | 5.4898 | 6.5996 | H19 | 3.9020 | 5.2419 | 3.4739 | 5.9929 | 2.1656 | 6.2855 | 1.0880 | 5.8563 | 5.4553 | 3.7710 | 4.3018 | 5.6932 | 6.9961 | 2.4957 | 2.4977 | 7.0478 | 5.4890 | 6.8415 | | 1.7625 | 1.7592 | H20 | 2.6186 | 3.8605 | 2.7868 | 4.3860 | 2.1587 | 4.7915 | 1.0857 | 4.6647 | 4.1055 | 3.1242 | 3.7975 | 3.9717 | 5.3858 | 2.5046 | 3.0596 | 5.4890 | 4.0977 | 5.4898 | 1.7625 | | 1.7595 | H21 | 3.3232 | 4.4078 | 2.7951 | 5.3293 | 2.1647 | 6.0302 | 1.0892 | 5.1061 | 4.2941 | 2.5931 | 3.7805 | 5.0018 | 6.1876 | 3.0625 | 2.5108 | 6.8415 | 5.4898 | 6.5996 | 1.7592 | 1.7595 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
108.735 |
|
O1 |
C2 |
H8 |
110.043 |
| O1 |
C2 |
H9 |
110.019 |
|
O1 |
C3 |
C5 |
108.735 |
| O1 |
C3 |
H10 |
110.043 |
|
O1 |
C3 |
H11 |
110.019 |
| C2 |
O1 |
C3 |
113.638 |
|
C2 |
C4 |
C6 |
112.883 |
| C2 |
C4 |
H12 |
108.362 |
|
C2 |
C4 |
H13 |
108.283 |
| C3 |
C5 |
C7 |
112.883 |
|
C3 |
C5 |
H14 |
108.362 |
| C3 |
C5 |
H15 |
108.283 |
|
C4 |
C2 |
H8 |
110.259 |
| C4 |
C2 |
H9 |
110.437 |
|
C4 |
C6 |
H16 |
111.048 |
| C4 |
C6 |
H17 |
110.633 |
|
C4 |
C6 |
H18 |
110.899 |
| C5 |
C3 |
H10 |
110.259 |
|
C5 |
C3 |
H11 |
110.437 |
| C5 |
C7 |
H19 |
111.048 |
|
C5 |
C7 |
H20 |
110.633 |
| C5 |
C7 |
H21 |
110.899 |
|
C6 |
C4 |
H12 |
110.021 |
| C6 |
C4 |
H13 |
110.162 |
|
C7 |
C5 |
H14 |
110.021 |
| C7 |
C5 |
H15 |
110.162 |
|
H8 |
C2 |
H9 |
107.343 |
| H10 |
C3 |
H11 |
107.343 |
|
H12 |
C4 |
H13 |
106.929 |
| H14 |
C5 |
H15 |
106.929 |
|
H16 |
C6 |
H17 |
108.350 |
| H16 |
C6 |
H18 |
107.800 |
|
H17 |
C6 |
H18 |
107.991 |
| H19 |
C7 |
H20 |
108.350 |
|
H19 |
C7 |
H21 |
107.800 |
| H20 |
C7 |
H21 |
107.991 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
