return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6OS (Vinyl sulfoxide)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-627.913248
Energy at 298.15K-627.919741
Nuclear repulsion energy291.306273
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3392 3133 0.70      
2 A' 3327 3073 6.20      
3 A' 3294 3043 2.00      
4 A' 1793 1656 0.33      
5 A' 1499 1384 12.75      
6 A' 1365 1261 0.45      
7 A' 1150 1063 80.28      
8 A' 1076 994 27.59      
9 A' 1067 985 66.67      
10 A' 1042 963 0.90      
11 A' 771 712 72.63      
12 A' 696 643 9.40      
13 A' 527 486 2.02      
14 A' 326 302 3.63      
15 A' 221 204 3.46      
16 A' 95 88 0.86      
17 A" 3392 3133 1.67      
18 A" 3322 3068 5.16      
19 A" 3293 3042 2.95      
20 A" 1783 1647 0.26      
21 A" 1495 1381 5.87      
22 A" 1348 1245 8.51      
23 A" 1070 988 8.25      
24 A" 1057 976 45.37      
25 A" 1039 959 0.95      
26 A" 729 673 16.23      
27 A" 608 561 4.09      
28 A" 514 475 13.62      
29 A" 270 249 8.59      
30 A" 164 152 0.50      

Unscaled Zero Point Vibrational Energy (zpe) 20861.2 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 19269.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.16976 0.07503 0.06642

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.578 -0.502 0.000
O2 1.282 0.817 0.000
C3 -0.607 -0.477 1.329
C4 -0.607 -0.477 -1.329
C5 -0.607 0.515 2.199
C6 -0.607 0.515 -2.199
H7 -1.254 -1.341 1.388
H8 -1.254 -1.341 -1.388
H9 -1.289 0.526 3.034
H10 -1.289 0.526 -3.034
H11 0.080 1.339 2.086
H12 0.080 1.339 -2.086

Atom - Atom Distances (Å)
  S1 O2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
S11.49581.78021.78022.69662.69662.44662.44663.70823.70822.82712.8271
O21.49582.64742.64742.91422.91423.60783.60783.98773.98772.46372.4637
C31.78022.64742.65711.31963.66401.08082.92342.09354.52862.08483.9285
C41.78022.64742.65713.66401.31962.92341.08084.52862.09353.92852.0848
C52.69662.91421.31963.66404.39722.12584.09011.07895.27731.07914.4172
C62.69662.91423.66401.31964.39724.09012.12585.27731.07894.41721.0791
H72.44663.60781.08082.92342.12584.09012.77672.48964.80113.07424.5866
H82.44663.60782.92341.08084.09012.12582.77674.80112.48964.58663.0742
H93.70823.98772.09354.52861.07895.27732.48964.80116.06901.85315.3625
H103.70823.98774.52862.09355.27731.07894.80112.48966.06905.36251.8531
H112.82712.46372.08483.92851.07914.41723.07424.58661.85315.36254.1725
H122.82712.46373.92852.08484.41721.07914.58663.07425.36251.85314.1725

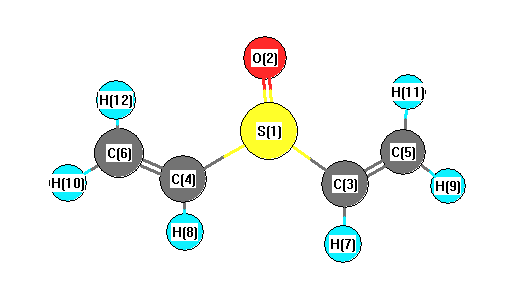
picture of Vinyl sulfoxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 C5 120.172 S1 C3 H7 115.369
S1 C4 C6 120.172 S1 C4 H8 115.369
O2 S1 C3 107.511 O2 S1 C4 107.511
C3 S1 C4 96.540 C3 C5 H9 121.260
C3 C5 H11 120.395 C4 C6 H10 121.260
C4 C6 H12 120.395 C5 C3 H7 124.357
C6 C4 H8 124.357 H9 C5 H11 118.344
H10 C6 H12 118.344
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability