return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Cyclobutane, methylene-)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-194.518817
Energy at 298.15K-194.527467
HF Energy-193.942459
Nuclear repulsion energy168.237792
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3266 3017 6.94      
2 A' 3250 3002 47.86      
3 A' 3222 2976 15.33      
4 A' 3193 2949 38.07      
5 A' 3171 2929 10.05      
6 A' 1869 1726 14.97      
7 A' 1602 1480 0.01      
8 A' 1571 1451 1.90      
9 A' 1523 1407 0.90      
10 A' 1350 1247 0.28      
11 A' 1291 1193 0.04      
12 A' 1161 1073 0.21      
13 A' 1030 951 0.41      
14 A' 963 889 45.23      
15 A' 941 869 0.70      
16 A' 790 730 1.44      
17 A' 697 643 1.51      
18 A' 398 368 5.04      
19 A' 108 100 0.03      
20 A" 3347 3092 13.08      
21 A" 3225 2979 15.41      
22 A" 3167 2925 42.15      
23 A" 1562 1443 2.53      
24 A" 1372 1267 0.12      
25 A" 1324 1223 0.22      
26 A" 1292 1193 1.12      
27 A" 1267 1171 1.84      
28 A" 1102 1018 0.10      
29 A" 990 915 1.35      
30 A" 963 889 0.76      
31 A" 860 794 0.14      
32 A" 683 631 0.13      
33 A" 368 340 0.09      

Unscaled Zero Point Vibrational Energy (zpe) 26457.8 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 24439.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.35022 0.15505 0.11660

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.178 -0.423 1.087
C2 0.178 -0.423 -1.087
C3 0.000 0.619 0.000
C4 -0.031 -1.506 0.000
C5 -0.326 1.899 0.000
H6 -0.525 -0.400 1.919
H7 -0.525 -0.400 -1.919
H8 1.191 -0.431 1.491
H9 1.191 -0.431 -1.491
H10 -0.467 2.447 0.921
H11 -0.467 2.447 -0.921
H12 0.661 -2.345 0.000
H13 -1.048 -1.891 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C12.17491.51641.54912.61261.08893.08741.09092.77042.94623.56192.26122.2006
C22.17491.51641.54912.61263.08741.08892.77041.09093.56192.94622.26122.2006
C31.51641.51642.12551.32052.23542.23542.17832.17832.09972.09973.03742.7206
C41.54911.54912.12553.41762.26922.26922.20712.20714.08264.08261.08771.0881
C52.61262.61261.32053.41763.00123.00123.15503.15501.08131.08134.35753.8582
H61.08893.08742.23542.26923.00123.83771.76923.81753.01794.02212.97872.4857
H73.08741.08892.23542.26923.00123.83773.81751.76924.02213.01792.97872.4857
H81.09092.77042.17832.20713.15501.76923.81752.98183.37054.10532.48363.0612
H92.77041.09092.17832.20713.15503.81751.76922.98184.10533.37052.48363.0612
H102.94623.56192.09974.08261.08133.01794.02213.37054.10531.84235.00914.4731
H113.56192.94622.09974.08261.08134.02213.01794.10533.37051.84235.00914.4731
H122.26122.26123.03741.08774.35752.97872.97872.48362.48365.00915.00911.7688
H132.20062.20062.72061.08813.85822.48572.48573.06123.06124.47314.47311.7688

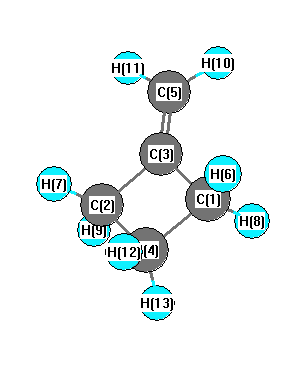
picture of Cyclobutane, methylene- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C2 91.631 C1 C3 C5 134.003
C1 C4 C2 89.174 C1 C4 H12 117.006
C1 C4 H13 111.926 C2 C3 C5 134.003
C2 C4 H12 117.006 C2 C4 H13 111.926
C3 C1 C4 87.784 C3 C1 H6 117.245
C3 C1 H8 112.294 C3 C2 C4 87.784
C3 C2 H7 117.245 C3 C2 H9 112.294
C3 C5 H10 121.584 C3 C5 H11 121.584
C4 C1 H6 117.614 C4 C1 H8 112.284
C4 C2 H7 117.614 C4 C2 H9 112.284
H6 C1 H8 108.509 H7 C2 H9 108.509
H10 C5 H11 116.832 H12 C4 H13 108.765
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability