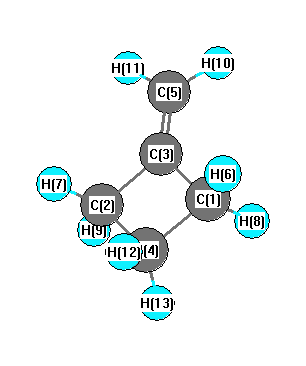
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3164 |
3027 |
9.00 |
|
|
|
| 2 |
A' |
3149 |
3013 |
41.85 |
|
|
|
| 3 |
A' |
3112 |
2977 |
19.62 |
|
|
|
| 4 |
A' |
3091 |
2958 |
41.32 |
|
|
|
| 5 |
A' |
3061 |
2929 |
10.59 |
|
|
|
| 6 |
A' |
1775 |
1699 |
19.94 |
|
|
|
| 7 |
A' |
1524 |
1458 |
0.01 |
|
|
|
| 8 |
A' |
1495 |
1430 |
1.60 |
|
|
|
| 9 |
A' |
1457 |
1394 |
0.65 |
|
|
|
| 10 |
A' |
1277 |
1222 |
0.22 |
|
|
|
| 11 |
A' |
1233 |
1180 |
0.08 |
|
|
|
| 12 |
A' |
1102 |
1055 |
0.06 |
|
|
|
| 13 |
A' |
990 |
947 |
0.46 |
|
|
|
| 14 |
A' |
901 |
862 |
33.54 |
|
|
|
| 15 |
A' |
896 |
857 |
11.41 |
|
|
|
| 16 |
A' |
756 |
724 |
2.30 |
|
|
|
| 17 |
A' |
666 |
637 |
1.65 |
|
|
|
| 18 |
A' |
381 |
364 |
5.51 |
|
|
|
| 19 |
A' |
93 |
89 |
0.02 |
|
|
|
| 20 |
A" |
3244 |
3104 |
13.61 |
|
|
|
| 21 |
A" |
3115 |
2980 |
16.67 |
|
|
|
| 22 |
A" |
3057 |
2925 |
45.27 |
|
|
|
| 23 |
A" |
1483 |
1419 |
2.87 |
|
|
|
| 24 |
A" |
1300 |
1244 |
0.16 |
|
|
|
| 25 |
A" |
1256 |
1201 |
0.12 |
|
|
|
| 26 |
A" |
1232 |
1179 |
1.43 |
|
|
|
| 27 |
A" |
1206 |
1154 |
2.64 |
|
|
|
| 28 |
A" |
1054 |
1008 |
0.12 |
|
|
|
| 29 |
A" |
952 |
911 |
1.00 |
|
|
|
| 30 |
A" |
920 |
880 |
0.87 |
|
|
|
| 31 |
A" |
818 |
782 |
0.14 |
|
|
|
| 32 |
A" |
642 |
615 |
0.14 |
|
|
|
| 33 |
A" |
353 |
338 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25377.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 24278.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.373 |
|
|
-0.119 |
| 2 |
C |
-0.373 |
|
|
-0.119 |
| 3 |
C |
0.203 |
|
|
0.237 |
| 4 |
C |
-0.339 |
|
|
-0.108 |
| 5 |
C |
-0.482 |
|
|
-0.677 |
| 6 |
H |
0.176 |
|
|
0.059 |
| 7 |
H |
0.176 |
|
|
0.059 |
| 8 |
H |
0.179 |
|
|
0.060 |
| 9 |
H |
0.179 |
|
|
0.060 |
| 10 |
H |
0.159 |
|
|
0.222 |
| 11 |
H |
0.159 |
|
|
0.222 |
| 12 |
H |
0.163 |
|
|
0.047 |
| 13 |
H |
0.175 |
|
|
0.059 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.086 |
-0.531 |
0.000 |
0.538 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.110 |
-0.528 |
0.000 |
0.540 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.619 |
-0.543 |
0.000 |
| y |
-0.543 |
-31.318 |
0.000 |
| z |
0.000 |
0.000 |
-30.057 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.932 |
-0.543 |
0.000 |
| y |
-0.543 |
-0.480 |
0.000 |
| z |
0.000 |
0.000 |
1.412 |
|
| Polar |
|---|
| 3z2-r2 | 2.824 |
|---|
| x2-y2 | -0.301 |
|---|
| xy | -0.543 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
115.709 |
| (<r2>)1/2 |
10.757 |