return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Cyclobutane, methylene-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-195.043693
Energy at 298.15K 
HF Energy-194.831354
Nuclear repulsion energy167.483691
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3184 3022 8.48      
2 A' 3170 3009 45.23      
3 A' 3136 2977 18.62      
4 A' 3112 2954 40.66      
5 A' 3086 2929 11.24      
6 A' 1771 1681 14.18      
7 A' 1550 1471 0.10      
8 A' 1524 1446 1.52      
9 A' 1478 1403 0.49      
10 A' 1294 1228 0.52      
11 A' 1247 1184 0.06      
12 A' 1118 1061 0.14      
13 A' 989 938 0.29      
14 A' 906 860 6.10      
15 A' 901 855 35.08      
16 A' 762 723 1.83      
17 A' 672 638 1.34      
18 A' 383 363 4.41      
19 A' 108 102 0.03      
20 A" 3265 3099 13.68      
21 A" 3139 2980 17.19      
22 A" 3082 2925 43.86      
23 A" 1513 1436 1.72      
24 A" 1313 1247 0.18      
25 A" 1273 1209 0.04      
26 A" 1245 1182 0.92      
27 A" 1215 1153 1.96      
28 A" 1065 1011 0.12      
29 A" 946 898 0.97      
30 A" 927 880 0.79      
31 A" 829 787 0.13      
32 A" 658 625 0.13      
33 A" 356 337 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 25606.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24306.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.262 -0.380 1.092
C2 0.262 -0.380 -1.092
C3 -0.115 0.610 0.000
C4 0.262 -1.489 0.000
C5 -0.702 1.804 0.000
H6 -0.431 -0.491 1.931
H7 -0.431 -0.491 -1.931
H8 1.265 -0.197 1.494
H9 1.265 -0.197 -1.494
H10 -0.954 2.314 0.925
H11 -0.954 2.314 -0.925
H12 1.103 -2.186 0.000
H13 -0.669 -2.060 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C12.18331.52141.55652.62531.09363.10261.09572.77902.96013.57802.27162.2095
C22.18331.52141.55652.62533.10261.09362.77901.09573.57802.96012.27162.2095
C31.52141.52142.13331.33052.24512.24512.18792.18792.11212.11213.04992.7275
C41.55651.55652.13333.43192.28112.28112.21492.21494.09854.09851.09211.0922
C52.62532.62531.33053.43193.01173.01173.17903.17901.08571.08574.37943.8647
H61.09363.10262.24512.28113.01173.86121.77543.83233.02524.03682.99202.4992
H73.10261.09362.24512.28113.01173.86123.83231.77544.03683.02522.99202.4992
H81.09572.77902.18792.21493.17901.77543.83232.98723.39874.13262.49203.0727
H92.77901.09572.18792.21493.17903.83231.77542.98724.13263.39872.49203.0727
H102.96013.57802.11214.09851.08573.02524.03683.39874.13261.85035.03314.4796
H113.57802.96012.11214.09851.08574.03683.02524.13263.39871.85035.03314.4796
H122.27162.27163.04991.09214.37942.99202.99202.49202.49205.03315.03311.7770
H132.20952.20952.72751.09223.86472.49922.49923.07273.07274.47964.47961.7770

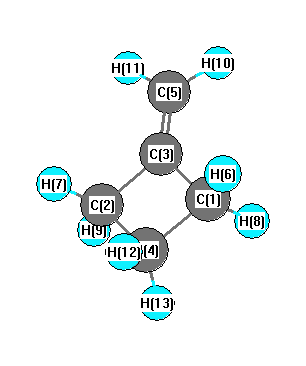
picture of Cyclobutane, methylene- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C2 91.705 C1 C3 C5 133.906
C1 C4 C2 89.076 C1 C4 H12 117.027
C1 C4 H13 111.870 C2 C3 C5 133.906
C2 C4 H12 117.027 C2 C4 H13 111.870
C3 C1 C4 87.748 C3 C1 H6 117.367
C3 C1 H8 112.423 C3 C2 C4 87.748
C3 C2 H7 117.367 C3 C2 H9 112.423
C3 C5 H10 121.558 C3 C5 H11 121.558
C4 C1 H6 117.748 C4 C1 H8 112.092
C4 C2 H7 117.748 C4 C2 H9 112.092
H6 C1 H8 108.377 H7 C2 H9 108.377
H10 C5 H11 116.884 H12 C4 H13 108.887
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.341      
2 C -0.341      
3 C 0.177      
4 C -0.309      
5 C -0.449      
6 H 0.162      
7 H 0.162      
8 H 0.164      
9 H 0.164      
10 H 0.151      
11 H 0.151      
12 H 0.150      
13 H 0.160      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 6.036 -1.628 0.000
y -1.628 8.805 0.000
z 0.000 0.000 7.205


<r2> (average value of r2) Å2
<r2> 115.987
(<r2>)1/2 10.770