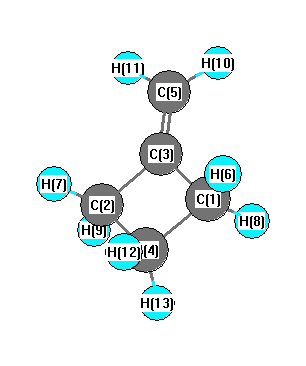
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3179 |
3023 |
7.83 |
|
|
|
| 2 |
A' |
3166 |
3010 |
40.37 |
|
|
|
| 3 |
A' |
3131 |
2978 |
16.49 |
|
|
|
| 4 |
A' |
3106 |
2954 |
40.13 |
|
|
|
| 5 |
A' |
3077 |
2926 |
11.79 |
|
|
|
| 6 |
A' |
1791 |
1703 |
20.81 |
|
|
|
| 7 |
A' |
1527 |
1452 |
0.00 |
|
|
|
| 8 |
A' |
1499 |
1425 |
2.06 |
|
|
|
| 9 |
A' |
1461 |
1389 |
0.81 |
|
|
|
| 10 |
A' |
1281 |
1219 |
0.11 |
|
|
|
| 11 |
A' |
1239 |
1178 |
0.06 |
|
|
|
| 12 |
A' |
1107 |
1053 |
0.06 |
|
|
|
| 13 |
A' |
998 |
949 |
0.40 |
|
|
|
| 14 |
A' |
908 |
864 |
21.75 |
|
|
|
| 15 |
A' |
904 |
860 |
23.63 |
|
|
|
| 16 |
A' |
762 |
725 |
2.29 |
|
|
|
| 17 |
A' |
668 |
635 |
1.77 |
|
|
|
| 18 |
A' |
386 |
367 |
5.43 |
|
|
|
| 19 |
A' |
110 |
105 |
0.03 |
|
|
|
| 20 |
A" |
3261 |
3101 |
12.50 |
|
|
|
| 21 |
A" |
3135 |
2981 |
18.54 |
|
|
|
| 22 |
A" |
3073 |
2922 |
40.49 |
|
|
|
| 23 |
A" |
1486 |
1413 |
3.13 |
|
|
|
| 24 |
A" |
1308 |
1244 |
0.13 |
|
|
|
| 25 |
A" |
1261 |
1199 |
0.15 |
|
|
|
| 26 |
A" |
1236 |
1175 |
1.40 |
|
|
|
| 27 |
A" |
1210 |
1151 |
2.53 |
|
|
|
| 28 |
A" |
1057 |
1005 |
0.15 |
|
|
|
| 29 |
A" |
959 |
912 |
1.12 |
|
|
|
| 30 |
A" |
926 |
880 |
0.74 |
|
|
|
| 31 |
A" |
821 |
781 |
0.17 |
|
|
|
| 32 |
A" |
650 |
618 |
0.16 |
|
|
|
| 33 |
A" |
355 |
338 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25518.3 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 24267.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.376 |
|
|
|
| 2 |
C |
-0.376 |
|
|
|
| 3 |
C |
0.196 |
|
|
|
| 4 |
C |
-0.344 |
|
|
|
| 5 |
C |
-0.482 |
|
|
|
| 6 |
H |
0.178 |
|
|
|
| 7 |
H |
0.178 |
|
|
|
| 8 |
H |
0.181 |
|
|
|
| 9 |
H |
0.181 |
|
|
|
| 10 |
H |
0.161 |
|
|
|
| 11 |
H |
0.161 |
|
|
|
| 12 |
H |
0.166 |
|
|
|
| 13 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.190 |
-0.502 |
0.000 |
0.537 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.436 |
-0.611 |
0.000 |
| y |
-0.611 |
-31.656 |
0.000 |
| z |
0.000 |
0.000 |
-30.144 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.536 |
-0.611 |
0.000 |
| y |
-0.611 |
-0.866 |
0.000 |
| z |
0.000 |
0.000 |
1.402 |
|
| Polar |
|---|
| 3z2-r2 | 2.804 |
|---|
| x2-y2 | 0.220 |
|---|
| xy | -0.611 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.117 |
-1.687 |
0.000 |
| y |
-1.687 |
8.926 |
0.000 |
| z |
0.000 |
0.000 |
7.268 |
<r2> (average value of r
2) Å
2
| <r2> |
115.236 |
| (<r2>)1/2 |
10.735 |