return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6S (Thiophene, 2,3-dihydro-)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-553.019801
Energy at 298.15K-553.026885
HF Energy-552.453345
Nuclear repulsion energy217.596809
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3356 3100 8.26      
2 A 3321 3068 2.13      
3 A 3265 3016 10.78      
4 A 3209 2964 37.34      
5 A 3194 2951 8.26      
6 A 3143 2903 33.39      
7 A 1765 1630 6.98      
8 A 1592 1470 0.73      
9 A 1572 1452 2.27      
10 A 1436 1327 0.45      
11 A 1416 1308 2.58      
12 A 1370 1265 16.46      
13 A 1286 1188 2.43      
14 A 1223 1129 0.42      
15 A 1191 1100 0.83      
16 A 1090 1006 5.66      
17 A 1063 982 2.77      
18 A 990 915 1.13      
19 A 973 899 2.46      
20 A 916 846 1.06      
21 A 843 778 19.65      
22 A 751 694 17.55      
23 A 733 677 30.94      
24 A 693 640 10.25      
25 A 531 491 0.62      
26 A 426 394 2.13      
27 A 173 160 1.35      

Unscaled Zero Point Vibrational Energy (zpe) 20760.1 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 19176.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.23033 0.15738 0.09869

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.055 -1.255 0.210
H2 -0.051 -1.511 1.267
H3 0.142 -2.153 -0.368
C4 -1.376 -0.578 -0.174
H5 -2.200 -0.987 0.411
H6 -1.610 -0.756 -1.228
C7 0.097 1.277 0.081
H8 0.457 2.289 0.177
C9 -1.173 0.897 0.056
H10 -2.001 1.587 0.127
S11 1.269 -0.032 -0.089

Atom - Atom Distances (Å)
  C1 H2 H3 C4 H5 H6 C7 H8 C9 H10 S11
C11.08791.08591.53312.17092.17562.53983.58072.42973.44511.8270
H21.08791.76722.16852.37153.03703.03363.98592.91953.83382.4019
H31.08591.76722.19592.72992.40013.45954.48613.34824.33862.4179
C41.53312.16852.19591.08981.09372.38263.42131.50662.27352.7024
H52.17092.37152.72991.08981.75693.24154.22432.17422.59673.6325
H62.17563.03702.40011.09371.75692.95953.93922.13782.73433.1798
C72.53983.03363.45952.38263.24152.95951.07841.32582.12091.7655
H83.58073.98594.48613.42134.22433.93921.07842.14742.55712.4729
C92.42972.91953.34821.50662.17422.13781.32582.14741.08002.6169
H103.44513.83384.33862.27352.59672.73432.12092.55711.08003.6551
S111.82702.40192.41792.70243.63253.17981.76552.47292.61693.6551

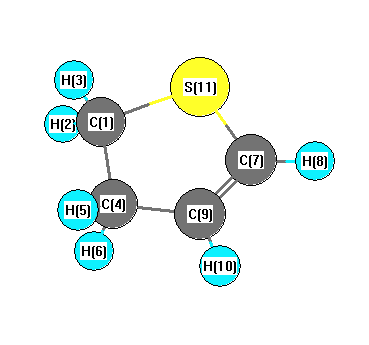
picture of Thiophene, 2,3-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H5 110.581 C1 C4 H6 110.723
C1 C4 C9 106.126 C1 S11 C7 89.960
H2 C1 H3 108.767 H2 C1 C4 110.504
H2 C1 S11 108.296 H3 C1 C4 112.843
H3 C1 S11 109.572 C4 C1 S11 106.753
C4 C9 C7 114.376 C4 C9 H10 122.166
H5 C4 H6 107.142 H5 C4 C9 112.741
H6 C4 C9 109.569 C7 C9 H10 123.337
H8 C7 C9 126.241 H8 C7 S11 118.817
C9 C7 S11 114.933
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability