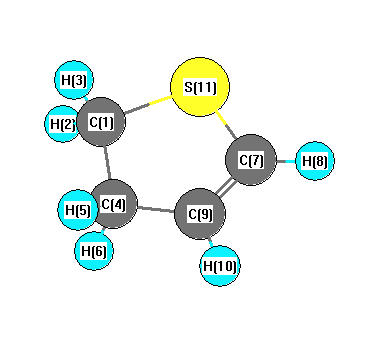
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3254 |
3113 |
7.39 |
|
|
|
| 2 |
A |
3221 |
3081 |
2.83 |
|
|
|
| 3 |
A |
3161 |
3025 |
9.88 |
|
|
|
| 4 |
A |
3098 |
2964 |
39.31 |
|
|
|
| 5 |
A |
3084 |
2950 |
7.17 |
|
|
|
| 6 |
A |
3023 |
2892 |
38.86 |
|
|
|
| 7 |
A |
1671 |
1599 |
12.34 |
|
|
|
| 8 |
A |
1515 |
1449 |
1.13 |
|
|
|
| 9 |
A |
1496 |
1431 |
1.87 |
|
|
|
| 10 |
A |
1359 |
1301 |
0.42 |
|
|
|
| 11 |
A |
1340 |
1282 |
3.49 |
|
|
|
| 12 |
A |
1294 |
1238 |
16.23 |
|
|
|
| 13 |
A |
1220 |
1167 |
2.20 |
|
|
|
| 14 |
A |
1161 |
1111 |
0.49 |
|
|
|
| 15 |
A |
1132 |
1083 |
1.35 |
|
|
|
| 16 |
A |
1035 |
990 |
5.58 |
|
|
|
| 17 |
A |
1010 |
966 |
2.78 |
|
|
|
| 18 |
A |
933 |
893 |
2.84 |
|
|
|
| 19 |
A |
922 |
882 |
1.48 |
|
|
|
| 20 |
A |
866 |
828 |
1.84 |
|
|
|
| 21 |
A |
801 |
767 |
23.52 |
|
|
|
| 22 |
A |
705 |
674 |
14.89 |
|
|
|
| 23 |
A |
685 |
656 |
29.99 |
|
|
|
| 24 |
A |
659 |
630 |
15.35 |
|
|
|
| 25 |
A |
505 |
483 |
0.61 |
|
|
|
| 26 |
A |
408 |
390 |
2.87 |
|
|
|
| 27 |
A |
152 |
145 |
1.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19854.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 18994.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.491 |
|
|
-0.140 |
| 2 |
H |
0.208 |
|
|
0.106 |
| 3 |
H |
0.204 |
|
|
0.114 |
| 4 |
C |
-0.352 |
|
|
0.034 |
| 5 |
H |
0.181 |
|
|
0.037 |
| 6 |
H |
0.186 |
|
|
0.040 |
| 7 |
C |
-0.337 |
|
|
-0.271 |
| 8 |
H |
0.199 |
|
|
0.226 |
| 9 |
C |
-0.117 |
|
|
-0.107 |
| 10 |
H |
0.169 |
|
|
0.100 |
| 11 |
S |
0.149 |
|
|
-0.138 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.692 |
-0.358 |
0.211 |
1.742 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-1.662 |
-0.364 |
0.250 |
1.719 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.918 |
0.574 |
0.054 |
| y |
0.574 |
-32.852 |
0.078 |
| z |
0.054 |
0.078 |
-39.332 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.825 |
0.574 |
0.054 |
| y |
0.574 |
5.773 |
0.078 |
| z |
0.054 |
0.078 |
-3.947 |
|
| Polar |
|---|
| 3z2-r2 | -7.895 |
|---|
| x2-y2 | -5.066 |
|---|
| xy | 0.574 |
|---|
| xz | 0.054 |
|---|
| yz | 0.078 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
127.605 |
| (<r2>)1/2 |
11.296 |