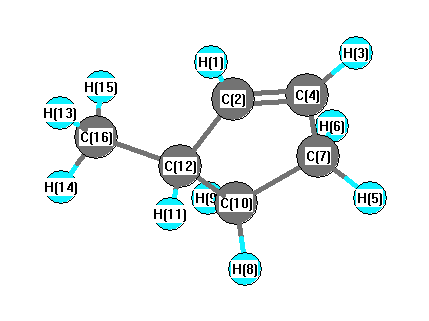
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3220 |
3081 |
28.84 |
|
|
|
| 2 |
A |
3194 |
3056 |
9.41 |
|
|
|
| 3 |
A |
3126 |
2991 |
51.64 |
|
|
|
| 4 |
A |
3123 |
2988 |
24.14 |
|
|
|
| 5 |
A |
3119 |
2984 |
26.90 |
|
|
|
| 6 |
A |
3081 |
2947 |
41.04 |
|
|
|
| 7 |
A |
3058 |
2926 |
25.57 |
|
|
|
| 8 |
A |
3047 |
2915 |
28.18 |
|
|
|
| 9 |
A |
3027 |
2896 |
45.92 |
|
|
|
| 10 |
A |
2994 |
2864 |
33.51 |
|
|
|
| 11 |
A |
1707 |
1633 |
1.02 |
|
|
|
| 12 |
A |
1527 |
1461 |
3.37 |
|
|
|
| 13 |
A |
1523 |
1457 |
4.27 |
|
|
|
| 14 |
A |
1523 |
1457 |
2.92 |
|
|
|
| 15 |
A |
1506 |
1440 |
0.99 |
|
|
|
| 16 |
A |
1431 |
1370 |
2.31 |
|
|
|
| 17 |
A |
1406 |
1345 |
4.78 |
|
|
|
| 18 |
A |
1356 |
1297 |
1.03 |
|
|
|
| 19 |
A |
1344 |
1286 |
0.57 |
|
|
|
| 20 |
A |
1328 |
1271 |
3.55 |
|
|
|
| 21 |
A |
1309 |
1253 |
1.20 |
|
|
|
| 22 |
A |
1243 |
1189 |
0.73 |
|
|
|
| 23 |
A |
1215 |
1162 |
0.29 |
|
|
|
| 24 |
A |
1155 |
1105 |
0.64 |
|
|
|
| 25 |
A |
1130 |
1081 |
4.09 |
|
|
|
| 26 |
A |
1116 |
1068 |
4.09 |
|
|
|
| 27 |
A |
1082 |
1035 |
2.18 |
|
|
|
| 28 |
A |
1015 |
971 |
1.97 |
|
|
|
| 29 |
A |
982 |
940 |
0.80 |
|
|
|
| 30 |
A |
968 |
926 |
1.51 |
|
|
|
| 31 |
A |
960 |
918 |
0.56 |
|
|
|
| 32 |
A |
924 |
884 |
3.02 |
|
|
|
| 33 |
A |
864 |
827 |
1.81 |
|
|
|
| 34 |
A |
834 |
798 |
1.45 |
|
|
|
| 35 |
A |
761 |
728 |
1.91 |
|
|
|
| 36 |
A |
743 |
711 |
29.36 |
|
|
|
| 37 |
A |
586 |
561 |
4.85 |
|
|
|
| 38 |
A |
497 |
475 |
2.61 |
|
|
|
| 39 |
A |
341 |
327 |
0.04 |
|
|
|
| 40 |
A |
285 |
273 |
0.15 |
|
|
|
| 41 |
A |
241 |
230 |
0.04 |
|
|
|
| 42 |
A |
117 |
112 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32002.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30616.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.150 |
|
|
0.163 |
| 2 |
C |
-0.122 |
|
|
-0.375 |
| 3 |
H |
0.153 |
|
|
0.114 |
| 4 |
C |
-0.145 |
|
|
-0.120 |
| 5 |
H |
0.169 |
|
|
0.048 |
| 6 |
H |
0.166 |
|
|
0.050 |
| 7 |
C |
-0.357 |
|
|
-0.057 |
| 8 |
H |
0.160 |
|
|
0.030 |
| 9 |
H |
0.166 |
|
|
0.063 |
| 10 |
C |
-0.333 |
|
|
-0.155 |
| 11 |
H |
0.162 |
|
|
-0.016 |
| 12 |
C |
-0.155 |
|
|
0.415 |
| 13 |
H |
0.164 |
|
|
0.132 |
| 14 |
H |
0.166 |
|
|
0.110 |
| 15 |
H |
0.168 |
|
|
0.141 |
| 16 |
C |
-0.513 |
|
|
-0.543 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.001 |
-0.132 |
0.116 |
0.176 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.010 |
-0.117 |
0.103 |
0.156 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.983 |
-0.226 |
-0.599 |
| y |
-0.226 |
-37.010 |
-0.140 |
| z |
-0.599 |
-0.140 |
-38.459 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.751 |
-0.226 |
-0.599 |
| y |
-0.226 |
0.711 |
-0.140 |
| z |
-0.599 |
-0.140 |
-1.462 |
|
| Polar |
|---|
| 3z2-r2 | -2.924 |
|---|
| x2-y2 | 0.026 |
|---|
| xy | -0.226 |
|---|
| xz | -0.599 |
|---|
| yz | -0.140 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
164.415 |
| (<r2>)1/2 |
12.822 |