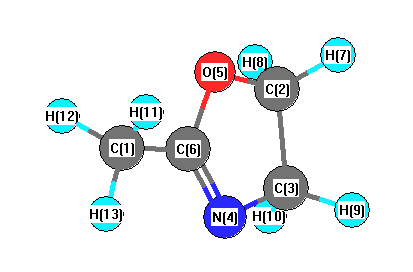
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3196 |
3058 |
4.68 |
|
|
|
| 2 |
A |
3082 |
2949 |
49.99 |
|
|
|
| 3 |
A |
3075 |
2942 |
8.82 |
|
|
|
| 4 |
A |
3064 |
2931 |
45.24 |
|
|
|
| 5 |
A |
1771 |
1694 |
141.37 |
|
|
|
| 6 |
A |
1553 |
1485 |
1.58 |
|
|
|
| 7 |
A |
1526 |
1460 |
0.18 |
|
|
|
| 8 |
A |
1502 |
1436 |
9.63 |
|
|
|
| 9 |
A |
1440 |
1377 |
47.60 |
|
|
|
| 10 |
A |
1396 |
1336 |
13.59 |
|
|
|
| 11 |
A |
1353 |
1294 |
13.86 |
|
|
|
| 12 |
A |
1270 |
1215 |
105.05 |
|
|
|
| 13 |
A |
1101 |
1053 |
2.83 |
|
|
|
| 14 |
A |
1045 |
1000 |
91.43 |
|
|
|
| 15 |
A |
997 |
954 |
1.46 |
|
|
|
| 16 |
A |
962 |
920 |
19.86 |
|
|
|
| 17 |
A |
924 |
884 |
18.02 |
|
|
|
| 18 |
A |
762 |
729 |
5.45 |
|
|
|
| 19 |
A |
669 |
640 |
2.59 |
|
|
|
| 20 |
A |
349 |
334 |
5.94 |
|
|
|
| 21 |
A |
3141 |
3005 |
7.92 |
|
|
|
| 22 |
A |
3132 |
2997 |
45.13 |
|
|
|
| 23 |
A |
3096 |
2962 |
8.44 |
|
|
|
| 24 |
A |
1499 |
1434 |
9.75 |
|
|
|
| 25 |
A |
1260 |
1206 |
0.05 |
|
|
|
| 26 |
A |
1229 |
1176 |
1.31 |
|
|
|
| 27 |
A |
1130 |
1081 |
0.01 |
|
|
|
| 28 |
A |
1072 |
1026 |
4.37 |
|
|
|
| 29 |
A |
839 |
802 |
0.19 |
|
|
|
| 30 |
A |
596 |
571 |
5.53 |
|
|
|
| 31 |
A |
238 |
228 |
10.64 |
|
|
|
| 32 |
A |
140 |
134 |
1.04 |
|
|
|
| 33 |
A |
79 |
75 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24243.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 23193.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.583 |
|
|
0.077 |
| 2 |
C |
-0.105 |
|
|
0.077 |
| 3 |
C |
-0.235 |
|
|
-0.022 |
| 4 |
N |
-0.450 |
|
|
0.040 |
| 5 |
O |
-0.479 |
|
|
0.040 |
| 6 |
C |
0.522 |
|
|
0.163 |
| 7 |
H |
0.179 |
|
|
-0.658 |
| 8 |
H |
0.179 |
|
|
-0.392 |
| 9 |
H |
0.182 |
|
|
0.815 |
| 10 |
H |
0.182 |
|
|
0.198 |
| 11 |
H |
0.200 |
|
|
0.200 |
| 12 |
H |
0.200 |
|
|
0.198 |
| 13 |
H |
0.211 |
|
|
-0.736 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.983 |
-0.899 |
0.000 |
1.332 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.798 |
-1.115 |
-0.001 |
1.371 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.426 |
1.775 |
0.000 |
| y |
1.775 |
-29.197 |
0.000 |
| z |
0.000 |
0.000 |
-34.785 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.435 |
1.775 |
0.000 |
| y |
1.775 |
8.409 |
0.000 |
| z |
0.000 |
0.000 |
0.027 |
|
| Polar |
|---|
| 3z2-r2 | 0.054 |
|---|
| x2-y2 | -11.229 |
|---|
| xy | 1.775 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
147.969 |
| (<r2>)1/2 |
12.164 |