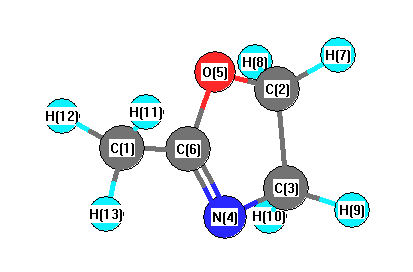
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3122 |
3070 |
4.36 |
|
|
|
| 2 |
A |
3002 |
2952 |
13.99 |
|
|
|
| 3 |
A |
3000 |
2950 |
45.37 |
|
|
|
| 4 |
A |
2983 |
2934 |
45.32 |
|
|
|
| 5 |
A |
1699 |
1671 |
118.76 |
|
|
|
| 6 |
A |
1504 |
1479 |
1.11 |
|
|
|
| 7 |
A |
1479 |
1454 |
0.32 |
|
|
|
| 8 |
A |
1459 |
1435 |
10.34 |
|
|
|
| 9 |
A |
1389 |
1366 |
37.37 |
|
|
|
| 10 |
A |
1344 |
1322 |
16.10 |
|
|
|
| 11 |
A |
1304 |
1283 |
13.29 |
|
|
|
| 12 |
A |
1222 |
1202 |
90.69 |
|
|
|
| 13 |
A |
1059 |
1041 |
2.65 |
|
|
|
| 14 |
A |
986 |
970 |
85.55 |
|
|
|
| 15 |
A |
958 |
942 |
2.27 |
|
|
|
| 16 |
A |
929 |
913 |
18.10 |
|
|
|
| 17 |
A |
888 |
874 |
18.30 |
|
|
|
| 18 |
A |
736 |
724 |
4.71 |
|
|
|
| 19 |
A |
651 |
640 |
2.19 |
|
|
|
| 20 |
A |
339 |
333 |
5.53 |
|
|
|
| 21 |
A |
3063 |
3013 |
7.47 |
|
|
|
| 22 |
A |
3048 |
2997 |
45.26 |
|
|
|
| 23 |
A |
3012 |
2963 |
7.54 |
|
|
|
| 24 |
A |
1456 |
1432 |
9.71 |
|
|
|
| 25 |
A |
1218 |
1198 |
0.08 |
|
|
|
| 26 |
A |
1185 |
1165 |
0.86 |
|
|
|
| 27 |
A |
1093 |
1075 |
0.04 |
|
|
|
| 28 |
A |
1037 |
1019 |
3.88 |
|
|
|
| 29 |
A |
814 |
800 |
0.04 |
|
|
|
| 30 |
A |
572 |
563 |
4.12 |
|
|
|
| 31 |
A |
231 |
227 |
9.84 |
|
|
|
| 32 |
A |
139 |
136 |
1.18 |
|
|
|
| 33 |
A |
71 |
70 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23494.9 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 23107.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.560 |
|
|
|
| 2 |
C |
-0.105 |
|
|
|
| 3 |
C |
-0.225 |
|
|
|
| 4 |
N |
-0.412 |
|
|
|
| 5 |
O |
-0.430 |
|
|
|
| 6 |
C |
0.483 |
|
|
|
| 7 |
H |
0.167 |
|
|
|
| 8 |
H |
0.167 |
|
|
|
| 9 |
H |
0.169 |
|
|
|
| 10 |
H |
0.169 |
|
|
|
| 11 |
H |
0.189 |
|
|
|
| 12 |
H |
0.189 |
|
|
|
| 13 |
H |
0.199 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.971 |
-0.840 |
0.000 |
1.284 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.350 |
1.720 |
0.000 |
| y |
1.720 |
-29.548 |
0.000 |
| z |
0.000 |
0.000 |
-34.990 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.081 |
1.720 |
0.000 |
| y |
1.720 |
8.122 |
0.000 |
| z |
0.000 |
0.000 |
-0.041 |
|
| Polar |
|---|
| 3z2-r2 | -0.081 |
|---|
| x2-y2 | -10.802 |
|---|
| xy | 1.720 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.545 |
0.674 |
0.000 |
| y |
0.674 |
9.249 |
0.000 |
| z |
0.000 |
0.000 |
5.638 |
<r2> (average value of r
2) Å
2
| <r2> |
149.778 |
| (<r2>)1/2 |
12.238 |