return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH3 (Propene)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-117.444959
Energy at 298.15K 
HF Energy-117.071164
Nuclear repulsion energy70.852876
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3331 3077 20.31      
2 A' 3253 3005 5.52      
3 A' 3241 2994 28.89      
4 A' 3217 2971 10.64      
5 A' 3130 2891 22.92      
6 A' 1810 1672 6.99      
7 A' 1579 1458 10.41      
8 A' 1529 1413 1.38      
9 A' 1491 1378 1.30      
10 A' 1386 1281 0.27      
11 A' 1250 1155 0.28      
12 A' 992 916 3.61      
13 A' 971 897 1.02      
14 A' 440 407 0.82      
15 A" 3193 2949 24.72      
16 A" 1564 1444 6.54      
17 A" 1130 1043 1.65      
18 A" 1066 985 13.12      
19 A" 980 905 43.28      
20 A" 610 563 10.41      
21 A" 202 186 0.31      

Unscaled Zero Point Vibrational Energy (zpe) 18181.8 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 16794.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
1.56280 0.31136 0.27271

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.135 -0.503 0.000
C2 0.000 0.477 0.000
C3 1.287 0.147 0.000
H4 1.610 -0.887 0.000
H5 2.066 0.898 0.000
H6 -0.275 1.528 0.000
H7 -0.768 -1.530 0.000
H8 -1.770 -0.367 0.879
H9 -1.770 -0.367 -0.879

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9
C11.49982.50772.77183.49462.20561.09061.09271.0927
C21.49981.32812.11002.10861.08632.14882.14932.1493
C32.50771.32811.08371.08212.08442.65213.22213.2221
H42.77182.11001.08371.84223.06352.46313.53103.5310
H53.49462.10861.08211.84222.42473.73174.13424.1342
H62.20561.08632.08443.06352.42473.09722.56902.5690
H71.09062.14882.65212.46313.73173.09721.76881.7688
H81.09272.14933.22213.53104.13422.56901.76881.7575
H91.09272.14933.22213.53104.13422.56901.76881.7575

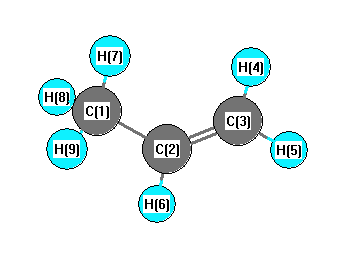
picture of Propene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.826 C1 C2 H6 116.131
C2 C1 H7 111.115 C2 C1 H8 111.026
C2 C1 H9 111.026 C2 C3 H4 121.726
C2 C3 H5 121.729 C3 C2 H6 119.043
H4 C3 H5 116.545 H7 C1 H8 108.224
H7 C1 H9 108.224 H8 C1 H9 107.073
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability