return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2C(CH3)CH3 (1-Propene, 2-methyl-)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-156.677708
Energy at 298.15K-156.685583
HF Energy-156.109344
Nuclear repulsion energy119.268074
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3170 3019 9.49      
2 A1 3148 2998 23.61      
3 A1 3054 2908 31.43      
4 A1 1760 1676 8.66      
5 A1 1552 1478 9.16      
6 A1 1490 1419 0.07      
7 A1 1460 1390 0.13      
8 A1 1111 1058 3.23      
9 A1 835 795 0.03      
10 A1 383 365 0.19      
11 A2 3108 2959 0.00      
12 A2 1523 1450 0.00      
13 A2 1046 996 0.00      
14 A2 712 678 0.00      
15 A2 170 162 0.00      
16 B1 3111 2962 51.31      
17 B1 1538 1465 15.19      
18 B1 1137 1082 0.04      
19 B1 907 863 44.33      
20 B1 431 411 4.84      
21 B1 202 193 0.43      
22 B2 3249 3094 20.83      
23 B2 3146 2996 22.20      
24 B2 3049 2903 19.28      
25 B2 1540 1466 4.15      
26 B2 1466 1396 8.85      
27 B2 1339 1276 2.26      
28 B2 1007 959 0.01      
29 B2 993 945 0.29      
30 B2 440 419 1.27      

Unscaled Zero Point Vibrational Energy (zpe) 24037.2 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 22890.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.30297 0.27829 0.15335

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.113
C2 0.000 0.000 1.458
H3 0.000 0.925 2.032
H4 0.000 -0.925 2.032
C5 0.000 1.291 -0.679
C6 0.000 -1.291 -0.679
H7 0.000 2.161 -0.014
H8 0.883 1.351 -1.330
H9 -0.883 1.351 -1.330
H10 0.000 -2.161 -0.014
H11 -0.883 -1.351 -1.330
H12 0.883 -1.351 -1.330

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 C6 H7 H8 H9 H10 H11 H12
C11.34452.13052.13051.51471.51472.16472.16512.16512.16472.16512.1651
C21.34451.08901.08902.49622.49622.61463.22123.22122.61463.22123.2212
H32.13051.08901.85022.73563.50162.39073.50233.50233.70294.15534.1553
H42.13051.08901.85023.50162.73563.70294.15534.15532.39073.50233.5023
C51.51472.49622.73563.50162.58241.09451.09861.09863.51552.86112.8611
C61.51472.49623.50162.73562.58243.51552.86112.86111.09451.09861.0986
H72.16472.61462.39073.70291.09453.51551.77941.77944.32193.85313.8531
H82.16513.22123.50234.15531.09862.86111.77941.76533.85313.22802.7025
H92.16513.22123.50234.15531.09862.86111.77941.76533.85312.70253.2280
H102.16472.61463.70292.39073.51551.09454.32193.85313.85311.77941.7794
H112.16513.22124.15533.50232.86111.09863.85313.22802.70251.77941.7653
H122.16513.22124.15533.50232.86111.09863.85312.70253.22801.77941.7653

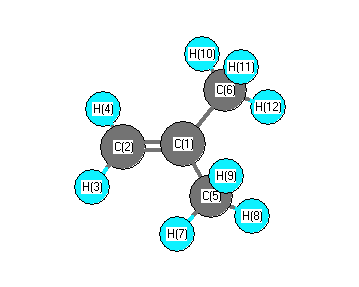
picture of 1-Propene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H3 121.846 C1 C2 H4 121.846
C1 C5 H7 111.102 C1 C5 H8 110.887
C1 C5 H9 110.887 C1 C6 H10 111.102
C1 C6 H11 110.887 C1 C6 H12 110.887
C2 C1 C5 121.518 C2 C1 C6 121.518
H3 C2 H4 116.309 C5 C1 C6 116.964
H7 C5 H8 108.453 H7 C5 H9 108.453
H8 C5 H9 106.915 H10 C6 H11 108.453
H10 C6 H12 108.453 H11 C6 H12 106.915
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability