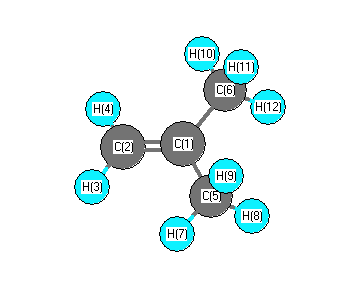
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3179 |
3023 |
8.42 |
|
|
|
| 2 |
A1 |
3159 |
3004 |
19.83 |
|
|
|
| 3 |
A1 |
3054 |
2904 |
36.77 |
|
|
|
| 4 |
A1 |
1764 |
1678 |
14.80 |
|
|
|
| 5 |
A1 |
1529 |
1454 |
11.70 |
|
|
|
| 6 |
A1 |
1468 |
1396 |
0.59 |
|
|
|
| 7 |
A1 |
1434 |
1364 |
0.26 |
|
|
|
| 8 |
A1 |
1097 |
1043 |
4.59 |
|
|
|
| 9 |
A1 |
838 |
797 |
0.09 |
|
|
|
| 10 |
A1 |
376 |
358 |
0.31 |
|
|
|
| 11 |
A2 |
3107 |
2955 |
0.00 |
|
|
|
| 12 |
A2 |
1496 |
1423 |
0.00 |
|
|
|
| 13 |
A2 |
1033 |
983 |
0.00 |
|
|
|
| 14 |
A2 |
710 |
676 |
0.00 |
|
|
|
| 15 |
A2 |
175 |
167 |
0.00 |
|
|
|
| 16 |
B1 |
3109 |
2957 |
45.47 |
|
|
|
| 17 |
B1 |
1515 |
1441 |
18.48 |
|
|
|
| 18 |
B1 |
1119 |
1064 |
1.23 |
|
|
|
| 19 |
B1 |
923 |
878 |
47.86 |
|
|
|
| 20 |
B1 |
437 |
416 |
6.33 |
|
|
|
| 21 |
B1 |
215 |
204 |
0.68 |
|
|
|
| 22 |
B2 |
3261 |
3101 |
19.09 |
|
|
|
| 23 |
B2 |
3157 |
3003 |
22.07 |
|
|
|
| 24 |
B2 |
3049 |
2899 |
19.20 |
|
|
|
| 25 |
B2 |
1511 |
1437 |
4.69 |
|
|
|
| 26 |
B2 |
1439 |
1368 |
10.72 |
|
|
|
| 27 |
B2 |
1322 |
1258 |
3.08 |
|
|
|
| 28 |
B2 |
998 |
950 |
0.03 |
|
|
|
| 29 |
B2 |
975 |
927 |
0.25 |
|
|
|
| 30 |
B2 |
436 |
415 |
1.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23942.8 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 22769.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.200 |
|
|
|
| 2 |
C |
-0.435 |
|
|
|
| 3 |
H |
0.156 |
|
|
|
| 4 |
H |
0.156 |
|
|
|
| 5 |
C |
-0.571 |
|
|
|
| 6 |
C |
-0.571 |
|
|
|
| 7 |
H |
0.178 |
|
|
|
| 8 |
H |
0.177 |
|
|
|
| 9 |
H |
0.177 |
|
|
|
| 10 |
H |
0.178 |
|
|
|
| 11 |
H |
0.177 |
|
|
|
| 12 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.519 |
0.519 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.664 |
0.000 |
0.000 |
| y |
0.000 |
-25.222 |
0.000 |
| z |
0.000 |
0.000 |
-25.628 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.238 |
0.000 |
0.000 |
| y |
0.000 |
1.424 |
0.000 |
| z |
0.000 |
0.000 |
0.815 |
|
| Polar |
|---|
| 3z2-r2 | 1.629 |
|---|
| x2-y2 | -2.441 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.245 |
0.000 |
0.000 |
| y |
0.000 |
6.397 |
0.000 |
| z |
0.000 |
0.000 |
8.213 |
<r2> (average value of r
2) Å
2
| <r2> |
90.487 |
| (<r2>)1/2 |
9.512 |