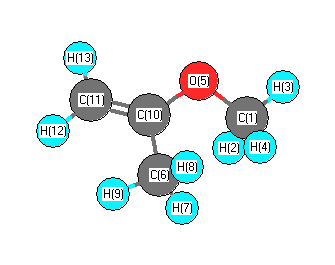
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3214 |
3161 |
8.27 |
|
|
|
| 2 |
A |
3124 |
3073 |
0.51 |
|
|
|
| 3 |
A |
3090 |
3039 |
10.14 |
|
|
|
| 4 |
A |
3082 |
3031 |
26.70 |
|
|
|
| 5 |
A |
3031 |
2981 |
18.25 |
|
|
|
| 6 |
A |
3008 |
2958 |
47.36 |
|
|
|
| 7 |
A |
2977 |
2928 |
23.03 |
|
|
|
| 8 |
A |
2946 |
2898 |
60.68 |
|
|
|
| 9 |
A |
1688 |
1660 |
99.25 |
|
|
|
| 10 |
A |
1488 |
1463 |
8.16 |
|
|
|
| 11 |
A |
1480 |
1455 |
11.35 |
|
|
|
| 12 |
A |
1467 |
1443 |
14.56 |
|
|
|
| 13 |
A |
1458 |
1434 |
5.03 |
|
|
|
| 14 |
A |
1453 |
1429 |
3.43 |
|
|
|
| 15 |
A |
1402 |
1379 |
5.71 |
|
|
|
| 16 |
A |
1380 |
1357 |
27.12 |
|
|
|
| 17 |
A |
1279 |
1258 |
176.69 |
|
|
|
| 18 |
A |
1177 |
1158 |
1.14 |
|
|
|
| 19 |
A |
1140 |
1121 |
3.72 |
|
|
|
| 20 |
A |
1103 |
1085 |
91.65 |
|
|
|
| 21 |
A |
1044 |
1026 |
2.29 |
|
|
|
| 22 |
A |
996 |
979 |
3.57 |
|
|
|
| 23 |
A |
911 |
896 |
4.54 |
|
|
|
| 24 |
A |
797 |
784 |
19.24 |
|
|
|
| 25 |
A |
792 |
779 |
45.84 |
|
|
|
| 26 |
A |
712 |
700 |
3.92 |
|
|
|
| 27 |
A |
519 |
511 |
6.32 |
|
|
|
| 28 |
A |
465 |
457 |
2.24 |
|
|
|
| 29 |
A |
411 |
404 |
0.67 |
|
|
|
| 30 |
A |
306 |
301 |
0.93 |
|
|
|
| 31 |
A |
240 |
236 |
0.66 |
|
|
|
| 32 |
A |
195 |
192 |
1.68 |
|
|
|
| 33 |
A |
80 |
79 |
5.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24225.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 23825.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.270 |
|
|
|
| 2 |
H |
0.158 |
|
|
|
| 3 |
H |
0.179 |
|
|
|
| 4 |
H |
0.156 |
|
|
|
| 5 |
O |
-0.431 |
|
|
|
| 6 |
C |
-0.578 |
|
|
|
| 7 |
H |
0.174 |
|
|
|
| 8 |
H |
0.178 |
|
|
|
| 9 |
H |
0.180 |
|
|
|
| 10 |
C |
0.406 |
|
|
|
| 11 |
C |
-0.441 |
|
|
|
| 12 |
H |
0.141 |
|
|
|
| 13 |
H |
0.149 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.981 |
1.536 |
0.419 |
1.870 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.107 |
-0.228 |
0.328 |
| y |
-0.228 |
-31.230 |
-0.396 |
| z |
0.328 |
-0.396 |
-32.898 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.957 |
-0.228 |
0.328 |
| y |
-0.228 |
-0.728 |
-0.396 |
| z |
0.328 |
-0.396 |
-3.229 |
|
| Polar |
|---|
| 3z2-r2 | -6.459 |
|---|
| x2-y2 | 3.123 |
|---|
| xy | -0.228 |
|---|
| xz | 0.328 |
|---|
| yz | -0.396 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.352 |
0.960 |
-0.225 |
| y |
0.960 |
7.335 |
-0.124 |
| z |
-0.225 |
-0.124 |
4.654 |
<r2> (average value of r
2) Å
2
| <r2> |
129.946 |
| (<r2>)1/2 |
11.399 |