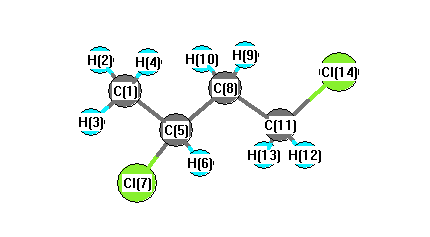
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3185 |
3047 |
5.70 |
|
|
|
| 2 |
A |
3165 |
3028 |
14.36 |
|
|
|
| 3 |
A |
3144 |
3008 |
19.35 |
|
|
|
| 4 |
A |
3124 |
2989 |
12.56 |
|
|
|
| 5 |
A |
3111 |
2976 |
10.42 |
|
|
|
| 6 |
A |
3100 |
2966 |
4.59 |
|
|
|
| 7 |
A |
3068 |
2935 |
21.60 |
|
|
|
| 8 |
A |
3063 |
2930 |
2.07 |
|
|
|
| 9 |
A |
1521 |
1455 |
4.14 |
|
|
|
| 10 |
A |
1515 |
1449 |
4.71 |
|
|
|
| 11 |
A |
1512 |
1446 |
7.86 |
|
|
|
| 12 |
A |
1499 |
1434 |
1.27 |
|
|
|
| 13 |
A |
1435 |
1373 |
6.82 |
|
|
|
| 14 |
A |
1412 |
1351 |
12.54 |
|
|
|
| 15 |
A |
1361 |
1302 |
7.35 |
|
|
|
| 16 |
A |
1340 |
1282 |
13.38 |
|
|
|
| 17 |
A |
1302 |
1246 |
19.28 |
|
|
|
| 18 |
A |
1285 |
1229 |
18.59 |
|
|
|
| 19 |
A |
1196 |
1145 |
8.50 |
|
|
|
| 20 |
A |
1141 |
1092 |
9.42 |
|
|
|
| 21 |
A |
1129 |
1080 |
2.29 |
|
|
|
| 22 |
A |
1080 |
1034 |
2.76 |
|
|
|
| 23 |
A |
1042 |
997 |
17.10 |
|
|
|
| 24 |
A |
971 |
929 |
2.23 |
|
|
|
| 25 |
A |
925 |
885 |
6.79 |
|
|
|
| 26 |
A |
790 |
755 |
11.27 |
|
|
|
| 27 |
A |
765 |
732 |
35.11 |
|
|
|
| 28 |
A |
631 |
603 |
31.42 |
|
|
|
| 29 |
A |
440 |
421 |
3.06 |
|
|
|
| 30 |
A |
411 |
393 |
5.76 |
|
|
|
| 31 |
A |
340 |
325 |
3.87 |
|
|
|
| 32 |
A |
257 |
246 |
0.43 |
|
|
|
| 33 |
A |
243 |
233 |
0.22 |
|
|
|
| 34 |
A |
152 |
145 |
2.34 |
|
|
|
| 35 |
A |
118 |
113 |
1.28 |
|
|
|
| 36 |
A |
78 |
75 |
3.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25423.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 24322.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.500 |
|
|
-0.441 |
| 2 |
H |
0.194 |
|
|
0.137 |
| 3 |
H |
0.198 |
|
|
0.143 |
| 4 |
H |
0.182 |
|
|
0.135 |
| 5 |
C |
-0.259 |
|
|
0.090 |
| 6 |
H |
0.215 |
|
|
0.099 |
| 7 |
Cl |
-0.082 |
|
|
-0.202 |
| 8 |
C |
-0.301 |
|
|
0.019 |
| 9 |
H |
0.195 |
|
|
0.063 |
| 10 |
H |
0.205 |
|
|
0.072 |
| 11 |
C |
-0.431 |
|
|
-0.243 |
| 12 |
H |
0.215 |
|
|
0.142 |
| 13 |
H |
0.241 |
|
|
0.151 |
| 14 |
Cl |
-0.072 |
|
|
-0.165 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.403 |
1.764 |
1.051 |
2.486 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-1.429 |
1.820 |
1.122 |
2.572 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-59.077 |
-3.874 |
0.090 |
| y |
-3.874 |
-50.519 |
0.043 |
| z |
0.090 |
0.043 |
-49.751 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.942 |
-3.874 |
0.090 |
| y |
-3.874 |
3.896 |
0.043 |
| z |
0.090 |
0.043 |
5.047 |
|
| Polar |
|---|
| 3z2-r2 | 10.094 |
|---|
| x2-y2 | -8.559 |
|---|
| xy | -3.874 |
|---|
| xz | 0.090 |
|---|
| yz | 0.043 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
368.424 |
| (<r2>)1/2 |
19.194 |