return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8Cl2 (Butane, 1,3-dichloro-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-1077.115997
Energy at 298.15K-1077.125134
HF Energy-1076.857211
Nuclear repulsion energy348.995318
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3214 3051 5.56      
2 A 3186 3024 15.45      
3 A 3166 3005 20.89      
4 A 3148 2988 15.26      
5 A 3138 2979 8.14      
6 A 3127 2968 3.68      
7 A 3092 2935 21.18      
8 A 3086 2929 1.50      
9 A 1546 1468 3.49      
10 A 1542 1463 4.02      
11 A 1537 1459 6.86      
12 A 1526 1449 0.89      
13 A 1460 1385 4.80      
14 A 1434 1361 11.79      
15 A 1382 1312 7.76      
16 A 1361 1292 15.77      
17 A 1324 1256 19.81      
18 A 1305 1239 19.94      
19 A 1212 1151 7.48      
20 A 1155 1096 8.39      
21 A 1141 1083 1.21      
22 A 1093 1038 3.55      
23 A 1052 998 13.54      
24 A 986 936 1.73      
25 A 937 889 6.69      
26 A 801 761 8.73      
27 A 768 729 35.30      
28 A 637 604 30.91      
29 A 447 424 2.83      
30 A 415 394 5.72      
31 A 344 327 3.58      
32 A 260 247 0.39      
33 A 246 233 0.24      
34 A 154 147 2.30      
35 A 119 113 1.23      
36 A 81 77 3.68      

Unscaled Zero Point Vibrational Energy (zpe) 25709.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24403.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.14763 0.03142 0.02701

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.172 1.527 -0.102
H2 -2.209 1.545 -1.193
H3 -3.185 1.374 0.276
H4 -1.809 2.495 0.253
C5 -1.230 0.417 0.378
H6 -1.228 0.407 1.471
Cl7 -1.888 -1.175 -0.137
C8 0.211 0.639 -0.125
H9 0.516 1.640 0.193
H10 0.220 0.617 -1.217
C11 1.241 -0.361 0.421
H12 1.257 -0.334 1.512
H13 1.007 -1.374 0.104
Cl14 2.895 0.016 -0.147

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 Cl7 C8 H9 H10 C11 H12 H13 Cl14
C11.09211.09201.09341.53302.14912.71642.54332.70642.79213.93484.22224.30835.2878
H21.09211.77201.77642.16773.05842.93492.79643.05872.60064.25884.78154.53245.4298
H31.09201.77201.77482.18002.48922.89003.49833.71163.79504.75634.91775.01596.2447
H41.09341.77641.77482.16102.48583.69102.76932.47773.13204.18144.35774.78755.3326
C51.53302.16772.18002.16101.09321.79721.54202.13962.16482.59032.83472.87824.1773
H62.14913.05842.48922.48581.09322.34972.16082.48873.06052.79012.59343.16774.4462
Cl72.71642.93492.89003.69101.79722.34972.77393.71612.96973.28043.64912.91154.9286
C82.54332.79643.49832.76931.54202.16082.77391.09371.09321.53522.17242.17642.7554
H92.70643.05873.71162.47772.13962.48873.71611.09371.76772.14042.48753.05522.9011
H102.79212.60063.79503.13202.16483.06052.96971.09321.76772.16353.07072.51512.9429
C113.93484.25884.75634.18142.59032.79013.28041.53522.14042.16351.09161.08711.7891
H124.22224.78154.91774.35772.83472.59343.64912.17242.48753.07071.09161.76852.3577
H134.30834.53245.01594.78752.87823.16772.91152.17643.05522.51511.08711.76852.3576
Cl145.28785.42986.24475.33264.17734.44624.92862.75542.90112.94291.78912.35772.3576

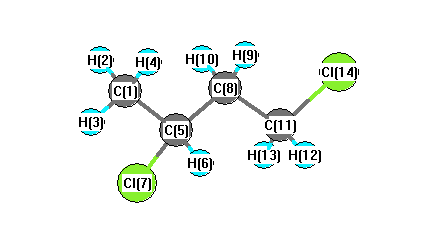
picture of Butane, 1,3-dichloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 108.683 C1 C5 Cl7 109.056
C1 C5 C8 111.605 H2 C1 H3 108.454
H2 C1 H4 108.742 H2 C1 C5 110.201
H3 C1 H4 108.606 H3 C1 C5 111.182
H4 C1 C5 109.601 C5 C8 H9 107.318
C5 C8 H10 109.286 C5 C8 C11 114.651
H6 C5 Cl7 106.203 H6 C5 C8 108.979
Cl7 C5 C8 112.121 C8 C11 H12 110.448
C8 C11 H13 111.033 C8 C11 Cl14 111.733
H9 C8 H10 107.867 H9 C8 C11 107.829
H10 C8 C11 109.654 H12 C11 H13 108.525
H12 C11 Cl14 107.368 H13 C11 Cl14 107.584
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.454      
2 H 0.179      
3 H 0.184      
4 H 0.167      
5 C -0.223      
6 H 0.200      
7 Cl -0.105      
8 C -0.275      
9 H 0.181      
10 H 0.191      
11 C -0.380      
12 H 0.200      
13 H 0.229      
14 Cl -0.093      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.989 0.476 -0.253
y 0.476 9.038 0.210
z -0.253 0.210 7.159


<r2> (average value of r2) Å2
<r2> 369.172
(<r2>)1/2 19.214