return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10S2 (1,4-Butanedithiol)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-954.282309
Energy at 298.15K-954.292990
HF Energy-954.028687
Nuclear repulsion energy334.445747
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3102 2944 0.00      
2 Ag 3069 2913 0.00      
3 Ag 2735 2596 0.00      
4 Ag 1543 1464 0.00      
5 Ag 1541 1462 0.00      
6 Ag 1441 1368 0.00      
7 Ag 1323 1256 0.00      
8 Ag 1130 1073 0.00      
9 Ag 1078 1023 0.00      
10 Ag 874 830 0.00      
11 Ag 779 740 0.00      
12 Ag 340 323 0.00      
13 Ag 208 198 0.00      
14 Au 3161 3000 47.32      
15 Au 3122 2963 14.80      
16 Au 1355 1286 1.44      
17 Au 1140 1082 3.73      
18 Au 922 875 2.17      
19 Au 761 723 3.97      
20 Au 190 180 36.96      
21 Au 102 97 12.55      
22 Au 56 53 6.90      
23 Bg 3157 2996 0.00      
24 Bg 3104 2946 0.00      
25 Bg 1366 1297 0.00      
26 Bg 1293 1227 0.00      
27 Bg 1080 1025 0.00      
28 Bg 803 762 0.00      
29 Bg 187 177 0.00      
30 Bg 137 130 0.00      
31 Bu 3104 2946 62.65      
32 Bu 3076 2920 16.66      
33 Bu 2735 2596 48.22      
34 Bu 1560 1480 3.14      
35 Bu 1537 1459 4.71      
36 Bu 1393 1322 50.86      
37 Bu 1270 1205 33.16      
38 Bu 1073 1019 1.07      
39 Bu 907 861 4.32      
40 Bu 743 705 4.51      
41 Bu 395 375 5.54      
42 Bu 102 97 3.90      

Unscaled Zero Point Vibrational Energy (zpe) 29494.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 27996.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.47858 0.01867 0.01821

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 1.445 2.773 0.000
S2 -1.445 -2.773 0.000
C3 1.445 1.282 0.000
C4 -1.445 -1.282 0.000
C5 0.002 0.764 0.000
C6 -0.002 -0.764 0.000
H7 2.477 3.039 0.000
H8 -2.477 -3.039 0.000
H9 -0.509 1.121 0.897
H10 -0.509 1.121 -0.897
H11 0.509 -1.121 0.897
H12 0.509 -1.121 -0.897
H13 -1.960 -0.922 -0.879
H14 -1.960 -0.922 0.879
H15 1.960 0.922 -0.879
H16 1.960 0.922 0.879

Atom - Atom Distances (Å)
  S1 S2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
S16.25391.49124.97952.47383.82141.06517.01162.71192.71194.10384.10385.10105.10102.11262.1126
S26.25394.97951.49123.82142.47387.01161.06514.10384.10382.71192.71192.11262.11265.10105.1010
C31.49124.97953.86361.53382.50572.03775.83542.15662.15662.72992.72994.15024.15021.08001.0800
C44.97951.49123.86362.50571.53385.83542.03772.72992.72992.15662.15661.08001.08004.15024.1502
C52.47383.82141.53382.50571.52803.36194.53941.09211.09212.14812.14812.73172.73172.15222.1522
C63.82142.47382.50571.53381.52804.53943.36192.14812.14811.09211.09212.15222.15222.73172.7317
H71.06517.01162.03775.83543.36194.53947.84103.66073.66074.68834.68836.01226.01222.34992.3499
H87.01161.06515.83542.03774.53943.36197.84104.68834.68833.66073.66072.34992.34996.01226.0122
H92.71194.10382.15662.72991.09212.14813.66074.68831.79382.46213.04623.07092.50553.04792.4772
H102.71194.10382.15662.72991.09212.14813.66074.68831.79383.04622.46212.50553.07092.47723.0479
H114.10382.71192.72992.15662.14811.09214.68833.66072.46213.04621.79383.04792.47723.07092.5055
H124.10382.71192.72992.15662.14811.09214.68833.66073.04622.46211.79382.47723.04792.50553.0709
H135.10102.11264.15021.08002.73172.15226.01222.34993.07092.50553.04792.47721.75764.33194.6749
H145.10102.11264.15021.08002.73172.15226.01222.34992.50553.07092.47723.04791.75764.67494.3319
H152.11265.10101.08004.15022.15222.73172.34996.01223.04792.47723.07092.50554.33194.67491.7576
H162.11265.10101.08004.15022.15222.73172.34996.01222.47723.04792.50553.07094.67494.33191.7576

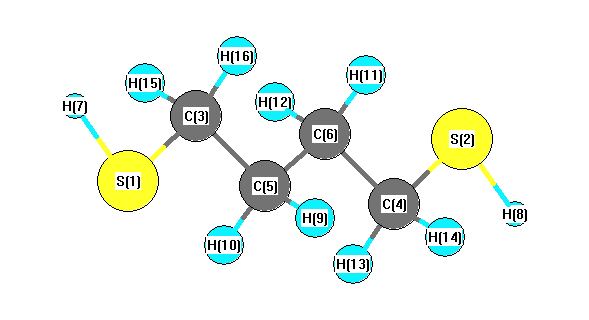
picture of 1,4-Butanedithiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 C5 109.723 S1 C3 H15 109.458
S1 C3 H16 109.458 S2 C4 C6 109.723
S2 C4 H13 109.458 S2 C4 H14 109.458
C3 S1 H7 104.475 C3 C5 C6 109.848
C3 C5 H9 109.275 C3 C5 H10 109.275
C4 S2 H8 104.475 C4 C6 C5 109.848
C4 C6 H11 109.275 C4 C6 H12 109.275
C5 C3 H15 109.634 C5 C3 H16 109.634
C5 C6 H11 109.005 C5 C6 H12 109.005
C6 C4 H13 109.634 C6 C4 H14 109.634
C6 C5 H9 109.005 C6 C5 H10 109.005
H9 C5 H10 110.422 H11 C6 H12 110.422
H13 C4 H14 108.918 H15 C3 H16 108.918
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 S -0.073      
2 S -0.073      
3 C -0.426      
4 C -0.426      
5 C -0.295      
6 C -0.295      
7 H 0.091      
8 H 0.091      
9 H 0.167      
10 H 0.167      
11 H 0.167      
12 H 0.167      
13 H 0.184      
14 H 0.184      
15 H 0.184      
16 H 0.184      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.806 2.306 0.000
y 2.306 13.949 0.000
z 0.000 0.000 8.177


<r2> (average value of r2) Å2
<r2> 535.564
(<r2>)1/2 23.142