return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (Furan, 2,3-dihydro-)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-230.395814
Energy at 298.15K-230.403176
HF Energy-229.788737
Nuclear repulsion energy175.194173
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3387 3129 8.50      
2 A 3361 3105 1.27      
3 A 3253 3005 38.02      
4 A 3208 2964 30.84      
5 A 3178 2936 38.46      
6 A 3158 2917 39.08      
7 A 1803 1665 55.59      
8 A 1621 1497 0.12      
9 A 1596 1474 0.20      
10 A 1498 1383 4.36      
11 A 1416 1308 5.11      
12 A 1390 1284 6.49      
13 A 1320 1220 7.68      
14 A 1286 1188 19.05      
15 A 1242 1148 70.74      
16 A 1169 1080 46.41      
17 A 1145 1058 5.50      
18 A 1077 995 13.98      
19 A 1009 932 31.94      
20 A 991 916 21.64      
21 A 977 903 1.77      
22 A 901 832 4.27      
23 A 877 810 6.84      
24 A 778 718 36.56      
25 A 715 661 1.54      
26 A 472 436 24.08      
27 A 126 116 2.50      

Unscaled Zero Point Vibrational Energy (zpe) 21476.8 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 19838.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.27281 0.26355 0.14203

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 1.130 2.014 0.107
C2 0.618 1.069 0.053
H3 2.231 -0.396 0.031
C4 1.189 -0.123 0.011
O5 0.329 -1.176 -0.088
H6 -1.235 1.224 -1.050
H7 -1.445 1.417 0.683
C8 -0.873 0.886 -0.077
H9 -1.645 -1.115 -0.633
H10 -1.323 -0.902 1.095
C11 -0.991 -0.638 0.091

Atom - Atom Distances (Å)
  H1 C2 H3 C4 O5 H6 H7 C8 H9 H10 C11
H11.07592.65042.13973.29412.74822.70532.30584.24663.93613.3957
C21.07592.17931.32272.26762.16192.18481.50783.21862.95572.3461
H32.65042.17931.07712.05943.97604.15053.36063.99783.74473.2321
C42.13971.32271.07711.36312.96973.12472.29813.07112.84512.2420
O53.29412.26762.05941.36313.02203.23462.38692.04882.05031.4366
H62.74822.16193.97602.96973.02201.75681.09182.41123.02132.1978
H72.70532.18484.15053.12473.23461.75681.08932.86052.35772.1862
C82.30581.50783.36062.29812.38691.09181.08932.21582.18461.5382
H94.24663.21863.99783.07112.04882.41122.86052.21581.77071.0858
H103.93612.95573.74472.84512.05033.02132.35772.18461.77071.0897
C113.39572.34613.23212.24201.43662.19782.18621.53821.08581.0897

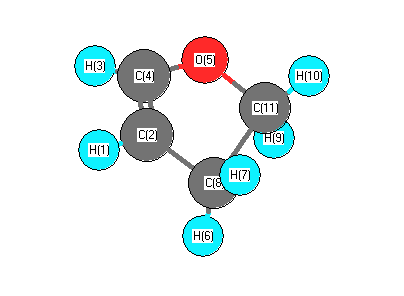
picture of Furan, 2,3-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 125.964 H1 C2 C8 125.538
C2 C4 H3 130.213 C2 C4 O5 115.178
C2 C8 H6 111.531 C2 C8 H7 113.572
C2 C8 C11 100.745 H3 C4 O5 114.605
C4 C2 C8 108.390 C4 O5 C11 106.386
O5 C11 C8 106.667 O5 C11 H9 107.818
O5 C11 H10 107.710 H6 C8 H7 107.309
H6 C8 C11 112.258 H7 C8 C11 111.474
C8 C11 H9 114.107 C8 C11 H10 111.318
H9 C11 H10 108.954
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability