return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (Furan, 2,3-dihydro-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-230.963755
Energy at 298.15K-230.970942
HF Energy-230.735679
Nuclear repulsion energy174.054527
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3300 3132 8.58      
2 A 3276 3109 2.32      
3 A 3167 3006 36.32      
4 A 3117 2958 34.93      
5 A 3091 2934 37.89      
6 A 3065 2909 42.26      
7 A 1701 1614 51.87      
8 A 1567 1488 0.09      
9 A 1545 1467 0.23      
10 A 1437 1364 2.94      
11 A 1354 1285 3.44      
12 A 1335 1267 6.91      
13 A 1270 1206 8.86      
14 A 1234 1171 8.62      
15 A 1187 1127 56.93      
16 A 1111 1055 37.41      
17 A 1096 1041 22.29      
18 A 1025 973 4.85      
19 A 952 904 11.61      
20 A 951 902 38.61      
21 A 908 862 3.51      
22 A 862 818 5.04      
23 A 843 800 5.03      
24 A 724 687 31.13      
25 A 689 654 2.25      
26 A 452 429 22.57      
27 A 111 105 1.69      

Unscaled Zero Point Vibrational Energy (zpe) 20684.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 19633.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.27036 0.25916 0.14002

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.934 1.909 0.541
C2 0.546 1.042 0.137
H3 2.239 -0.211 0.056
C4 1.195 -0.110 -0.001
O5 0.308 -1.163 -0.205
H6 -0.875 0.672 -1.346
H7 -1.545 1.550 0.068
C8 -0.825 0.809 -0.262
H9 -1.904 -1.057 -0.049
H10 -0.965 -0.553 1.385
C11 -0.973 -0.576 0.290

Atom - Atom Distances (Å)
  H1 C2 H3 C4 O5 H6 H7 C8 H9 H10 C11
H11.03192.53682.10683.22262.89202.54942.22494.14733.22233.1430
C21.03192.10801.32932.24402.08752.15321.44733.23182.52692.2252
H32.53682.10801.05092.16853.52774.17383.24514.22963.48503.2410
C42.10681.32931.05091.39152.58913.20392.23413.24042.60412.2363
O53.22262.24402.16851.39152.46323.29612.27452.21982.12551.4930
H62.89202.08753.52772.58912.46321.79351.09372.39352.99452.0604
H72.54942.15324.17383.20393.29611.79351.08442.63372.54872.2131
C82.22491.44733.24512.23412.27451.09371.08442.16552.14171.4985
H94.14733.23184.22963.24042.21982.39352.63372.16551.78661.1012
H103.22232.52693.48502.60412.12552.99452.54872.14171.78661.0950
C113.14302.22523.24102.23631.49302.06042.21311.49851.10121.0950

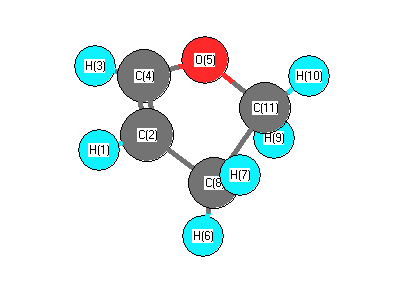
picture of Furan, 2,3-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 125.857 H1 C2 C8 126.831
C2 C4 H3 124.250 C2 C4 O5 111.110
C2 C8 H6 109.689 C2 C8 H7 115.789
C2 C8 C11 98.102 H3 C4 O5 124.628
C4 C2 C8 107.071 C4 O5 C11 101.605
O5 C11 C8 98.985 O5 C11 H9 116.860
O5 C11 H10 109.474 H6 C8 H7 110.856
H6 C8 C11 104.188 H7 C8 C11 117.024
C8 C11 H9 111.903 C8 C11 H10 110.372
H9 C11 H10 108.882
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.161      
2 C -0.242      
3 H 0.174      
4 C 0.160      
5 O -0.538      
6 H 0.163      
7 H 0.162      
8 C -0.342      
9 H 0.164      
10 H 0.159      
11 C -0.021      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 6.840 -0.198 0.043
y -0.198 6.857 0.016
z 0.043 0.016 4.234


<r2> (average value of r2) Å2
<r2> 94.148
(<r2>)1/2 9.703