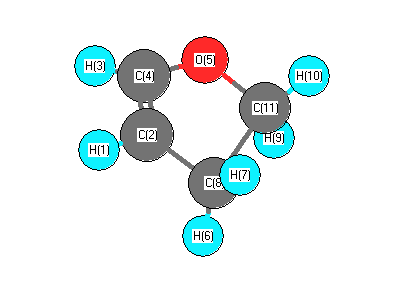
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3298 |
3136 |
7.40 |
|
|
|
| 2 |
A |
3270 |
3110 |
3.33 |
|
|
|
| 3 |
A |
3152 |
2997 |
36.04 |
|
|
|
| 4 |
A |
3104 |
2952 |
33.95 |
|
|
|
| 5 |
A |
3078 |
2927 |
38.57 |
|
|
|
| 6 |
A |
3051 |
2901 |
43.86 |
|
|
|
| 7 |
A |
1724 |
1640 |
60.69 |
|
|
|
| 8 |
A |
1547 |
1471 |
0.20 |
|
|
|
| 9 |
A |
1522 |
1448 |
0.11 |
|
|
|
| 10 |
A |
1434 |
1364 |
4.71 |
|
|
|
| 11 |
A |
1345 |
1279 |
3.20 |
|
|
|
| 12 |
A |
1321 |
1257 |
7.52 |
|
|
|
| 13 |
A |
1265 |
1203 |
7.88 |
|
|
|
| 14 |
A |
1229 |
1169 |
17.38 |
|
|
|
| 15 |
A |
1192 |
1133 |
77.18 |
|
|
|
| 16 |
A |
1118 |
1064 |
31.24 |
|
|
|
| 17 |
A |
1086 |
1033 |
3.17 |
|
|
|
| 18 |
A |
1036 |
985 |
8.53 |
|
|
|
| 19 |
A |
967 |
920 |
26.90 |
|
|
|
| 20 |
A |
957 |
910 |
26.62 |
|
|
|
| 21 |
A |
924 |
879 |
2.83 |
|
|
|
| 22 |
A |
858 |
816 |
1.23 |
|
|
|
| 23 |
A |
846 |
805 |
5.16 |
|
|
|
| 24 |
A |
726 |
690 |
34.29 |
|
|
|
| 25 |
A |
697 |
662 |
2.07 |
|
|
|
| 26 |
A |
457 |
435 |
23.28 |
|
|
|
| 27 |
A |
94 |
89 |
1.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20649.1 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 19637.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.167 |
|
|
|
| 2 |
C |
-0.241 |
|
|
|
| 3 |
H |
0.179 |
|
|
|
| 4 |
C |
0.122 |
|
|
|
| 5 |
O |
-0.469 |
|
|
|
| 6 |
H |
0.180 |
|
|
|
| 7 |
H |
0.179 |
|
|
|
| 8 |
C |
-0.386 |
|
|
|
| 9 |
H |
0.179 |
|
|
|
| 10 |
H |
0.175 |
|
|
|
| 11 |
C |
-0.083 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.730 |
0.881 |
0.111 |
1.149 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.255 |
1.206 |
0.067 |
| y |
1.206 |
-30.864 |
-0.123 |
| z |
0.067 |
-0.123 |
-30.683 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.518 |
1.206 |
0.067 |
| y |
1.206 |
-2.395 |
-0.123 |
| z |
0.067 |
-0.123 |
-2.123 |
|
| Polar |
|---|
| 3z2-r2 | -4.247 |
|---|
| x2-y2 | 4.609 |
|---|
| xy | 1.206 |
|---|
| xz | 0.067 |
|---|
| yz | -0.123 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.889 |
-0.227 |
0.042 |
| y |
-0.227 |
6.911 |
0.003 |
| z |
0.042 |
0.003 |
4.271 |
<r2> (average value of r
2) Å
2
| <r2> |
93.530 |
| (<r2>)1/2 |
9.671 |