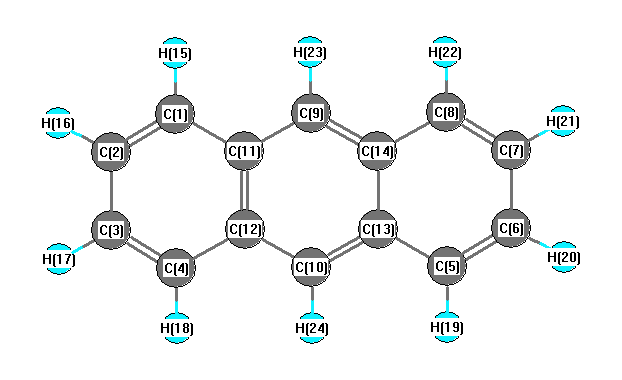
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3221 |
3081 |
0.00 |
|
|
|
| 2 |
Ag |
3194 |
3056 |
0.00 |
|
|
|
| 3 |
Ag |
3186 |
3048 |
0.00 |
|
|
|
| 4 |
Ag |
1623 |
1552 |
0.00 |
|
|
|
| 5 |
Ag |
1539 |
1473 |
0.00 |
|
|
|
| 6 |
Ag |
1463 |
1400 |
0.00 |
|
|
|
| 7 |
Ag |
1311 |
1254 |
0.00 |
|
|
|
| 8 |
Ag |
1198 |
1146 |
0.00 |
|
|
|
| 9 |
Ag |
1047 |
1001 |
0.00 |
|
|
|
| 10 |
Ag |
771 |
737 |
0.00 |
|
|
|
| 11 |
Ag |
637 |
610 |
0.00 |
|
|
|
| 12 |
Ag |
399 |
382 |
0.00 |
|
|
|
| 13 |
Au |
991 |
948 |
0.00 |
|
|
|
| 14 |
Au |
870 |
832 |
0.00 |
|
|
|
| 15 |
Au |
757 |
724 |
0.00 |
|
|
|
| 16 |
Au |
504 |
483 |
0.00 |
|
|
|
| 17 |
Au |
123 |
118 |
0.00 |
|
|
|
| 18 |
B1g |
960 |
918 |
0.00 |
|
|
|
| 19 |
B1g |
774 |
740 |
0.00 |
|
|
|
| 20 |
B1g |
486 |
465 |
0.00 |
|
|
|
| 21 |
B1g |
237 |
227 |
0.00 |
|
|
|
| 22 |
B1u |
3208 |
3069 |
63.74 |
|
|
|
| 23 |
B1u |
3189 |
3051 |
12.52 |
|
|
|
| 24 |
B1u |
3184 |
3046 |
13.84 |
|
|
|
| 25 |
B1u |
1698 |
1625 |
6.77 |
|
|
|
| 26 |
B1u |
1504 |
1439 |
1.56 |
|
|
|
| 27 |
B1u |
1350 |
1292 |
5.71 |
|
|
|
| 28 |
B1u |
1297 |
1240 |
7.67 |
|
|
|
| 29 |
B1u |
1181 |
1130 |
6.18 |
|
|
|
| 30 |
B1u |
914 |
875 |
1.62 |
|
|
|
| 31 |
B1u |
661 |
632 |
0.69 |
|
|
|
| 32 |
B1u |
234 |
224 |
1.32 |
|
|
|
| 33 |
B2g |
992 |
949 |
0.00 |
|
|
|
| 34 |
B2g |
917 |
877 |
0.00 |
|
|
|
| 35 |
B2g |
849 |
812 |
0.00 |
|
|
|
| 36 |
B2g |
780 |
746 |
0.00 |
|
|
|
| 37 |
B2g |
590 |
565 |
0.00 |
|
|
|
| 38 |
B2g |
272 |
260 |
0.00 |
|
|
|
| 39 |
B2u |
3220 |
3081 |
68.58 |
|
|
|
| 40 |
B2u |
3193 |
3055 |
0.01 |
|
|
|
| 41 |
B2u |
1607 |
1538 |
5.11 |
|
|
|
| 42 |
B2u |
1500 |
1435 |
2.26 |
|
|
|
| 43 |
B2u |
1450 |
1387 |
1.06 |
|
|
|
| 44 |
B2u |
1407 |
1346 |
4.07 |
|
|
|
| 45 |
B2u |
1203 |
1151 |
2.46 |
|
|
|
| 46 |
B2u |
1178 |
1127 |
1.44 |
|
|
|
| 47 |
B2u |
1043 |
998 |
5.02 |
|
|
|
| 48 |
B2u |
831 |
795 |
0.01 |
|
|
|
| 49 |
B2u |
615 |
588 |
8.39 |
|
|
|
| 50 |
B3g |
3208 |
3069 |
0.00 |
|
|
|
| 51 |
B3g |
3189 |
3051 |
0.00 |
|
|
|
| 52 |
B3g |
1698 |
1624 |
0.00 |
|
|
|
| 53 |
B3g |
1649 |
1577 |
0.00 |
|
|
|
| 54 |
B3g |
1425 |
1363 |
0.00 |
|
|
|
| 55 |
B3g |
1305 |
1249 |
0.00 |
|
|
|
| 56 |
B3g |
1221 |
1168 |
0.00 |
|
|
|
| 57 |
B3g |
1138 |
1089 |
0.00 |
|
|
|
| 58 |
B3g |
925 |
885 |
0.00 |
|
|
|
| 59 |
B3g |
531 |
508 |
0.00 |
|
|
|
| 60 |
B3g |
391 |
374 |
0.00 |
|
|
|
| 61 |
B3u |
965 |
923 |
7.60 |
|
|
|
| 62 |
B3u |
893 |
854 |
63.92 |
|
|
|
| 63 |
B3u |
742 |
710 |
70.00 |
|
|
|
| 64 |
B3u |
482 |
461 |
16.18 |
|
|
|
| 65 |
B3u |
386 |
370 |
0.08 |
|
|
|
| 66 |
B3u |
93 |
89 |
1.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42797.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 40944.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.204 |
|
|
|
| 2 |
C |
-0.174 |
|
|
|
| 3 |
C |
-0.174 |
|
|
|
| 4 |
C |
-0.204 |
|
|
|
| 5 |
C |
-0.204 |
|
|
|
| 6 |
C |
-0.174 |
|
|
|
| 7 |
C |
-0.174 |
|
|
|
| 8 |
C |
-0.204 |
|
|
|
| 9 |
C |
-0.304 |
|
|
|
| 10 |
C |
-0.304 |
|
|
|
| 11 |
C |
0.121 |
|
|
|
| 12 |
C |
0.121 |
|
|
|
| 13 |
C |
0.121 |
|
|
|
| 14 |
C |
0.121 |
|
|
|
| 15 |
H |
0.164 |
|
|
|
| 16 |
H |
0.164 |
|
|
|
| 17 |
H |
0.164 |
|
|
|
| 18 |
H |
0.164 |
|
|
|
| 19 |
H |
0.164 |
|
|
|
| 20 |
H |
0.164 |
|
|
|
| 21 |
H |
0.164 |
|
|
|
| 22 |
H |
0.164 |
|
|
|
| 23 |
H |
0.163 |
|
|
|
| 24 |
H |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-86.067 |
0.000 |
0.000 |
| y |
0.000 |
-68.345 |
0.000 |
| z |
0.000 |
0.000 |
-69.288 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-17.251 |
0.000 |
0.000 |
| y |
0.000 |
9.332 |
0.000 |
| z |
0.000 |
0.000 |
7.919 |
|
| Polar |
|---|
| 3z2-r2 | 15.837 |
|---|
| x2-y2 | -17.722 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.616 |
0.000 |
0.000 |
| y |
0.000 |
39.689 |
0.000 |
| z |
0.000 |
0.000 |
22.227 |
<r2> (average value of r
2) Å
2
| <r2> |
798.128 |
| (<r2>)1/2 |
28.251 |