return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H15N (triethylamine)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3 1A
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-291.188099
Energy at 298.15K-291.205634
HF Energy-290.372148
Nuclear repulsion energy338.040957
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3266 3017 48.34      
2 A 3249 3002 9.84      
3 A 3232 2985 0.47      
4 A 3178 2936 5.68      
5 A 3095 2859 174.18      
6 A 1625 1501 7.81      
7 A 1598 1476 0.00      
8 A 1593 1472 2.57      
9 A 1533 1416 14.39      
10 A 1504 1389 0.00      
11 A 1404 1297 4.22      
12 A 1255 1160 11.56      
13 A 1165 1077 3.43      
14 A 1099 1015 5.54      
15 A 864 798 0.19      
16 A 804 743 10.66      
17 A 467 431 4.24      
18 A 323 298 0.98      
19 A 205 189 0.83      
20 A 103 95 0.01      
21 E 3269 3019 45.67      
22 E 3254 3005 47.45      
23 E 3228 2981 21.84      
24 E 3178 2935 27.84      
25 E 3085 2850 10.46      
26 E 1621 1498 3.68      
26 E 1621 1498 3.68      
27 E 1605 1482 0.33      
27 E 1605 1482 0.33      
28 E 1593 1472 3.29      
28 E 1593 1472 3.29      
29 E 1529 1413 25.49      
30 E 1492 1378 8.65      
30 E 1492 1378 8.65      
31 E 1420 1312 21.58      
32 E 1331 1229 19.29      
33 E 1180 1090 14.83      
34 E 1167 1078 12.65      
35 E 991 916 1.26      
35 E 991 916 1.26      
36 E 849 784 2.79      
36 E 849 784 2.79      
37 E 500 461 2.27      
37 E 500 461 2.27      
38 E 336 311 0.48      
38 E 336 311 0.48      
39 E 230 212 0.04      
39 E 230 212 0.04      
40 E 94 87 0.21      
40 E 94 87 0.21      

Unscaled Zero Point Vibrational Energy (zpe) 47730.7 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 44088.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.07770 0.07770 0.04487

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C3

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 0.026
C2 -0.324 1.355 0.454
C3 -1.011 -0.958 0.454
C4 1.335 -0.397 0.454
C5 0.290 2.414 -0.446
C6 -2.236 -0.955 -0.446
C7 1.945 -1.458 -0.446
H8 -1.404 1.471 0.426
H9 -0.025 1.521 1.498
H10 -0.572 -1.951 0.426
H11 -1.305 -0.783 1.498
H12 1.976 0.480 0.426
H13 1.330 -0.739 1.498
H14 1.377 2.372 -0.448
H15 -0.051 2.273 -1.468
H16 0.000 3.410 -0.116
H17 -2.743 0.006 -0.448
H18 -1.943 -1.181 -1.468
H19 -2.953 -1.705 -0.116
H20 1.366 -2.379 -0.448
H21 1.994 -1.092 -1.468
H22 2.953 -1.705 -0.116

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22
N11.45741.45741.45742.47672.47672.47672.07242.11752.07242.11752.07242.11752.78352.72073.41262.78352.72073.41262.78352.72073.4126
C21.45742.41252.41251.52003.13113.72471.08651.09823.31522.57302.46042.86492.17752.14832.15702.91293.57064.07414.19603.88054.5191
C31.45742.41252.41253.72471.52003.13112.46042.86491.08651.09823.31522.57304.19603.88054.51912.17752.14832.15702.91293.57064.0741
C41.45742.41252.41253.13113.72471.52003.31522.57302.46042.86491.08651.09822.91293.57064.07414.19603.88054.51912.17752.14832.1570
C52.47671.52003.72473.13114.21124.21122.12602.16274.53424.06742.70963.84721.08721.08681.08883.87274.35385.25294.91174.03014.9155
C62.47673.13111.52003.72474.21124.21122.70963.84722.12602.16274.53424.06744.91174.03014.91551.08721.08681.08883.87274.35385.2529
C72.47673.72473.13111.52004.21124.21124.53424.06742.70963.84722.12602.16273.87274.35385.25294.91174.03014.91551.08721.08681.0888
H82.07241.08652.46043.31522.12602.70964.53421.74663.52172.49763.52173.67503.05072.46222.45392.16883.30373.57504.82234.65905.4184
H92.11751.09822.86492.57302.16273.84724.06741.74663.67502.63552.49762.63552.54493.06052.48403.66984.44754.64574.57514.43974.6779
H102.07243.31521.08652.46044.53422.12602.70963.52173.67501.74663.52172.49764.82234.65905.41843.05072.46222.45392.16883.30373.5750
H112.11752.57301.09822.86494.06742.16273.84722.49762.63551.74663.67502.63554.57514.43974.67792.54493.06052.48403.66984.44754.6457
H122.07242.46043.31521.08652.70964.53422.12603.52172.49763.52173.67501.74662.16883.30373.57504.82234.65905.41843.05072.46222.4539
H132.11752.86492.57301.09823.84724.06742.16273.67502.63552.49762.63551.74663.66984.44754.64574.57514.43974.67792.54493.06052.4840
H142.78352.17754.19602.91291.08724.91173.87273.05072.54494.82234.57512.16883.66981.75801.75574.75094.96885.95664.75093.66434.3838
H152.72072.14833.88053.57061.08684.03014.35382.46223.06054.65904.43973.30374.44751.75801.76773.66433.93835.10654.96883.93835.1652
H163.41262.15704.51914.07411.08884.91555.25292.45392.48405.41844.67793.57504.64571.75571.76774.38385.16525.90585.95665.10655.9058
H172.78352.91292.17754.19603.87271.08724.91172.16883.66983.05072.54494.82234.57514.75093.66434.38381.75801.75574.75094.96885.9566
H182.72073.57062.14833.88054.35381.08684.03013.30374.44752.46223.06054.65904.43974.96883.93835.16521.75801.76773.66433.93835.1065
H193.41264.07412.15704.51915.25291.08884.91553.57504.64572.45392.48405.41844.67795.95665.10655.90581.75571.76774.38385.16525.9058
H202.78354.19602.91292.17754.91173.87271.08724.82234.57512.16883.66983.05072.54494.75094.96885.95664.75093.66434.38381.75801.7557
H212.72073.88053.57062.14834.03014.35381.08684.65904.43973.30374.44752.46223.06053.66433.93835.10654.96883.93835.16521.75801.7677
H223.41264.51914.07412.15704.91555.25291.08885.41844.67793.57504.64572.45392.48404.38385.16525.90585.95665.10655.90581.75571.7677

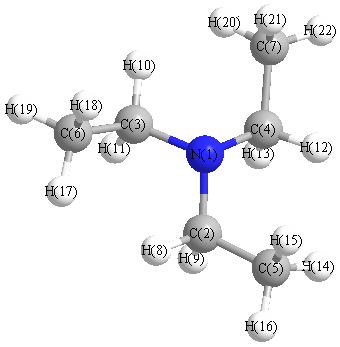
picture of triethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C5 112.560 N1 C2 H8 108.225
N1 C2 H9 111.122 N1 C3 C6 112.560
N1 C3 H10 108.225 N1 C3 H11 111.122
N1 C4 C7 112.560 N1 C4 H12 108.225
N1 C4 H13 111.122 C2 N1 C3 111.724
C2 N1 C4 111.724 C2 C5 H14 112.203
C2 C5 H15 109.886 C2 C5 H16 110.458
C3 N1 C4 111.724 C3 C6 H17 112.203
C3 C6 H18 109.886 C3 C6 H19 110.458
C4 C7 H20 112.203 C4 C7 H21 109.886
C4 C7 H22 110.458 C5 C2 H8 108.157
C5 C2 H9 110.344 C6 C3 H10 108.157
C6 C3 H11 110.344 C7 C4 H12 108.157
C7 C4 H13 110.344 H8 C2 H9 106.159
H10 C3 H11 106.159 H12 C4 H13 106.159
H14 C5 H15 107.929 H14 C5 H16 107.574
H15 C5 H16 108.680 H17 C6 H18 107.929
H17 C6 H19 107.574 H18 C6 H19 108.680
H20 C7 H21 107.929 H20 C7 H22 107.574
H21 C7 H22 108.680
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability