return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C7H8 (Norbornadiene)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-270.606313
Energy at 298.15K-270.615701
Nuclear repulsion energy287.392700
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3256 3100 8.48      
2 A1 3144 2994 10.17      
3 A1 3087 2940 37.93      
4 A1 1676 1596 0.07      
5 A1 1547 1473 0.30      
6 A1 1294 1233 2.50      
7 A1 1157 1102 0.17      
8 A1 972 926 2.75      
9 A1 922 878 6.97      
10 A1 807 768 0.49      
11 A1 750 714 51.03      
12 A1 434 413 5.12      
13 A2 3228 3074 0.00      
14 A2 1346 1282 0.00      
15 A2 1298 1236 0.00      
16 A2 1161 1106 0.00      
17 A2 948 903 0.00      
18 A2 897 855 0.00      
19 A2 728 693 0.00      
20 A2 452 431 0.00      
21 B1 3253 3098 23.93      
22 B1 3145 2995 35.17      
23 B1 1640 1562 3.01      
24 B1 1283 1222 0.58      
25 B1 1113 1060 0.45      
26 B1 1065 1014 0.56      
27 B1 947 902 1.63      
28 B1 682 650 31.08      
29 B1 518 494 6.07      
30 B2 3230 3076 12.82      
31 B2 3141 2991 56.58      
32 B2 1377 1312 20.84      
33 B2 1308 1245 0.00      
34 B2 1215 1157 0.54      
35 B2 991 944 0.02      
36 B2 924 880 0.39      
37 B2 898 855 3.59      
38 B2 833 793 7.37      
39 B2 560 533 1.37      

Unscaled Zero Point Vibrational Energy (zpe) 28613.9 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 27249.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.14279 0.12030 0.10603

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.351
C2 0.000 1.121 0.275
C3 0.000 -1.121 0.275
C4 1.240 0.671 -0.521
C5 1.240 -0.671 -0.521
C6 -1.240 0.671 -0.521
C7 -1.240 -0.671 -0.521
H8 -0.901 0.000 1.976
H9 0.901 0.000 1.976
H10 0.000 2.160 0.620
H11 0.000 -2.160 0.620
H12 1.931 1.342 -1.025
H13 1.931 -1.342 -1.025
H14 -1.931 1.342 -1.025
H15 -1.931 -1.342 -1.025

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15
C11.55441.55442.34362.34362.34362.34361.09691.09692.28072.28073.34283.34283.34283.3428
C21.55442.24291.54092.32051.54092.32052.22832.22831.09453.29982.33813.38892.33813.3889
C31.55442.24292.32051.54092.32051.54092.22832.22833.29981.09453.38892.33813.38892.3381
C42.34361.54092.32051.34212.48102.82073.35752.60772.24883.29481.08662.18673.28073.7900
C52.34362.32051.54091.34212.82072.48103.35752.60773.29482.24882.18671.08663.79003.2807
C62.34361.54092.32052.48102.82071.34212.60773.35752.24883.29483.28073.79001.08662.1867
C72.34362.32051.54092.82072.48101.34212.60773.35753.29482.24883.79003.28072.18671.0866
H81.09692.22832.22833.35753.35752.60772.60771.80262.70542.70544.33934.33933.44493.4449
H91.09692.22832.22832.60772.60773.35753.35751.80262.70542.70543.44493.44494.33934.3393
H102.28071.09453.29982.24883.29482.24883.29482.70542.70544.32042.66544.32392.66544.3239
H112.28073.29981.09453.29482.24883.29482.24882.70542.70544.32044.32392.66544.32392.6654
H123.34282.33813.38891.08662.18673.28073.79004.33933.44492.66544.32392.68303.86234.7027
H133.34283.38892.33812.18671.08663.79003.28074.33933.44494.32392.66542.68304.70273.8623
H143.34282.33813.38893.28073.79001.08662.18673.44494.33932.66544.32393.86234.70272.6830
H153.34283.38892.33813.79003.28072.18671.08663.44494.33934.32392.66544.70273.86232.6830

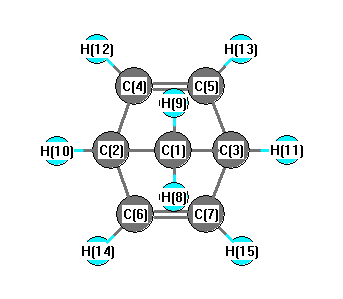
picture of Norbornadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 98.427 C1 C2 C6 98.427
C1 C2 H10 117.808 C1 C3 C5 98.427
C1 C3 C7 98.427 C1 C3 H11 117.808
C2 C1 C3 92.356 C2 C1 H8 113.245
C2 C1 H9 113.245 C2 C4 C5 106.996
C2 C4 H12 124.808 C2 C6 C7 106.996
C2 C6 H14 124.808 C3 C1 H8 113.245
C3 C1 H9 113.245 C3 C5 C4 106.996
C3 C5 H13 124.808 C3 C7 C6 106.996
C3 C7 H15 124.808 C4 C2 C6 107.225
C4 C2 H10 116.111 C4 C5 H13 128.099
C5 C3 C7 107.225 C5 C3 H11 116.111
C5 C4 H12 128.099 C6 C2 H10 116.111
C6 C7 H15 128.099 C7 C3 H11 116.111
C7 C6 H14 128.099 H8 C1 H9 110.503
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability