return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C7H8 (Norbornadiene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-271.157011
Energy at 298.15K-271.166356
HF Energy-270.856347
Nuclear repulsion energy287.659062
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3272 3106 8.67      
2 A1 3160 2999 10.14      
3 A1 3099 2941 42.72      
4 A1 1660 1576 0.12      
5 A1 1540 1461 0.10      
6 A1 1284 1219 2.33      
7 A1 1148 1089 0.21      
8 A1 962 914 2.68      
9 A1 904 859 6.09      
10 A1 794 754 0.33      
11 A1 751 713 48.89      
12 A1 432 410 4.16      
13 A2 3245 3081 0.00      
14 A2 1326 1259 0.00      
15 A2 1292 1227 0.00      
16 A2 1155 1096 0.00      
17 A2 932 884 0.00      
18 A2 904 858 0.00      
19 A2 726 689 0.00      
20 A2 456 432 0.00      
21 B1 3269 3103 24.39      
22 B1 3159 2998 38.00      
23 B1 1622 1539 4.98      
24 B1 1264 1200 0.78      
25 B1 1106 1049 0.28      
26 B1 1058 1004 0.26      
27 B1 923 876 1.67      
28 B1 679 645 30.01      
29 B1 513 487 5.90      
30 B2 3247 3082 13.40      
31 B2 3158 2997 60.41      
32 B2 1371 1301 19.70      
33 B2 1301 1235 0.01      
34 B2 1198 1137 0.78      
35 B2 978 928 0.02      
36 B2 925 878 2.01      
37 B2 898 852 2.67      
38 B2 821 779 6.58      
39 B2 555 526 1.56      

Unscaled Zero Point Vibrational Energy (zpe) 28541.3 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 27091.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.14321 0.12037 0.10608

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.351
C2 0.000 1.121 0.272
C3 0.000 -1.121 0.272
C4 1.241 0.670 -0.519
C5 1.241 -0.670 -0.519
C6 -1.241 0.670 -0.519
C7 -1.241 -0.670 -0.519
H8 -0.899 0.000 1.973
H9 0.899 0.000 1.973
H10 0.000 2.157 0.614
H11 0.000 -2.157 0.614
H12 1.930 1.339 -1.020
H13 1.930 -1.339 -1.020
H14 -1.930 1.339 -1.020
H15 -1.930 -1.339 -1.020

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15
C11.55621.55622.34282.34282.34282.34281.09351.09352.27962.27963.33793.33793.33793.3379
C21.55622.24221.53992.31821.53992.31822.22722.22721.09073.29582.33303.38322.33303.3832
C31.55622.24222.31821.53992.31821.53992.22722.22723.29581.09073.38322.33303.38322.3330
C42.34281.53992.31821.33922.48282.82093.35342.60372.24433.28841.08302.18143.27993.7872
C52.34282.31821.53991.33922.82092.48283.35342.60373.28842.24432.18141.08303.78723.2799
C62.34281.53992.31822.48282.82091.33922.60373.35342.24433.28843.27993.78721.08302.1814
C72.34282.31821.53992.82092.48281.33922.60373.35343.28842.24433.78723.27992.18141.0830
H81.09352.22722.22723.35343.35342.60372.60371.79872.70372.70374.33134.33133.43753.4375
H91.09352.22722.22722.60372.60373.35343.35341.79872.70372.70373.43753.43754.33134.3313
H102.27961.09073.29582.24433.28842.24433.28842.70372.70374.31382.65794.31442.65794.3144
H112.27963.29581.09073.28842.24433.28842.24432.70372.70374.31384.31442.65794.31442.6579
H123.33792.33303.38321.08302.18143.27993.78724.33133.43752.65794.31442.67743.86044.6979
H133.33793.38322.33302.18141.08303.78723.27994.33133.43754.31442.65792.67744.69793.8604
H143.33792.33303.38323.27993.78721.08302.18143.43754.33132.65794.31443.86044.69792.6774
H153.33793.38322.33303.78723.27992.18141.08303.43754.33134.31442.65794.69793.86042.6774

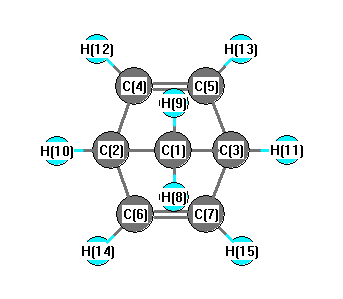
picture of Norbornadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 98.349 C1 C2 C6 98.349
C1 C2 H10 117.837 C1 C3 C5 98.349
C1 C3 C7 98.349 C1 C3 H11 117.837
C2 C1 C3 92.177 C2 C1 H8 113.239
C2 C1 H9 113.239 C2 C4 C5 107.050
C2 C4 H12 124.696 C2 C6 C7 107.050
C2 C6 H14 124.696 C3 C1 H8 113.239
C3 C1 H9 113.239 C3 C5 C4 107.050
C3 C5 H13 124.696 C3 C7 C6 107.050
C3 C7 H15 124.696 C4 C2 C6 107.447
C4 C2 H10 116.066 C4 C5 H13 128.155
C5 C3 C7 107.447 C5 C3 H11 116.066
C5 C4 H12 128.155 C6 C2 H10 116.066
C6 C7 H15 128.155 C7 C3 H11 116.066
C7 C6 H14 128.155 H8 C1 H9 110.652
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.258      
2 C -0.227      
3 C -0.227      
4 C -0.128      
5 C -0.128      
6 C -0.128      
7 C -0.128      
8 H 0.151      
9 H 0.151      
10 H 0.146      
11 H 0.146      
12 H 0.157      
13 H 0.157      
14 H 0.157      
15 H 0.157      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.957 0.000 0.000
y 0.000 10.484 0.000
z 0.000 0.000 7.466


<r2> (average value of r2) Å2
<r2> 152.450
(<r2>)1/2 12.347