return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CHO (Propanal)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-192.552239
Energy at 298.15K-192.558628
HF Energy-191.949749
Nuclear repulsion energy118.150444
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3158 3007 21.89      
2 A' 3082 2935 16.05      
3 A' 3049 2904 24.59      
4 A' 2958 2817 120.19      
5 A' 1824 1737 106.60      
6 A' 1549 1475 8.12      
7 A' 1518 1445 9.27      
8 A' 1474 1404 8.86      
9 A' 1448 1379 3.91      
10 A' 1416 1349 8.16      
11 A' 1146 1091 14.26      
12 A' 1032 983 1.09      
13 A' 888 845 17.06      
14 A' 682 650 7.12      
15 A' 270 257 9.12      
16 A" 3165 3014 21.76      
17 A" 3081 2934 12.18      
18 A" 1546 1472 5.88      
19 A" 1314 1251 0.32      
20 A" 1181 1125 0.18      
21 A" 922 878 2.05      
22 A" 700 667 2.67      
23 A" 239 228 0.30      
24 A" 137 131 2.73      

Unscaled Zero Point Vibrational Energy (zpe) 18888.9 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 17987.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.54937 0.19731 0.15340

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.456 0.450 0.000
C2 0.000 0.917 0.000
C3 -1.008 -0.209 0.000
O4 -0.709 -1.392 0.000
H5 2.141 1.306 0.000
H6 1.676 -0.160 0.881
H7 1.676 -0.160 -0.881
H8 -0.216 1.556 0.871
H9 -0.216 1.556 -0.871
H10 -2.077 0.090 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.52902.55042.84341.09661.09421.09422.18562.18563.5517
C21.52901.51042.41552.17652.17802.17801.10201.10202.2358
C32.55041.51041.22083.49452.82472.82472.12122.12121.1104
O42.84342.41551.22083.92562.82552.82553.11373.11372.0168
H51.09662.17653.49453.92561.77321.77322.52542.52544.3905
H61.09422.17802.82472.82551.77321.76242.55393.09743.8630
H71.09422.17802.82472.82551.77321.76243.09742.55393.8630
H82.18561.10202.12123.11372.52542.55393.09741.74262.5246
H92.18561.10202.12123.11372.52543.09742.55391.74262.5246
H103.55172.23581.11042.01684.39053.86303.86302.52462.5246

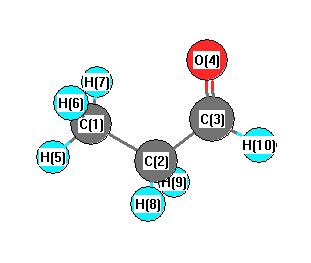
picture of Propanal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 114.088 C1 C2 H8 111.311
C1 C2 H9 111.311 C2 C1 H5 110.915
C2 C1 H6 111.171 C2 C1 H7 111.171
C2 C3 O4 124.014 C2 C3 H10 116.262
C3 C2 H8 107.547 C3 C2 H9 107.547
O4 C3 H10 119.723 H5 C1 H6 108.072
H5 C1 H7 108.072 H6 C1 H7 107.284
H8 C2 H9 104.494
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability