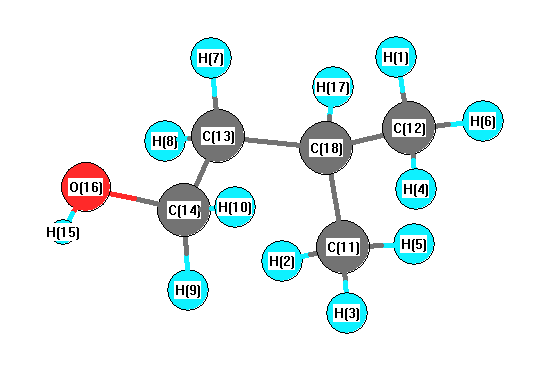
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3793 |
3629 |
14.59 |
|
|
|
| 2 |
A' |
3126 |
2991 |
47.24 |
|
|
|
| 3 |
A' |
3125 |
2989 |
79.64 |
|
|
|
| 4 |
A' |
3054 |
2922 |
11.93 |
|
|
|
| 5 |
A' |
3050 |
2918 |
46.36 |
|
|
|
| 6 |
A' |
3029 |
2897 |
14.40 |
|
|
|
| 7 |
A' |
2994 |
2864 |
50.84 |
|
|
|
| 8 |
A' |
1555 |
1488 |
1.96 |
|
|
|
| 9 |
A' |
1536 |
1470 |
7.76 |
|
|
|
| 10 |
A' |
1535 |
1468 |
10.18 |
|
|
|
| 11 |
A' |
1514 |
1449 |
2.65 |
|
|
|
| 12 |
A' |
1480 |
1416 |
4.03 |
|
|
|
| 13 |
A' |
1443 |
1380 |
3.87 |
|
|
|
| 14 |
A' |
1388 |
1328 |
1.42 |
|
|
|
| 15 |
A' |
1358 |
1300 |
31.46 |
|
|
|
| 16 |
A' |
1262 |
1208 |
32.11 |
|
|
|
| 17 |
A' |
1218 |
1166 |
4.58 |
|
|
|
| 18 |
A' |
1140 |
1091 |
0.11 |
|
|
|
| 19 |
A' |
1092 |
1044 |
99.37 |
|
|
|
| 20 |
A' |
1013 |
969 |
3.00 |
|
|
|
| 21 |
A' |
967 |
926 |
11.83 |
|
|
|
| 22 |
A' |
810 |
775 |
4.97 |
|
|
|
| 23 |
A' |
558 |
533 |
1.18 |
|
|
|
| 24 |
A' |
402 |
385 |
6.66 |
|
|
|
| 25 |
A' |
326 |
312 |
5.43 |
|
|
|
| 26 |
A' |
257 |
246 |
0.10 |
|
|
|
| 27 |
A' |
209 |
200 |
2.04 |
|
|
|
| 28 |
A" |
3124 |
2989 |
22.64 |
|
|
|
| 29 |
A" |
3119 |
2984 |
1.24 |
|
|
|
| 30 |
A" |
3096 |
2962 |
43.24 |
|
|
|
| 31 |
A" |
3047 |
2915 |
45.35 |
|
|
|
| 32 |
A" |
3023 |
2892 |
43.66 |
|
|
|
| 33 |
A" |
1523 |
1457 |
1.08 |
|
|
|
| 34 |
A" |
1512 |
1446 |
0.66 |
|
|
|
| 35 |
A" |
1424 |
1363 |
8.71 |
|
|
|
| 36 |
A" |
1383 |
1323 |
1.13 |
|
|
|
| 37 |
A" |
1333 |
1275 |
0.28 |
|
|
|
| 38 |
A" |
1261 |
1206 |
0.01 |
|
|
|
| 39 |
A" |
1191 |
1139 |
4.83 |
|
|
|
| 40 |
A" |
1018 |
974 |
0.43 |
|
|
|
| 41 |
A" |
975 |
932 |
0.00 |
|
|
|
| 42 |
A" |
935 |
895 |
1.16 |
|
|
|
| 43 |
A" |
801 |
766 |
0.02 |
|
|
|
| 44 |
A" |
369 |
353 |
0.13 |
|
|
|
| 45 |
A" |
296 |
283 |
122.07 |
|
|
|
| 46 |
A" |
241 |
231 |
1.17 |
|
|
|
| 47 |
A" |
105 |
101 |
0.01 |
|
|
|
| 48 |
A" |
70 |
67 |
9.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36538.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 34956.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.167 |
|
|
0.099 |
| 2 |
H |
0.167 |
|
|
0.099 |
| 3 |
H |
0.162 |
|
|
0.096 |
| 4 |
H |
0.162 |
|
|
0.096 |
| 5 |
H |
0.165 |
|
|
0.103 |
| 6 |
H |
0.165 |
|
|
0.103 |
| 7 |
H |
0.169 |
|
|
0.072 |
| 8 |
H |
0.169 |
|
|
0.072 |
| 9 |
H |
0.141 |
|
|
-0.015 |
| 10 |
H |
0.141 |
|
|
-0.015 |
| 11 |
C |
-0.508 |
|
|
-0.413 |
| 12 |
C |
-0.508 |
|
|
-0.413 |
| 13 |
C |
-0.315 |
|
|
-0.211 |
| 14 |
C |
-0.078 |
|
|
0.273 |
| 15 |
H |
0.403 |
|
|
0.391 |
| 16 |
O |
-0.631 |
|
|
-0.625 |
| 17 |
H |
0.158 |
|
|
-0.016 |
| 18 |
C |
-0.127 |
|
|
0.303 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.265 |
-1.441 |
0.000 |
1.465 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.240 |
-1.453 |
0.000 |
1.473 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.469 |
3.427 |
0.000 |
| y |
3.427 |
-42.644 |
0.000 |
| z |
0.000 |
0.000 |
-39.776 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.741 |
3.427 |
0.000 |
| y |
3.427 |
-5.522 |
0.000 |
| z |
0.000 |
0.000 |
-1.219 |
|
| Polar |
|---|
| 3z2-r2 | -2.439 |
|---|
| x2-y2 | 8.175 |
|---|
| xy | 3.427 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
223.359 |
| (<r2>)1/2 |
14.945 |