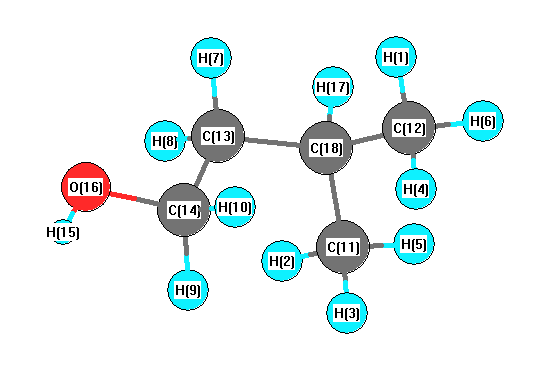
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3817 |
3630 |
16.90 |
|
|
|
| 2 |
A' |
3142 |
2988 |
47.44 |
|
|
|
| 3 |
A' |
3139 |
2985 |
73.29 |
|
|
|
| 4 |
A' |
3067 |
2917 |
16.64 |
|
|
|
| 5 |
A' |
3063 |
2913 |
40.52 |
|
|
|
| 6 |
A' |
3045 |
2896 |
12.97 |
|
|
|
| 7 |
A' |
3008 |
2861 |
49.46 |
|
|
|
| 8 |
A' |
1557 |
1480 |
1.97 |
|
|
|
| 9 |
A' |
1537 |
1462 |
7.42 |
|
|
|
| 10 |
A' |
1535 |
1460 |
11.51 |
|
|
|
| 11 |
A' |
1515 |
1441 |
2.41 |
|
|
|
| 12 |
A' |
1485 |
1412 |
3.86 |
|
|
|
| 13 |
A' |
1447 |
1376 |
4.35 |
|
|
|
| 14 |
A' |
1389 |
1321 |
1.34 |
|
|
|
| 15 |
A' |
1362 |
1296 |
33.89 |
|
|
|
| 16 |
A' |
1268 |
1206 |
32.51 |
|
|
|
| 17 |
A' |
1224 |
1164 |
4.64 |
|
|
|
| 18 |
A' |
1145 |
1089 |
1.66 |
|
|
|
| 19 |
A' |
1102 |
1048 |
97.57 |
|
|
|
| 20 |
A' |
1020 |
970 |
3.26 |
|
|
|
| 21 |
A' |
976 |
928 |
11.29 |
|
|
|
| 22 |
A' |
819 |
779 |
4.78 |
|
|
|
| 23 |
A' |
562 |
535 |
1.16 |
|
|
|
| 24 |
A' |
406 |
386 |
6.71 |
|
|
|
| 25 |
A' |
328 |
312 |
5.33 |
|
|
|
| 26 |
A' |
250 |
238 |
0.14 |
|
|
|
| 27 |
A' |
210 |
200 |
2.10 |
|
|
|
| 28 |
A" |
3140 |
2986 |
19.78 |
|
|
|
| 29 |
A" |
3134 |
2980 |
2.49 |
|
|
|
| 30 |
A" |
3111 |
2959 |
42.02 |
|
|
|
| 31 |
A" |
3060 |
2910 |
44.99 |
|
|
|
| 32 |
A" |
3039 |
2890 |
40.79 |
|
|
|
| 33 |
A" |
1524 |
1449 |
1.04 |
|
|
|
| 34 |
A" |
1512 |
1438 |
0.59 |
|
|
|
| 35 |
A" |
1430 |
1360 |
10.12 |
|
|
|
| 36 |
A" |
1386 |
1319 |
0.70 |
|
|
|
| 37 |
A" |
1335 |
1270 |
0.24 |
|
|
|
| 38 |
A" |
1262 |
1201 |
0.02 |
|
|
|
| 39 |
A" |
1196 |
1138 |
4.51 |
|
|
|
| 40 |
A" |
1023 |
973 |
0.60 |
|
|
|
| 41 |
A" |
978 |
930 |
0.00 |
|
|
|
| 42 |
A" |
938 |
892 |
1.18 |
|
|
|
| 43 |
A" |
802 |
762 |
0.01 |
|
|
|
| 44 |
A" |
369 |
351 |
0.13 |
|
|
|
| 45 |
A" |
297 |
282 |
122.39 |
|
|
|
| 46 |
A" |
234 |
222 |
1.35 |
|
|
|
| 47 |
A" |
104 |
99 |
0.00 |
|
|
|
| 48 |
A" |
62 |
59 |
9.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36675.7 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 34878.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.169 |
|
|
|
| 2 |
H |
0.169 |
|
|
|
| 3 |
H |
0.163 |
|
|
|
| 4 |
H |
0.163 |
|
|
|
| 5 |
H |
0.168 |
|
|
|
| 6 |
H |
0.168 |
|
|
|
| 7 |
H |
0.171 |
|
|
|
| 8 |
H |
0.171 |
|
|
|
| 9 |
H |
0.142 |
|
|
|
| 10 |
H |
0.142 |
|
|
|
| 11 |
C |
-0.512 |
|
|
|
| 12 |
C |
-0.512 |
|
|
|
| 13 |
C |
-0.320 |
|
|
|
| 14 |
C |
-0.080 |
|
|
|
| 15 |
H |
0.405 |
|
|
|
| 16 |
O |
-0.634 |
|
|
|
| 17 |
H |
0.161 |
|
|
|
| 18 |
C |
-0.135 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.393 |
-1.408 |
0.000 |
1.462 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.034 |
2.636 |
0.000 |
| y |
2.636 |
-43.266 |
0.000 |
| z |
0.000 |
0.000 |
-39.878 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.538 |
2.636 |
0.000 |
| y |
2.636 |
-6.310 |
0.000 |
| z |
0.000 |
0.000 |
-1.228 |
|
| Polar |
|---|
| 3z2-r2 | -2.455 |
|---|
| x2-y2 | 9.232 |
|---|
| xy | 2.636 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.078 |
-0.677 |
0.000 |
| y |
-0.677 |
8.221 |
0.000 |
| z |
0.000 |
0.000 |
8.121 |
<r2> (average value of r
2) Å
2
| <r2> |
222.046 |
| (<r2>)1/2 |
14.901 |