Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B2PLYP/6-31G*
Geometric Data calculated at B2PLYP/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B2PLYP/6-31G*
Geometric Data calculated at B2PLYP/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -345.426082 |
| Energy at 298.15K | |
| HF Energy | -345.094681 |
| Nuclear repulsion energy | 296.085168 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3205 |
3042 |
7.06 |
|
|
|
| 2 |
A |
3151 |
2991 |
5.61 |
|
|
|
| 3 |
A |
3118 |
2959 |
2.67 |
|
|
|
| 4 |
A |
3083 |
2926 |
0.33 |
|
|
|
| 5 |
A |
1807 |
1715 |
16.67 |
|
|
|
| 6 |
A |
1523 |
1446 |
5.60 |
|
|
|
| 7 |
A |
1521 |
1444 |
16.05 |
|
|
|
| 8 |
A |
1508 |
1431 |
7.20 |
|
|
|
| 9 |
A |
1432 |
1359 |
6.43 |
|
|
|
| 10 |
A |
1300 |
1234 |
27.03 |
|
|
|
| 11 |
A |
1168 |
1108 |
1.57 |
|
|
|
| 12 |
A |
1100 |
1044 |
0.61 |
|
|
|
| 13 |
A |
962 |
913 |
1.18 |
|
|
|
| 14 |
A |
802 |
762 |
0.20 |
|
|
|
| 15 |
A |
631 |
599 |
1.29 |
|
|
|
| 16 |
A |
489 |
464 |
6.91 |
|
|
|
| 17 |
A |
323 |
306 |
0.78 |
|
|
|
| 18 |
A |
175 |
167 |
0.13 |
|
|
|
| 19 |
A |
147 |
139 |
0.90 |
|
|
|
| 20 |
A |
61 |
58 |
7.16 |
|
|
|
| 21 |
B |
3205 |
3042 |
7.33 |
|
|
|
| 22 |
B |
3186 |
3024 |
6.03 |
|
|
|
| 23 |
B |
3150 |
2990 |
1.10 |
|
|
|
| 24 |
B |
3083 |
2926 |
3.00 |
|
|
|
| 25 |
B |
1783 |
1693 |
248.61 |
|
|
|
| 26 |
B |
1521 |
1444 |
0.80 |
|
|
|
| 27 |
B |
1516 |
1439 |
26.97 |
|
|
|
| 28 |
B |
1433 |
1360 |
81.77 |
|
|
|
| 29 |
B |
1283 |
1218 |
114.57 |
|
|
|
| 30 |
B |
1218 |
1156 |
151.61 |
|
|
|
| 31 |
B |
1083 |
1028 |
6.10 |
|
|
|
| 32 |
B |
1024 |
972 |
0.66 |
|
|
|
| 33 |
B |
915 |
869 |
18.12 |
|
|
|
| 34 |
B |
814 |
773 |
8.62 |
|
|
|
| 35 |
B |
548 |
520 |
25.77 |
|
|
|
| 36 |
B |
499 |
474 |
1.30 |
|
|
|
| 37 |
B |
415 |
394 |
1.25 |
|
|
|
| 38 |
B |
175 |
166 |
0.36 |
|
|
|
| 39 |
B |
56 |
53 |
11.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27205.2 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 25823.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.020 |
| C2 |
0.000 |
1.230 |
0.107 |
| C3 |
0.000 |
-1.230 |
0.107 |
| C4 |
-1.357 |
1.750 |
-0.302 |
| C5 |
1.357 |
-1.750 |
-0.302 |
| O6 |
1.049 |
1.706 |
-0.295 |
| O7 |
-1.049 |
-1.706 |
-0.295 |
| H8 |
-0.898 |
-0.031 |
1.639 |
| H9 |
0.898 |
0.031 |
1.639 |
| H10 |
-1.249 |
2.511 |
-1.075 |
| H11 |
-1.975 |
0.921 |
-0.661 |
| H12 |
-1.867 |
2.179 |
0.567 |
| H13 |
1.249 |
-2.511 |
-1.075 |
| H14 |
1.975 |
-0.921 |
-0.661 |
| H15 |
1.867 |
-2.179 |
0.567 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
O6 |
O7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.5322 | 1.5322 | 2.5793 | 2.5793 | 2.3956 | 2.3956 | 1.0916 | 1.0916 | 3.5006 | 2.7528 | 2.9048 | 3.5006 | 2.7528 | 2.9048 |
C2 | 1.5322 | | 2.4608 | 1.5097 | 3.3001 | 1.2198 | 3.1436 | 2.1786 | 2.1434 | 2.1440 | 2.1421 | 2.1439 | 4.1175 | 3.0200 | 3.9140 | C3 | 1.5322 | 2.4608 | | 3.3001 | 1.5097 | 3.1436 | 1.2198 | 2.1434 | 2.1786 | 4.1175 | 3.0200 | 3.9140 | 2.1440 | 2.1421 | 2.1439 | C4 | 2.5793 | 1.5097 | 3.3001 | | 4.4288 | 2.4066 | 3.4692 | 2.6744 | 3.4371 | 1.0898 | 1.0946 | 1.0953 | 5.0538 | 4.2857 | 5.1559 | C5 | 2.5793 | 3.3001 | 1.5097 | 4.4288 | | 3.4692 | 2.4066 | 3.4371 | 2.6744 | 5.0538 | 4.2857 | 5.1559 | 1.0898 | 1.0946 | 1.0953 | O6 | 2.3956 | 1.2198 | 3.1436 | 2.4066 | 3.4692 | | 4.0049 | 3.2480 | 2.5630 | 2.5569 | 3.1460 | 3.0771 | 4.2927 | 2.8092 | 4.0621 | O7 | 2.3956 | 3.1436 | 1.2198 | 3.4692 | 2.4066 | 4.0049 | | 2.5630 | 3.2480 | 4.2927 | 2.8092 | 4.0621 | 2.5569 | 3.1460 | 3.0771 | H8 | 1.0916 | 2.1786 | 2.1434 | 2.6744 | 3.4371 | 3.2480 | 2.5630 | | 1.7980 | 3.7352 | 2.7128 | 2.6403 | 4.2580 | 3.7875 | 3.6621 | H9 | 1.0916 | 2.1434 | 2.1786 | 3.4371 | 2.6744 | 2.5630 | 3.2480 | 1.7980 | | 4.2580 | 3.7875 | 3.6621 | 3.7352 | 2.7128 | 2.6403 | H10 | 3.5006 | 2.1440 | 4.1175 | 1.0898 | 5.0538 | 2.5569 | 4.2927 | 3.7352 | 4.2580 | | 1.7963 | 1.7854 | 5.6085 | 4.7270 | 5.8649 | H11 | 2.7528 | 2.1421 | 3.0200 | 1.0946 | 4.2857 | 3.1460 | 2.8092 | 2.7128 | 3.7875 | 1.7963 | | 1.7614 | 4.7270 | 4.3590 | 5.0872 | H12 | 2.9048 | 2.1439 | 3.9140 | 1.0953 | 5.1559 | 3.0771 | 4.0621 | 2.6403 | 3.6621 | 1.7854 | 1.7614 | | 5.8649 | 5.0872 | 5.7384 | H13 | 3.5006 | 4.1175 | 2.1440 | 5.0538 | 1.0898 | 4.2927 | 2.5569 | 4.2580 | 3.7352 | 5.6085 | 4.7270 | 5.8649 | | 1.7963 | 1.7854 | H14 | 2.7528 | 3.0200 | 2.1421 | 4.2857 | 1.0946 | 2.8092 | 3.1460 | 3.7875 | 2.7128 | 4.7270 | 4.3590 | 5.0872 | 1.7963 | | 1.7614 | H15 | 2.9048 | 3.9140 | 2.1439 | 5.1559 | 1.0953 | 4.0621 | 3.0771 | 3.6621 | 2.6403 | 5.8649 | 5.0872 | 5.7384 | 1.7854 | 1.7614 | |
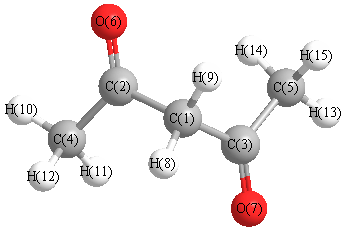 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
115.971 |
|
C1 |
C2 |
O6 |
120.613 |
| C1 |
C3 |
C5 |
115.971 |
|
C1 |
C3 |
O7 |
120.613 |
| C2 |
C1 |
C3 |
106.836 |
|
C2 |
C1 |
H8 |
111.149 |
| C2 |
C1 |
H9 |
108.383 |
|
C2 |
C4 |
H10 |
110.081 |
| C2 |
C4 |
H11 |
109.644 |
|
C2 |
C4 |
H12 |
109.742 |
| C3 |
C1 |
H8 |
108.383 |
|
C3 |
C1 |
H9 |
111.149 |
| C3 |
C5 |
H13 |
110.081 |
|
C3 |
C5 |
H14 |
109.644 |
| C3 |
C5 |
H15 |
109.742 |
|
C4 |
C2 |
O6 |
123.344 |
| C5 |
C3 |
O7 |
123.344 |
|
H8 |
C1 |
H9 |
110.879 |
| H10 |
C4 |
H11 |
110.635 |
|
H10 |
C4 |
H12 |
109.592 |
| H11 |
C4 |
H12 |
107.095 |
|
H13 |
C5 |
H14 |
110.635 |
| H13 |
C5 |
H15 |
109.592 |
|
H14 |
C5 |
H15 |
107.095 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.446 |
|
|
|
| 2 |
C |
0.471 |
|
|
|
| 3 |
C |
0.471 |
|
|
|
| 4 |
C |
-0.554 |
|
|
|
| 5 |
C |
-0.554 |
|
|
|
| 6 |
O |
-0.472 |
|
|
|
| 7 |
O |
-0.472 |
|
|
|
| 8 |
H |
0.194 |
|
|
|
| 9 |
H |
0.194 |
|
|
|
| 10 |
H |
0.195 |
|
|
|
| 11 |
H |
0.206 |
|
|
|
| 12 |
H |
0.183 |
|
|
|
| 13 |
H |
0.195 |
|
|
|
| 14 |
H |
0.206 |
|
|
|
| 15 |
H |
0.183 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.916 |
-0.162 |
0.000 |
| y |
-0.162 |
8.806 |
0.000 |
| z |
0.000 |
0.000 |
6.738 |
<r2> (average value of r
2) Å
2
| <r2> |
229.506 |
| (<r2>)1/2 |
15.149 |
