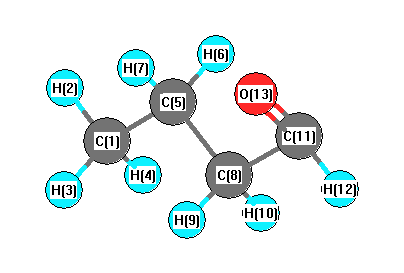
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3134 |
2998 |
33.38 |
|
|
|
| 2 |
A' |
3073 |
2940 |
18.45 |
|
|
|
| 3 |
A' |
3052 |
2919 |
26.28 |
|
|
|
| 4 |
A' |
3022 |
2891 |
22.79 |
|
|
|
| 5 |
A' |
2901 |
2776 |
159.36 |
|
|
|
| 6 |
A' |
1848 |
1768 |
144.41 |
|
|
|
| 7 |
A' |
1532 |
1465 |
6.13 |
|
|
|
| 8 |
A' |
1513 |
1448 |
2.83 |
|
|
|
| 9 |
A' |
1475 |
1411 |
13.67 |
|
|
|
| 10 |
A' |
1441 |
1379 |
3.54 |
|
|
|
| 11 |
A' |
1434 |
1372 |
6.82 |
|
|
|
| 12 |
A' |
1416 |
1355 |
18.96 |
|
|
|
| 13 |
A' |
1325 |
1268 |
5.33 |
|
|
|
| 14 |
A' |
1147 |
1098 |
12.16 |
|
|
|
| 15 |
A' |
1070 |
1024 |
0.11 |
|
|
|
| 16 |
A' |
974 |
931 |
6.56 |
|
|
|
| 17 |
A' |
869 |
831 |
8.79 |
|
|
|
| 18 |
A' |
700 |
670 |
13.50 |
|
|
|
| 19 |
A' |
347 |
332 |
0.78 |
|
|
|
| 20 |
A' |
195 |
187 |
8.52 |
|
|
|
| 21 |
A" |
3129 |
2994 |
55.81 |
|
|
|
| 22 |
A" |
3105 |
2970 |
0.11 |
|
|
|
| 23 |
A" |
3051 |
2919 |
9.27 |
|
|
|
| 24 |
A" |
1527 |
1461 |
7.66 |
|
|
|
| 25 |
A" |
1335 |
1277 |
0.16 |
|
|
|
| 26 |
A" |
1264 |
1210 |
0.07 |
|
|
|
| 27 |
A" |
1162 |
1111 |
0.15 |
|
|
|
| 28 |
A" |
955 |
914 |
0.02 |
|
|
|
| 29 |
A" |
794 |
759 |
1.98 |
|
|
|
| 30 |
A" |
673 |
644 |
4.36 |
|
|
|
| 31 |
A" |
253 |
242 |
0.03 |
|
|
|
| 32 |
A" |
178 |
170 |
1.23 |
|
|
|
| 33 |
A" |
80 |
76 |
1.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24986.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 23904.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.513 |
|
|
-0.410 |
| 2 |
H |
0.174 |
|
|
0.100 |
| 3 |
H |
0.165 |
|
|
0.100 |
| 4 |
H |
0.165 |
|
|
0.100 |
| 5 |
C |
-0.303 |
|
|
0.228 |
| 6 |
H |
0.180 |
|
|
-0.010 |
| 7 |
H |
0.180 |
|
|
-0.010 |
| 8 |
C |
-0.405 |
|
|
-0.188 |
| 9 |
H |
0.187 |
|
|
0.055 |
| 10 |
H |
0.187 |
|
|
0.055 |
| 11 |
C |
0.242 |
|
|
0.458 |
| 12 |
H |
0.125 |
|
|
-0.033 |
| 13 |
O |
-0.385 |
|
|
-0.444 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.749 |
1.840 |
0.000 |
2.538 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.774 |
1.856 |
0.000 |
2.567 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.193 |
-3.466 |
0.000 |
| y |
-3.466 |
-30.424 |
0.000 |
| z |
0.000 |
0.000 |
-30.307 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.827 |
-3.466 |
0.000 |
| y |
-3.466 |
2.825 |
0.000 |
| z |
0.000 |
0.000 |
3.002 |
|
| Polar |
|---|
| 3z2-r2 | 6.003 |
|---|
| x2-y2 | -5.768 |
|---|
| xy | -3.466 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
154.841 |
| (<r2>)1/2 |
12.444 |