return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCH2CH2CH3 (Butanal)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-232.186882
Energy at 298.15K-232.195510
HF Energy-231.956605
Nuclear repulsion energy171.754910
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3155 2995 34.16      
2 A' 3096 2939 17.88      
3 A' 3076 2919 24.46      
4 A' 3046 2891 21.14      
5 A' 2940 2791 149.58      
6 A' 1810 1718 118.81      
7 A' 1557 1478 5.00      
8 A' 1539 1461 2.44      
9 A' 1504 1427 12.23      
10 A' 1464 1390 1.83      
11 A' 1450 1376 4.65      
12 A' 1435 1362 17.63      
13 A' 1344 1276 4.13      
14 A' 1159 1100 11.41      
15 A' 1075 1020 0.31      
16 A' 983 933 6.55      
17 A' 876 831 7.30      
18 A' 702 666 13.82      
19 A' 351 334 0.86      
20 A' 201 191 8.44      
21 A" 3151 2991 58.27      
22 A" 3128 2969 0.18      
23 A" 3075 2918 10.20      
24 A" 1552 1473 6.41      
25 A" 1353 1284 0.20      
26 A" 1284 1219 0.05      
27 A" 1181 1121 0.07      
28 A" 966 917 0.06      
29 A" 806 766 1.44      
30 A" 688 653 3.05      
31 A" 255 242 0.03      
32 A" 174 165 1.39      
33 A" 82 78 1.75      

Unscaled Zero Point Vibrational Energy (zpe) 25228.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23947.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.49986 0.08519 0.07583

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.289 -0.363 0.000
H2 2.821 -1.316 0.000
H3 2.596 0.201 0.885
H4 2.596 0.201 -0.885
C5 0.765 -0.585 0.000
H6 0.477 -1.161 -0.880
H7 0.477 -1.161 0.880
C8 0.000 0.749 0.000
H9 0.296 1.334 0.880
H10 0.296 1.334 -0.880
C11 -1.495 0.624 0.000
H12 -2.073 1.539 0.000
O13 -2.106 -0.440 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 C11 H12 O13
C11.09161.09321.09321.53942.16652.16652.54462.76132.76133.90974.75794.3949
H21.09161.77011.77012.18162.50832.50833.49623.76463.76464.73135.66555.0039
H31.09321.77011.76952.18003.07582.51902.79732.56443.11294.20684.93694.8271
H41.09321.77011.76952.18002.51903.07582.79733.11292.56444.20684.93694.8271
C51.53942.18162.18002.18001.09091.09091.53842.16292.16292.56293.54492.8747
H62.16652.50833.07582.51901.09091.76072.15723.05932.50212.80173.81702.8226
H72.16652.50832.51903.07581.09091.76072.15722.50213.05932.80173.81702.8226
C82.54463.49622.79732.79731.53842.15722.15721.09731.09731.49992.21792.4181
H92.76133.76462.56443.11292.16293.05932.50211.09731.75982.11792.53513.1128
H102.76133.76463.11292.56442.16292.50213.05931.09731.75982.11792.53513.1128
C113.90974.73134.20684.20682.56292.80172.80171.49992.11792.11791.08271.2264
H124.75795.66554.93694.93693.54493.81703.81702.21792.53512.53511.08271.9792
O134.39495.00394.82714.82712.87472.82262.82262.41813.11283.11281.22641.9792

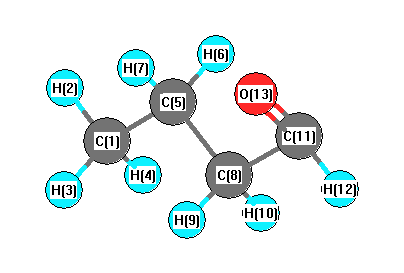
picture of Butanal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 109.730 C1 C5 H7 109.730
C1 C5 C8 111.539 H2 C1 H3 108.230
H2 C1 H4 108.230 H2 C1 C5 110.883
H3 C1 H4 108.063 H3 C1 C5 110.667
H4 C1 C5 110.667 C5 C8 H9 109.152
C5 C8 H10 109.152 C5 C8 C11 115.031
H6 C5 H7 107.602 H6 C5 C8 109.075
H7 C5 C8 109.075 C8 C11 H12 117.453
C8 C11 O13 124.691 H9 C8 H10 106.627
H9 C8 C11 108.276 H10 C8 C11 108.276
H12 C11 O13 117.856
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.465      
2 H 0.159      
3 H 0.149      
4 H 0.149      
5 C -0.268      
6 H 0.165      
7 H 0.165      
8 C -0.378      
9 H 0.171      
10 H 0.171      
11 C 0.285      
12 H 0.127      
13 O -0.430      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.420 -0.055 0.000
y -0.055 6.853 0.000
z 0.000 0.000 5.265


<r2> (average value of r2) Å2
<r2> 154.941
(<r2>)1/2 12.448