return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9N (Pyrrolidine)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-211.757618
Energy at 298.15K-211.769287
HF Energy-211.145032
Nuclear repulsion energy191.070090
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3661 3382 0.65      
2 A' 3248 3000 63.74      
3 A' 3215 2969 7.59      
4 A' 3196 2952 43.64      
5 A' 3083 2847 123.58      
6 A' 1633 1509 0.68      
7 A' 1607 1484 3.64      
8 A' 1490 1377 2.37      
9 A' 1409 1301 1.88      
10 A' 1329 1228 1.04      
11 A' 1306 1206 10.15      
12 A' 1137 1050 0.50      
13 A' 1062 981 5.40      
14 A' 1006 929 8.42      
15 A' 961 887 12.87      
16 A' 943 871 60.76      
17 A' 816 754 21.61      
18 A' 611 564 44.98      
19 A' 309 285 4.81      
20 A" 3228 2982 3.73      
21 A" 3209 2964 68.74      
22 A" 3185 2942 17.44      
23 A" 3077 2843 36.06      
24 A" 1615 1492 2.12      
25 A" 1584 1463 0.76      
26 A" 1530 1413 8.37      
27 A" 1414 1306 11.27      
28 A" 1392 1286 18.63      
29 A" 1325 1224 5.73      
30 A" 1278 1180 0.44      
31 A" 1210 1117 10.09      
32 A" 1174 1085 0.41      
33 A" 992 916 0.17      
34 A" 922 851 2.31      
35 A" 664 613 0.80      
36 A" 64 59 0.20      

Unscaled Zero Point Vibrational Energy (zpe) 29941.0 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 27656.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.23151 0.22924 0.13138

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.533 -1.091 0.000
H2 0.416 -2.094 0.000
C3 -0.102 -0.467 1.153
C4 -0.102 -0.467 -1.153
C5 -0.102 1.018 0.773
C6 -0.102 1.018 -0.773
H7 -1.133 -0.814 1.304
H8 -1.133 -0.814 -1.304
H9 0.456 -0.674 2.064
H10 0.456 -0.674 -2.064
H11 0.793 1.504 1.154
H12 0.793 1.504 -1.154
H13 -0.960 1.541 1.192
H14 -0.960 1.541 -1.192

Atom - Atom Distances (Å)
  N1 H2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
N11.01031.45691.45692.33442.33442.13402.13402.10722.10722.85132.85133.25193.2519
H21.01032.06082.06083.24873.24872.39562.39562.50592.50593.79723.79724.06544.0654
C31.45692.06082.30551.53262.43171.09852.68681.08863.27152.16433.16302.18313.2035
C41.45692.06082.30552.43171.53262.68681.09853.27151.08863.16302.16433.20352.1831
C52.33443.24871.53262.43171.54592.16872.95542.20033.35011.08772.17961.08822.2068
C62.33443.24872.43171.53261.54592.95542.16873.35012.20032.17961.08772.20681.0882
H72.13402.39561.09852.68682.16872.95542.60781.76733.72673.01803.88922.36403.4360
H82.13402.39562.68681.09852.95542.16872.60783.72671.76733.88923.01803.43602.3640
H92.10722.50591.08863.27152.20033.35011.76733.72674.12812.38403.90002.76934.1846
H102.10722.50593.27151.08863.35012.20033.72671.76734.12813.90002.38404.18462.7693
H112.85133.79722.16433.16301.08772.17963.01803.88922.38403.90002.30771.75432.9291
H122.85133.79723.16302.16432.17961.08773.88923.01803.90002.38402.30772.92911.7543
H133.25194.06542.18313.20351.08822.20682.36403.43602.76934.18461.75432.92912.3841
H143.25194.06543.20352.18312.20681.08823.43602.36404.18462.76932.92911.75432.3841

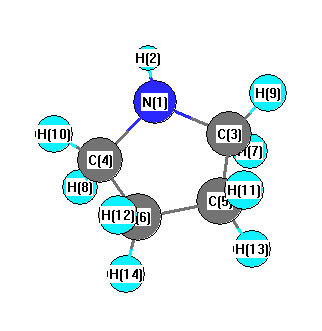
picture of Pyrrolidine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C3 C5 102.650 N1 C3 H7 112.497
N1 C3 H9 110.921 N1 C4 C6 102.650
N1 C4 H8 112.497 N1 C4 H10 110.921
H2 N1 C3 112.023 H2 N1 C4 112.023
C3 N1 C4 104.606 C3 C5 C6 104.348
C3 C5 H11 110.219 C3 C5 H13 111.692
C4 C6 C5 104.348 C4 C6 H12 110.219
C4 C6 H14 111.692 C5 C3 H7 109.931
C5 C3 H9 113.068 C5 C6 H12 110.499
C5 C6 H14 112.650 C6 C4 H8 109.931
C6 C4 H10 113.068 C6 C5 H11 110.499
C6 C5 H13 112.650 H7 C3 H9 107.812
H8 C4 H10 107.812 H11 C5 H13 107.458
H12 C6 H14 107.458
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability