return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9N (Pyrrolidine)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-211.869016
Energy at 298.15K-211.880603
HF Energy-211.144142
Nuclear repulsion energy190.197969
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3560 3342 0.65      
2 A' 3186 2991 54.79      
3 A' 3152 2958 7.91      
4 A' 3134 2942 38.18      
5 A' 3017 2831 113.76      
6 A' 1594 1496 0.80      
7 A' 1570 1474 4.13      
8 A' 1446 1357 1.97      
9 A' 1368 1284 1.76      
10 A' 1290 1211 0.60      
11 A' 1271 1193 9.18      
12 A' 1112 1044 0.74      
13 A' 1041 977 6.93      
14 A' 981 921 3.06      
15 A' 940 883 13.81      
16 A' 926 869 59.55      
17 A' 799 750 19.98      
18 A' 599 562 42.26      
19 A' 307 288 4.94      
20 A" 3168 2973 1.79      
21 A" 3147 2954 60.96      
22 A" 3126 2934 15.47      
23 A" 3014 2829 32.96      
24 A" 1576 1479 1.29      
25 A" 1549 1454 0.89      
26 A" 1485 1394 7.41      
27 A" 1372 1288 11.36      
28 A" 1347 1264 14.29      
29 A" 1288 1209 5.69      
30 A" 1240 1164 0.45      
31 A" 1174 1102 9.62      
32 A" 1142 1072 0.26      
33 A" 969 910 0.24      
34 A" 899 844 2.30      
35 A" 646 606 0.71      
36 A" 64 60 0.21      

Unscaled Zero Point Vibrational Energy (zpe) 29249.1 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 27453.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.22949 0.22707 0.13044

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.546 -1.093 0.000
H2 0.416 -2.102 0.000
C3 -0.105 -0.469 1.156
C4 -0.105 -0.469 -1.156
C5 -0.105 1.022 0.776
C6 -0.105 1.022 -0.776
H7 -1.142 -0.819 1.299
H8 -1.142 -0.819 -1.299
H9 0.451 -0.677 2.076
H10 0.451 -0.677 -2.076
H11 0.796 1.509 1.159
H12 0.796 1.509 -1.159
H13 -0.968 1.546 1.197
H14 -0.968 1.546 -1.197

Atom - Atom Distances (Å)
  N1 H2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
N11.01781.46641.46642.34472.34472.14792.14792.11892.11892.85902.85903.26913.2691
H21.01782.06812.06813.26103.26102.40062.40062.51792.51793.81143.81144.08124.0812
C31.46642.06812.31271.53832.44051.10452.68841.09433.28602.17293.17532.19193.2158
C41.46642.06812.31272.44051.53832.68841.10453.28601.09433.17532.17293.21582.1919
C52.34473.26101.53832.44051.55212.17702.96162.21023.36571.09322.18931.09372.2164
C62.34473.26102.44051.53831.55212.96162.17703.36572.21022.18931.09322.21641.0937
H72.14792.40061.10452.68842.17702.96162.59771.77833.73443.03253.90102.37363.4429
H82.14792.40062.68841.10452.96162.17702.59773.73441.77833.90103.03253.44292.3736
H92.11892.51791.09433.28602.21023.36571.77833.73444.15122.39543.91932.77964.2031
H102.11892.51793.28601.09433.36572.21023.73441.77834.15123.91932.39544.20312.7796
H112.85903.81142.17293.17531.09322.18933.03253.90102.39543.91932.31821.76442.9434
H122.85903.81143.17532.17292.18931.09323.90103.03253.91932.39542.31822.94341.7644
H133.26914.08122.19193.21581.09372.21642.37363.44292.77964.20311.76442.94342.3943
H143.26914.08123.21582.19192.21641.09373.44292.37364.20312.77962.94341.76442.3943

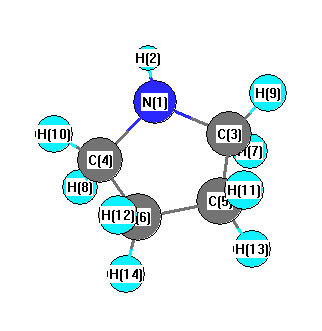
picture of Pyrrolidine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C3 C5 102.558 N1 C3 H7 112.570
N1 C3 H9 110.844 N1 C4 C6 102.558
N1 C4 H8 112.570 N1 C4 H10 110.844
H2 N1 C3 111.434 H2 N1 C4 111.434
C3 N1 C4 104.099 C3 C5 C6 104.312
C3 C5 H11 110.176 C3 C5 H13 111.652
C4 C6 C5 104.312 C4 C6 H12 110.176
C4 C6 H14 111.652 C5 C3 H7 109.837
C5 C3 H9 113.104 C5 C6 H12 110.510
C5 C6 H14 112.644 C6 C4 H8 109.837
C6 C4 H10 113.104 C6 C5 H11 110.510
C6 C5 H13 112.644 H7 C3 H9 107.955
H8 C4 H10 107.955 H11 C5 H13 107.564
H12 C6 H14 107.564
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability