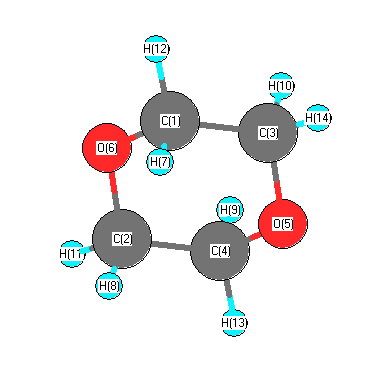
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3120 |
2985 |
0.00 |
|
|
|
| 2 |
Ag |
3001 |
2871 |
0.00 |
|
|
|
| 3 |
Ag |
1517 |
1452 |
0.00 |
|
|
|
| 4 |
Ag |
1444 |
1381 |
0.00 |
|
|
|
| 5 |
Ag |
1346 |
1288 |
0.00 |
|
|
|
| 6 |
Ag |
1157 |
1107 |
0.00 |
|
|
|
| 7 |
Ag |
1043 |
998 |
0.00 |
|
|
|
| 8 |
Ag |
867 |
829 |
0.00 |
|
|
|
| 9 |
Ag |
438 |
419 |
0.00 |
|
|
|
| 10 |
Ag |
419 |
401 |
0.00 |
|
|
|
| 11 |
Au |
3118 |
2983 |
93.64 |
|
|
|
| 12 |
Au |
2994 |
2864 |
48.50 |
|
|
|
| 13 |
Au |
1503 |
1438 |
0.13 |
|
|
|
| 14 |
Au |
1409 |
1348 |
28.02 |
|
|
|
| 15 |
Au |
1298 |
1242 |
47.90 |
|
|
|
| 16 |
Au |
1178 |
1127 |
175.40 |
|
|
|
| 17 |
Au |
1121 |
1072 |
3.15 |
|
|
|
| 18 |
Au |
908 |
869 |
15.98 |
|
|
|
| 19 |
Au |
251 |
240 |
1.20 |
|
|
|
| 20 |
Bg |
3118 |
2983 |
0.00 |
|
|
|
| 21 |
Bg |
3005 |
2875 |
0.00 |
|
|
|
| 22 |
Bg |
1501 |
1436 |
0.00 |
|
|
|
| 23 |
Bg |
1372 |
1312 |
0.00 |
|
|
|
| 24 |
Bg |
1252 |
1198 |
0.00 |
|
|
|
| 25 |
Bg |
1170 |
1119 |
0.00 |
|
|
|
| 26 |
Bg |
875 |
837 |
0.00 |
|
|
|
| 27 |
Bg |
488 |
467 |
0.00 |
|
|
|
| 28 |
Bu |
3119 |
2984 |
57.98 |
|
|
|
| 29 |
Bu |
3010 |
2880 |
173.50 |
|
|
|
| 30 |
Bu |
1511 |
1446 |
11.71 |
|
|
|
| 31 |
Bu |
1429 |
1367 |
5.12 |
|
|
|
| 32 |
Bu |
1333 |
1275 |
10.28 |
|
|
|
| 33 |
Bu |
1077 |
1031 |
10.08 |
|
|
|
| 34 |
Bu |
915 |
875 |
66.75 |
|
|
|
| 35 |
Bu |
609 |
583 |
13.03 |
|
|
|
| 36 |
Bu |
270 |
259 |
18.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27092.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 25919.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.094 |
|
|
0.060 |
| 2 |
C |
-0.094 |
|
|
0.060 |
| 3 |
C |
-0.094 |
|
|
0.052 |
| 4 |
C |
-0.094 |
|
|
0.052 |
| 5 |
O |
-0.464 |
|
|
-0.350 |
| 6 |
O |
-0.464 |
|
|
-0.354 |
| 7 |
H |
0.154 |
|
|
0.056 |
| 8 |
H |
0.154 |
|
|
0.056 |
| 9 |
H |
0.154 |
|
|
0.057 |
| 10 |
H |
0.154 |
|
|
0.057 |
| 11 |
H |
0.173 |
|
|
0.063 |
| 12 |
H |
0.173 |
|
|
0.063 |
| 13 |
H |
0.173 |
|
|
0.064 |
| 14 |
H |
0.173 |
|
|
0.064 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.001 |
0.007 |
0.000 |
0.007 |
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
139.074 |
| (<r2>)1/2 |
11.793 |