return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (1,4-Dioxane)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-307.325363
Energy at 298.15K-307.336612
HF Energy-307.038484
Nuclear repulsion energy264.157799
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3146 2986 0.00      
2 Ag 3036 2882 0.00      
3 Ag 1543 1465 0.00      
4 Ag 1460 1386 0.00      
5 Ag 1354 1285 0.00      
6 Ag 1175 1115 0.00      
7 Ag 1043 990 0.00      
8 Ag 863 819 0.00      
9 Ag 441 419 0.00      
10 Ag 426 405 0.00      
11 Au 3144 2984 88.17      
12 Au 3029 2876 45.09      
13 Au 1529 1451 0.00      
14 Au 1425 1353 30.42      
15 Au 1307 1241 37.69      
16 Au 1173 1114 165.95      
17 Au 1135 1077 11.57      
18 Au 914 867 21.78      
19 Au 255 243 1.27      
20 Bg 3145 2985 0.00      
21 Bg 3038 2884 0.00      
22 Bg 1527 1449 0.00      
23 Bg 1388 1318 0.00      
24 Bg 1261 1197 0.00      
25 Bg 1163 1104 0.00      
26 Bg 884 839 0.00      
27 Bg 491 466 0.00      
28 Bu 3146 2986 59.91      
29 Bu 3043 2889 156.43      
30 Bu 1537 1459 8.92      
31 Bu 1446 1372 6.09      
32 Bu 1341 1273 9.29      
33 Bu 1083 1028 10.23      
34 Bu 905 859 64.48      
35 Bu 619 588 14.12      
36 Bu 275 261 19.99      

Unscaled Zero Point Vibrational Energy (zpe) 27344.9 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 25955.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.16908 0.15668 0.09151

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.765 1.163
C2 0.000 0.765 -1.163
C3 0.000 -0.765 1.163
C4 0.000 -0.765 -1.163
O5 -0.662 -1.257 0.000
O6 0.662 1.257 0.000
H7 -1.035 1.121 1.186
H8 -1.035 1.121 -1.186
H9 1.035 -1.121 -1.186
H10 1.035 -1.121 1.186
H11 0.527 1.154 -2.039
H12 0.527 1.154 2.039
H13 -0.527 -1.154 -2.039
H14 -0.527 -1.154 2.039

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12 H13 H14
C12.32541.53092.78412.42461.42521.09492.59153.18542.15183.26801.09403.77012.1749
C22.32542.78411.53092.42461.42522.59151.09492.15183.18541.09403.26802.17493.7701
C31.53092.78412.32541.42522.42462.15183.18542.59151.09493.77012.17493.26801.0940
C42.78411.53092.32541.42522.42463.18542.15181.09492.59152.17493.77011.09403.2680
O52.42462.42461.42521.42522.84062.68312.68312.07512.07513.37393.37392.04602.0460
O61.42521.42522.42462.42462.84062.07512.07512.68312.68312.04602.04603.37393.3739
H71.09492.59152.15183.18542.68312.07512.37243.86533.05163.58401.78053.97932.4820
H82.59151.09493.18542.15182.68312.07512.37243.05163.86531.78053.58402.48203.9793
H93.18542.15182.59151.09492.07512.68313.86533.05162.37242.48203.97931.78053.5840
H102.15183.18541.09492.59152.07512.68313.05163.86532.37243.97932.48203.58401.7805
H113.26801.09403.77012.17493.37392.04603.58401.78052.48203.97934.07802.53764.8031
H121.09403.26802.17493.77013.37392.04601.78053.58403.97932.48204.07804.80312.5376
H133.77012.17493.26801.09402.04603.37393.97932.48201.78053.58402.53764.80314.0780
H142.17493.77011.09403.26802.04603.37392.48203.97933.58401.78054.80312.53764.0780

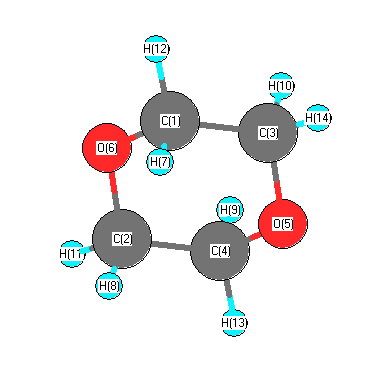
picture of 1,4-Dioxane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 O5 110.165 C1 C3 H10 108.934
C1 C3 H14 110.810 C1 O6 C2 109.338
C2 C4 O5 110.165 C2 C4 H9 108.934
C2 C4 H13 110.810 C3 C1 O6 110.165
C3 C1 H7 108.934 C3 C1 H12 110.810
C3 O5 C4 109.338 C4 C2 O6 110.165
C4 C2 H8 108.934 C4 C2 H11 110.810
O5 C3 H10 110.166 O5 C3 H14 107.892
O5 C4 H9 110.166 O5 C4 H13 107.892
O6 C1 H7 110.166 O6 C1 H12 107.892
O6 C2 H8 110.166 O6 C2 H11 107.892
H7 C1 H12 108.859 H8 C2 H11 108.859
H9 C4 H13 108.859 H10 C3 H14 108.859
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.034      
2 C -0.034      
3 C -0.034      
4 C -0.034      
5 O -0.539      
6 O -0.539      
7 H 0.143      
8 H 0.143      
9 H 0.143      
10 H 0.143      
11 H 0.160      
12 H 0.160      
13 H 0.160      
14 H 0.160      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 6.257 -0.105 0.000
y -0.105 6.381 0.000
z 0.000 0.000 8.161


<r2> (average value of r2) Å2
<r2> 139.652
(<r2>)1/2 11.817