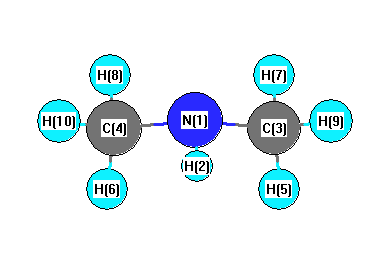
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3495 |
3356 |
1.54 |
|
|
|
| 2 |
A' |
3117 |
2993 |
41.71 |
|
|
|
| 3 |
A' |
3069 |
2947 |
54.90 |
|
|
|
| 4 |
A' |
2945 |
2828 |
161.82 |
|
|
|
| 5 |
A' |
1548 |
1487 |
0.76 |
|
|
|
| 6 |
A' |
1525 |
1465 |
8.81 |
|
|
|
| 7 |
A' |
1496 |
1437 |
1.04 |
|
|
|
| 8 |
A' |
1284 |
1233 |
0.28 |
|
|
|
| 9 |
A' |
1203 |
1155 |
6.93 |
|
|
|
| 10 |
A' |
956 |
918 |
1.25 |
|
|
|
| 11 |
A' |
796 |
765 |
125.08 |
|
|
|
| 12 |
A' |
387 |
371 |
6.13 |
|
|
|
| 13 |
A' |
272 |
261 |
2.76 |
|
|
|
| 14 |
A" |
3116 |
2993 |
28.86 |
|
|
|
| 15 |
A" |
3068 |
2946 |
28.33 |
|
|
|
| 16 |
A" |
2938 |
2821 |
45.80 |
|
|
|
| 17 |
A" |
1545 |
1483 |
11.12 |
|
|
|
| 18 |
A" |
1513 |
1453 |
5.47 |
|
|
|
| 19 |
A" |
1501 |
1441 |
6.64 |
|
|
|
| 20 |
A" |
1467 |
1408 |
3.46 |
|
|
|
| 21 |
A" |
1184 |
1137 |
33.33 |
|
|
|
| 22 |
A" |
1112 |
1068 |
0.09 |
|
|
|
| 23 |
A" |
1046 |
1005 |
9.98 |
|
|
|
| 24 |
A" |
246 |
236 |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20413.8 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 19603.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.524 |
0.349 |
0.337 |
0.351 |
| 2 |
H |
0.294 |
-0.698 |
-1.004 |
-0.644 |
| 3 |
C |
-0.300 |
0.011 |
0.016 |
0.046 |
| 4 |
C |
-0.300 |
0.038 |
0.014 |
0.077 |
| 5 |
H |
0.117 |
-0.013 |
-0.015 |
0.027 |
| 6 |
H |
0.117 |
0.139 |
0.319 |
-0.009 |
| 7 |
H |
0.151 |
0.038 |
0.014 |
0.067 |
| 8 |
H |
0.151 |
0.011 |
0.016 |
0.034 |
| 9 |
H |
0.147 |
-0.013 |
-0.015 |
0.015 |
| 10 |
H |
0.147 |
0.139 |
0.319 |
0.035 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.961 |
-0.361 |
0.000 |
1.027 |
| CHELPG |
0.000 |
-0.182 |
0.998 |
1.015 |
| AIM |
0.000 |
0.433 |
-0.137 |
0.454 |
| ESP |
-0.004 |
-0.201 |
0.989 |
1.009 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.568 |
0.000 |
0.000 |
| y |
0.000 |
-20.230 |
2.034 |
| z |
0.000 |
2.034 |
-21.515 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.305 |
0.000 |
0.000 |
| y |
0.000 |
0.312 |
2.034 |
| z |
0.000 |
2.034 |
-1.617 |
|
| Polar |
|---|
| 3z2-r2 | -3.233 |
|---|
| x2-y2 | 0.662 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 2.034 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.053 |
-0.000 |
0.000 |
| y |
-0.000 |
4.314 |
0.083 |
| z |
0.000 |
0.083 |
3.989 |
<r2> (average value of r
2) Å
2
| <r2> |
57.292 |
| (<r2>)1/2 |
7.569 |