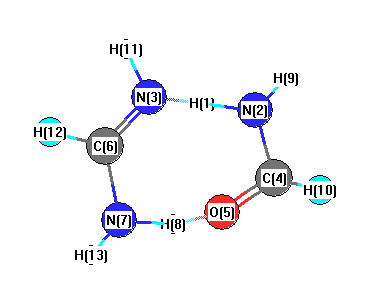
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3751 |
3564 |
90.99 |
|
|
|
| 2 |
A |
3705 |
3521 |
60.68 |
|
|
|
| 3 |
A |
3578 |
3400 |
3.04 |
|
|
|
| 4 |
A |
3370 |
3202 |
853.03 |
|
|
|
| 5 |
A |
3131 |
2975 |
712.02 |
|
|
|
| 6 |
A |
3098 |
2944 |
215.16 |
|
|
|
| 7 |
A |
3007 |
2857 |
237.34 |
|
|
|
| 8 |
A |
1841 |
1750 |
576.80 |
|
|
|
| 9 |
A |
1784 |
1695 |
119.12 |
|
|
|
| 10 |
A |
1706 |
1621 |
7.12 |
|
|
|
| 11 |
A |
1682 |
1598 |
5.89 |
|
|
|
| 12 |
A |
1465 |
1392 |
28.24 |
|
|
|
| 13 |
A |
1447 |
1375 |
14.13 |
|
|
|
| 14 |
A |
1392 |
1323 |
14.17 |
|
|
|
| 15 |
A |
1383 |
1315 |
107.91 |
|
|
|
| 16 |
A |
1180 |
1121 |
67.57 |
|
|
|
| 17 |
A |
1140 |
1084 |
1.09 |
|
|
|
| 18 |
A |
1124 |
1068 |
6.91 |
|
|
|
| 19 |
A |
1081 |
1028 |
15.67 |
|
|
|
| 20 |
A |
1064 |
1011 |
0.24 |
|
|
|
| 21 |
A |
934 |
888 |
106.34 |
|
|
|
| 22 |
A |
833 |
792 |
14.12 |
|
|
|
| 23 |
A |
759 |
721 |
164.36 |
|
|
|
| 24 |
A |
660 |
627 |
11.36 |
|
|
|
| 25 |
A |
597 |
567 |
7.40 |
|
|
|
| 26 |
A |
513 |
487 |
113.81 |
|
|
|
| 27 |
A |
333 |
317 |
62.48 |
|
|
|
| 28 |
A |
253 |
240 |
84.38 |
|
|
|
| 29 |
A |
198 |
188 |
1.24 |
|
|
|
| 30 |
A |
166 |
158 |
0.41 |
|
|
|
| 31 |
A |
155 |
147 |
20.39 |
|
|
|
| 32 |
A |
76 |
72 |
84.22 |
|
|
|
| 33 |
A |
10i |
9i |
47.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23697.0 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 22519.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.438 |
|
|
|
| 2 |
N |
-0.780 |
|
|
|
| 3 |
N |
-0.688 |
|
|
|
| 4 |
C |
0.351 |
|
|
|
| 5 |
O |
-0.502 |
|
|
|
| 6 |
C |
0.253 |
|
|
|
| 7 |
N |
-0.788 |
|
|
|
| 8 |
H |
0.424 |
|
|
|
| 9 |
H |
0.346 |
|
|
|
| 10 |
H |
0.130 |
|
|
|
| 11 |
H |
0.311 |
|
|
|
| 12 |
H |
0.151 |
|
|
|
| 13 |
H |
0.353 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.391 |
1.513 |
-0.002 |
2.055 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.723 |
7.093 |
0.129 |
| y |
7.093 |
-31.716 |
0.070 |
| z |
0.129 |
0.070 |
-38.843 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
11.557 |
7.093 |
0.129 |
| y |
7.093 |
-0.433 |
0.070 |
| z |
0.129 |
0.070 |
-11.124 |
|
| Polar |
|---|
| 3z2-r2 | -22.247 |
|---|
| x2-y2 | 7.993 |
|---|
| xy | 7.093 |
|---|
| xz | 0.129 |
|---|
| yz | 0.070 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.410 |
0.490 |
0.018 |
| y |
0.490 |
8.909 |
0.001 |
| z |
0.018 |
0.001 |
2.915 |
<r2> (average value of r
2) Å
2
| <r2> |
215.236 |
| (<r2>)1/2 |
14.671 |