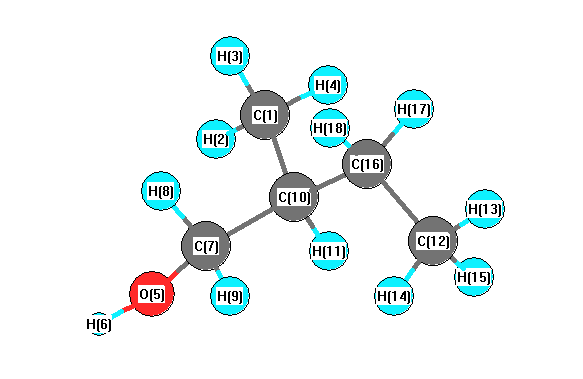
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3797 |
3633 |
17.78 |
|
|
|
| 2 |
A |
3159 |
3022 |
16.92 |
|
|
|
| 3 |
A |
3134 |
2998 |
47.56 |
|
|
|
| 4 |
A |
3127 |
2991 |
39.69 |
|
|
|
| 5 |
A |
3117 |
2982 |
37.35 |
|
|
|
| 6 |
A |
3074 |
2941 |
21.61 |
|
|
|
| 7 |
A |
3056 |
2923 |
43.14 |
|
|
|
| 8 |
A |
3050 |
2918 |
57.58 |
|
|
|
| 9 |
A |
3041 |
2909 |
36.00 |
|
|
|
| 10 |
A |
3031 |
2899 |
9.32 |
|
|
|
| 11 |
A |
3023 |
2892 |
37.04 |
|
|
|
| 12 |
A |
2976 |
2847 |
54.17 |
|
|
|
| 13 |
A |
1546 |
1479 |
2.28 |
|
|
|
| 14 |
A |
1536 |
1469 |
6.04 |
|
|
|
| 15 |
A |
1532 |
1465 |
9.53 |
|
|
|
| 16 |
A |
1527 |
1461 |
5.17 |
|
|
|
| 17 |
A |
1520 |
1454 |
7.14 |
|
|
|
| 18 |
A |
1510 |
1444 |
1.38 |
|
|
|
| 19 |
A |
1475 |
1411 |
3.70 |
|
|
|
| 20 |
A |
1438 |
1376 |
3.65 |
|
|
|
| 21 |
A |
1427 |
1365 |
6.31 |
|
|
|
| 22 |
A |
1406 |
1345 |
1.03 |
|
|
|
| 23 |
A |
1379 |
1319 |
4.18 |
|
|
|
| 24 |
A |
1342 |
1284 |
2.35 |
|
|
|
| 25 |
A |
1315 |
1258 |
13.54 |
|
|
|
| 26 |
A |
1282 |
1227 |
6.17 |
|
|
|
| 27 |
A |
1262 |
1208 |
29.94 |
|
|
|
| 28 |
A |
1198 |
1146 |
1.22 |
|
|
|
| 29 |
A |
1184 |
1133 |
4.84 |
|
|
|
| 30 |
A |
1140 |
1091 |
0.79 |
|
|
|
| 31 |
A |
1099 |
1051 |
87.95 |
|
|
|
| 32 |
A |
1060 |
1014 |
2.72 |
|
|
|
| 33 |
A |
1035 |
990 |
15.47 |
|
|
|
| 34 |
A |
986 |
943 |
7.14 |
|
|
|
| 35 |
A |
936 |
896 |
1.95 |
|
|
|
| 36 |
A |
926 |
886 |
1.81 |
|
|
|
| 37 |
A |
834 |
798 |
4.00 |
|
|
|
| 38 |
A |
769 |
736 |
5.90 |
|
|
|
| 39 |
A |
496 |
475 |
6.64 |
|
|
|
| 40 |
A |
444 |
425 |
1.57 |
|
|
|
| 41 |
A |
389 |
372 |
2.27 |
|
|
|
| 42 |
A |
278 |
266 |
6.53 |
|
|
|
| 43 |
A |
271 |
259 |
70.90 |
|
|
|
| 44 |
A |
250 |
239 |
39.86 |
|
|
|
| 45 |
A |
235 |
224 |
3.34 |
|
|
|
| 46 |
A |
222 |
212 |
4.59 |
|
|
|
| 47 |
A |
118 |
113 |
3.99 |
|
|
|
| 48 |
A |
76 |
73 |
4.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36512.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 34931.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.512 |
|
|
-0.493 |
| 2 |
H |
0.194 |
|
|
0.151 |
| 3 |
H |
0.154 |
|
|
0.108 |
| 4 |
H |
0.160 |
|
|
0.100 |
| 5 |
O |
-0.631 |
|
|
-0.631 |
| 6 |
H |
0.405 |
|
|
0.419 |
| 7 |
C |
-0.068 |
|
|
0.055 |
| 8 |
H |
0.132 |
|
|
0.029 |
| 9 |
H |
0.142 |
|
|
0.025 |
| 10 |
C |
-0.131 |
|
|
0.295 |
| 11 |
H |
0.162 |
|
|
0.020 |
| 12 |
C |
-0.515 |
|
|
-0.281 |
| 13 |
H |
0.168 |
|
|
0.089 |
| 14 |
H |
0.166 |
|
|
0.067 |
| 15 |
H |
0.168 |
|
|
0.068 |
| 16 |
C |
-0.308 |
|
|
-0.119 |
| 17 |
H |
0.162 |
|
|
0.056 |
| 18 |
H |
0.153 |
|
|
0.041 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.395 |
-1.016 |
0.999 |
1.479 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.399 |
-1.016 |
1.002 |
1.482 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.339 |
3.236 |
-2.512 |
| y |
3.236 |
-36.867 |
-1.590 |
| z |
-2.512 |
-1.590 |
-39.517 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.147 |
3.236 |
-2.512 |
| y |
3.236 |
2.561 |
-1.590 |
| z |
-2.512 |
-1.590 |
-1.414 |
|
| Polar |
|---|
| 3z2-r2 | -2.828 |
|---|
| x2-y2 | -2.472 |
|---|
| xy | 3.236 |
|---|
| xz | -2.512 |
|---|
| yz | -1.590 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
215.534 |
| (<r2>)1/2 |
14.681 |